Jump to
S1C2
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -958.626174 |
| Energy at 298.15K | |
| HF Energy | -958.626174 |
| Nuclear repulsion energy | 124.631974 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3187 |
3169 |
0.23 |
|
|
|
| 2 |
A1 |
740 |
736 |
8.98 |
|
|
|
| 3 |
A1 |
302 |
301 |
0.25 |
|
|
|
| 4 |
B1 |
368i |
365i |
52.94 |
|
|
|
| 5 |
B2 |
1167 |
1160 |
44.55 |
|
|
|
| 6 |
B2 |
903 |
897 |
158.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 2965.4 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 2948.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.675 |
| H2 |
0.000 |
0.000 |
1.770 |
| Cl3 |
0.000 |
1.489 |
-0.171 |
| Cl4 |
0.000 |
-1.489 |
-0.171 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
Cl3 |
Cl4 |
| C1 | | 1.0958 | 1.7126 | 1.7126 |
H2 | 1.0958 | | 2.4469 | 2.4469 | Cl3 | 1.7126 | 2.4469 | | 2.9784 | Cl4 | 1.7126 | 2.4469 | 2.9784 | |
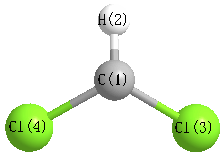 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl3 |
C1 |
H2 |
119.592 |
|
Cl3 |
C1 |
Cl4 |
120.816 |
| Cl4 |
C1 |
H2 |
119.592 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.153 |
|
|
|
| 2 |
H |
0.109 |
|
|
|
| 3 |
Cl |
0.022 |
|
|
|
| 4 |
Cl |
0.022 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.787 |
0.787 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.642 |
0.000 |
0.000 |
| y |
0.000 |
-30.269 |
0.000 |
| z |
0.000 |
0.000 |
-28.734 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.140 |
0.000 |
0.000 |
| y |
0.000 |
-0.081 |
0.000 |
| z |
0.000 |
0.000 |
2.222 |
|
| Polar |
|---|
| 3z2-r2 | 4.443 |
|---|
| x2-y2 | -1.373 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.258 |
0.000 |
0.000 |
| y |
0.000 |
6.941 |
0.000 |
| z |
0.000 |
0.000 |
3.731 |
<r2> (average value of r
2) Å
2
| <r2> |
101.135 |
| (<r2>)1/2 |
10.057 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -958.627162 |
| Energy at 298.15K | -958.627927 |
| HF Energy | -958.627162 |
| Nuclear repulsion energy | 124.428867 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3138 |
3119 |
1.67 |
|
|
|
| 2 |
A' |
745 |
741 |
14.93 |
|
|
|
| 3 |
A' |
483 |
480 |
26.15 |
|
|
|
| 4 |
A' |
297 |
295 |
0.46 |
|
|
|
| 5 |
A" |
1171 |
1164 |
34.20 |
|
|
|
| 6 |
A" |
865 |
860 |
185.79 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3348.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 3329.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.013 |
0.703 |
0.000 |
| H2 |
-0.528 |
1.660 |
0.000 |
| Cl3 |
0.013 |
-0.173 |
1.485 |
| Cl4 |
0.013 |
-0.173 |
-1.485 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
Cl3 |
Cl4 |
| C1 | | 1.0999 | 1.7240 | 1.7240 |
H2 | 1.0999 | | 2.4205 | 2.4205 | Cl3 | 1.7240 | 2.4205 | | 2.9700 | Cl4 | 1.7240 | 2.4205 | 2.9700 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl3 |
C1 |
H2 |
116.244 |
|
Cl3 |
C1 |
Cl4 |
118.936 |
| Cl4 |
C1 |
H2 |
116.244 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.145 |
|
|
|
| 2 |
H |
0.115 |
|
|
|
| 3 |
Cl |
0.015 |
|
|
|
| 4 |
Cl |
0.015 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.537 |
0.679 |
0.000 |
0.866 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.317 |
-0.941 |
0.000 |
| y |
-0.941 |
-29.237 |
0.000 |
| z |
0.000 |
0.000 |
-30.442 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.478 |
-0.941 |
0.000 |
| y |
-0.941 |
1.643 |
0.000 |
| z |
0.000 |
0.000 |
-0.165 |
|
| Polar |
|---|
| 3z2-r2 | -0.331 |
|---|
| x2-y2 | -2.080 |
|---|
| xy | -0.941 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.387 |
-0.215 |
0.000 |
| y |
-0.215 |
3.753 |
0.000 |
| z |
0.000 |
0.000 |
7.028 |
<r2> (average value of r
2) Å
2
| <r2> |
100.945 |
| (<r2>)1/2 |
10.047 |
