Jump to
S1C2
Energy calculated at ROHF/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -957.396410 |
| Energy at 298.15K | |
| HF Energy | -957.396410 |
| Nuclear repulsion energy | 125.257126 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at ROHF/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3446 |
2966 |
0.55 |
|
|
|
| 2 |
A1 |
786 |
676 |
14.98 |
|
|
|
| 3 |
A1 |
324 |
279 |
0.87 |
|
|
|
| 4 |
B1 |
645i |
555i |
40.14 |
|
|
|
| 5 |
B2 |
1314 |
1131 |
73.48 |
|
|
|
| 6 |
B2 |
986 |
848 |
129.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3105.2 cm
-1
Scaled (by 0.8607) Zero Point Vibrational Energy (zpe) 2672.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at ROHF/cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.681 |
| H2 |
0.000 |
0.000 |
1.756 |
| Cl3 |
0.000 |
1.479 |
-0.172 |
| Cl4 |
0.000 |
-1.479 |
-0.172 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
Cl3 |
Cl4 |
| C1 | | 1.0745 | 1.7077 | 1.7077 |
H2 | 1.0745 | | 2.4298 | 2.4298 | Cl3 | 1.7077 | 2.4298 | | 2.9590 | Cl4 | 1.7077 | 2.4298 | 2.9590 | |
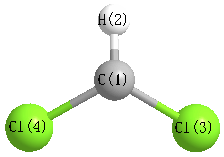 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl3 |
C1 |
H2 |
119.962 |
|
Cl3 |
C1 |
Cl4 |
120.075 |
| Cl4 |
C1 |
H2 |
119.962 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at ROHF/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.049 |
|
|
|
| 2 |
H |
0.129 |
|
|
|
| 3 |
Cl |
-0.040 |
|
|
|
| 4 |
Cl |
-0.040 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.228 |
1.228 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.015 |
0.000 |
0.000 |
| y |
0.000 |
-31.812 |
-0.000 |
| z |
0.000 |
-0.000 |
-28.841 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.688 |
0.000 |
0.000 |
| y |
0.000 |
-1.384 |
-0.000 |
| z |
0.000 |
-0.000 |
3.072 |
|
| Polar |
|---|
| 3z2-r2 | 6.145 |
|---|
| x2-y2 | -0.203 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
100.585 |
| (<r2>)1/2 |
10.029 |
Jump to
S1C1
Energy calculated at ROHF/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -957.399967 |
| Energy at 298.15K | -957.400947 |
| HF Energy | -957.399967 |
| Nuclear repulsion energy | 124.997423 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at ROHF/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3384 |
2913 |
3.17 |
|
|
|
| 2 |
A' |
814 |
701 |
23.16 |
|
|
|
| 3 |
A' |
705 |
607 |
8.58 |
|
|
|
| 4 |
A' |
319 |
275 |
0.57 |
|
|
|
| 5 |
A" |
1324 |
1140 |
54.51 |
|
|
|
| 6 |
A" |
935 |
805 |
175.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3740.5 cm
-1
Scaled (by 0.8607) Zero Point Vibrational Energy (zpe) 3219.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at ROHF/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.015 |
0.719 |
0.000 |
| H2 |
-0.597 |
1.609 |
0.000 |
| Cl3 |
0.015 |
-0.174 |
1.474 |
| Cl4 |
0.015 |
-0.174 |
-1.474 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
Cl3 |
Cl4 |
| C1 | | 1.0798 | 1.7232 | 1.7232 |
H2 | 1.0798 | | 2.3927 | 2.3927 | Cl3 | 1.7232 | 2.3927 | | 2.9472 | Cl4 | 1.7232 | 2.3927 | 2.9472 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl3 |
C1 |
H2 |
115.291 |
|
Cl3 |
C1 |
Cl4 |
117.557 |
| Cl4 |
C1 |
H2 |
115.291 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at ROHF/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.014 |
|
|
|
| 2 |
H |
0.127 |
|
|
|
| 3 |
Cl |
-0.057 |
|
|
|
| 4 |
Cl |
-0.057 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.545 |
1.160 |
0.000 |
1.281 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.592 |
-0.971 |
0.000 |
| y |
-0.971 |
-29.439 |
0.000 |
| z |
0.000 |
0.000 |
-32.145 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.800 |
-0.971 |
0.000 |
| y |
-0.971 |
2.430 |
0.000 |
| z |
0.000 |
0.000 |
-1.629 |
|
| Polar |
|---|
| 3z2-r2 | -3.258 |
|---|
| x2-y2 | -2.153 |
|---|
| xy | -0.971 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
100.318 |
| (<r2>)1/2 |
10.016 |
