return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SSSCH3 (dimethyl trisulfide)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-1272.582655
Energy at 298.15K-1272.588850
Nuclear repulsion energy361.636555
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3183 3054 7.86      
2 A 3170 3042 0.44      
3 A 3068 2944 5.36      
4 A 1482 1422 5.66      
5 A 1462 1402 6.40      
6 A 1347 1292 0.89      
7 A 981 941 0.06      
8 A 978 938 2.97      
9 A 719 690 0.15      
10 A 486 466 0.13      
11 A 270 259 0.15      
12 A 189 181 1.19      
13 A 160 153 0.96      
14 A 62 60 1.73      
15 B 3183 3053 4.19      
16 B 3170 3041 7.10      
17 B 3068 2943 38.44      
18 B 1481 1420 9.37      
19 B 1461 1402 7.25      
20 B 1350 1295 14.12      
21 B 981 941 1.44      
22 B 974 934 11.39      
23 B 717 688 1.39      
24 B 480 461 11.38      
25 B 275 264 0.74      
26 B 183 176 0.70      
27 B 96 92 3.12      

Unscaled Zero Point Vibrational Energy (zpe) 17487.0 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 16777.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.14179 0.04601 0.04595

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 0.000 1.123
S2 0.000 1.663 -0.156
S3 0.000 -1.663 -0.156
C4 1.751 1.686 -0.681
C5 -1.751 -1.686 -0.681
H6 1.861 2.535 -1.380
H7 -1.861 -2.535 -1.380
H8 2.416 1.830 0.186
H9 1.999 0.748 -1.205
H10 -2.416 -1.830 0.186
H11 -1.999 -0.748 -1.205

Atom - Atom Distances (Å)
  S1 S2 S3 C4 C5 H6 H7 H8 H9 H10 H11
S12.09852.09853.02763.02764.01894.01893.17293.15823.17293.1582
S22.09853.32651.82843.81602.39144.75202.44622.43574.26143.3028
S32.09853.32653.81601.82844.75202.39144.26143.30282.44622.4357
C43.02761.82843.81604.86241.10435.59921.10191.10275.52144.5015
C53.02763.81601.82844.86245.59921.10435.52144.50151.10191.1027
H64.01892.39144.75201.10435.59926.28871.80411.80066.30835.0698
H74.01894.75202.39145.59921.10436.28876.30835.06981.80411.8006
H83.17292.44624.26141.10195.52141.80416.30831.81066.06245.2985
H93.15822.43573.30281.10274.50151.80065.06981.81065.29854.2685
H103.17294.26142.44625.52141.10196.30831.80416.06245.29851.8106
H113.15823.30282.43574.50151.10275.06981.80065.29854.26851.8106

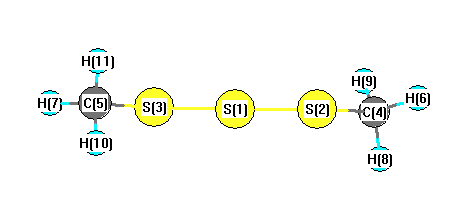
picture of dimethyl trisulfide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 S2 C4 100.662 S1 S3 C5 100.662
S2 S1 S3 104.861 S2 C4 H6 106.635
S2 C4 H8 110.732 S2 C4 H9 109.916
S3 C5 H7 106.635 S3 C5 H10 110.732
S3 C5 H11 109.916 H6 C4 H8 109.718
H6 C4 H9 109.341 H7 C5 H10 109.718
H7 C5 H11 109.341 H8 C4 H9 110.417
H10 C5 H11 110.417
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability