return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4F2 (orthodifluorobenzene)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-429.591746
Energy at 298.15K-429.596561
Nuclear repulsion energy345.612524
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3255 3114 1.70      
2 A1 3239 3099 7.81      
3 A1 1699 1626 31.14      
4 A1 1586 1518 184.16      
5 A1 1331 1273 70.16      
6 A1 1323 1265 27.32      
7 A1 1168 1118 0.14      
8 A1 1057 1011 5.60      
9 A1 786 752 33.56      
10 A1 581 556 3.69      
11 A1 285 273 0.37      
12 A2 969 927 0.00      
13 A2 872 834 0.00      
14 A2 687 658 0.00      
15 A2 556 532 0.00      
16 A2 192 183 0.00      
17 B1 953 912 7.06      
18 B1 779 746 73.18      
19 B1 471 450 1.79      
20 B1 303 289 0.05      
21 B2 3250 3109 3.44      
22 B2 3225 3086 0.89      
23 B2 1707 1633 8.22      
24 B2 1512 1446 11.97      
25 B2 1302 1246 12.41      
26 B2 1248 1194 31.70      
27 B2 1127 1078 16.91      
28 B2 862 825 15.40      
29 B2 553 529 3.78      
30 B2 441 422 0.33      

Unscaled Zero Point Vibrational Energy (zpe) 18658.6 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 17850.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.10777 0.07340 0.04366

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.698 -0.542
C2 0.000 -0.698 -0.542
C3 0.000 -1.404 0.661
C4 0.000 -0.702 1.877
C5 0.000 0.702 1.877
C6 0.000 1.404 0.661
F7 0.000 1.354 -1.714
F8 0.000 -1.354 -1.714
H9 0.000 -2.497 0.627
H10 0.000 -1.255 2.821
H11 0.000 1.255 2.821
H12 0.000 2.497 0.627

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 F7 F8 H9 H10 H11 H12
C11.39562.42172.79562.41971.39531.34262.36283.40193.88983.40982.1456
C21.39561.39532.41972.79562.42172.36281.34262.14563.40983.88983.4019
C32.42171.39531.40392.43202.80743.63952.37521.09372.16553.42613.9007
C42.79562.41971.40391.40482.43204.13813.64952.18711.09422.17363.4349
C52.41972.79562.43201.40481.40393.64954.13813.43492.17361.09422.1871
C61.39532.42172.80742.43201.40392.37523.63953.90073.42612.16551.0937
F71.34262.36283.63954.13813.64952.37522.70864.50655.23234.53622.6044
F82.36281.34262.37523.64954.13813.63952.70862.60444.53625.23234.5065
H93.40192.14561.09372.18713.43493.90074.50652.60442.52154.34694.9937
H103.88983.40982.16551.09422.17363.42615.23234.53622.52152.51074.3469
H113.40983.88983.42612.17361.09422.16554.53625.23234.34692.51072.5215
H122.14563.40193.90073.43492.18711.09372.60444.50654.99374.34692.5215

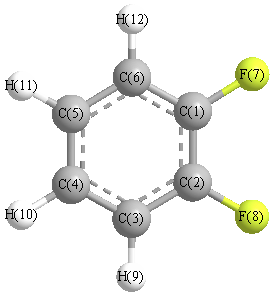
picture of orthodifluorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.394 C1 C2 F8 119.272
C1 C6 C5 119.637 C1 C6 H12 118.596
C2 C1 C6 120.394 C2 C1 F7 119.272
C2 C3 C4 119.637 C2 C3 H9 118.596
C3 C2 F8 120.334 C3 C4 C5 119.970
C3 C4 H10 119.679 C4 C3 H9 121.767
C4 C5 C6 119.970 C4 C5 H11 120.351
C5 C4 H10 120.351 C5 C6 H12 121.767
C6 C1 F7 120.334 C6 C5 H11 119.679
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability