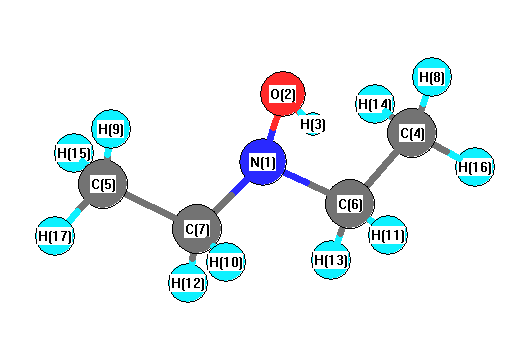
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3313 |
3318 |
15.66 |
|
|
|
| 2 |
A |
3052 |
3057 |
38.07 |
|
|
|
| 3 |
A |
3034 |
3039 |
40.60 |
|
|
|
| 4 |
A |
2980 |
2985 |
42.39 |
|
|
|
| 5 |
A |
2965 |
2970 |
5.76 |
|
|
|
| 6 |
A |
2799 |
2804 |
166.43 |
|
|
|
| 7 |
A |
1447 |
1450 |
6.43 |
|
|
|
| 8 |
A |
1418 |
1421 |
0.78 |
|
|
|
| 9 |
A |
1414 |
1416 |
19.71 |
|
|
|
| 10 |
A |
1373 |
1375 |
24.67 |
|
|
|
| 11 |
A |
1358 |
1360 |
23.73 |
|
|
|
| 12 |
A |
1338 |
1341 |
0.02 |
|
|
|
| 13 |
A |
1253 |
1255 |
2.72 |
|
|
|
| 14 |
A |
1128 |
1130 |
0.96 |
|
|
|
| 15 |
A |
1114 |
1115 |
10.06 |
|
|
|
| 16 |
A |
1026 |
1027 |
4.06 |
|
|
|
| 17 |
A |
883 |
884 |
9.30 |
|
|
|
| 18 |
A |
806 |
808 |
1.36 |
|
|
|
| 19 |
A |
719 |
720 |
22.88 |
|
|
|
| 20 |
A |
449 |
450 |
3.71 |
|
|
|
| 21 |
A |
411 |
412 |
0.11 |
|
|
|
| 22 |
A |
209 |
210 |
0.24 |
|
|
|
| 23 |
A |
206 |
206 |
1.74 |
|
|
|
| 24 |
A |
110 |
110 |
0.24 |
|
|
|
| 25 |
A |
3051 |
3056 |
0.53 |
|
|
|
| 26 |
A |
3034 |
3039 |
13.89 |
|
|
|
| 27 |
A |
2969 |
2974 |
5.23 |
|
|
|
| 28 |
A |
2964 |
2968 |
44.69 |
|
|
|
| 29 |
A |
2783 |
2788 |
8.87 |
|
|
|
| 30 |
A |
1445 |
1447 |
0.38 |
|
|
|
| 31 |
A |
1414 |
1416 |
1.01 |
|
|
|
| 32 |
A |
1406 |
1408 |
0.87 |
|
|
|
| 33 |
A |
1348 |
1350 |
12.72 |
|
|
|
| 34 |
A |
1317 |
1319 |
45.11 |
|
|
|
| 35 |
A |
1248 |
1250 |
2.12 |
|
|
|
| 36 |
A |
1150 |
1152 |
0.66 |
|
|
|
| 37 |
A |
1051 |
1053 |
0.32 |
|
|
|
| 38 |
A |
1024 |
1025 |
24.75 |
|
|
|
| 39 |
A |
922 |
924 |
0.00 |
|
|
|
| 40 |
A |
786 |
787 |
0.06 |
|
|
|
| 41 |
A |
528 |
529 |
29.87 |
|
|
|
| 42 |
A |
408 |
409 |
76.62 |
|
|
|
| 43 |
A |
336 |
336 |
2.66 |
|
|
|
| 44 |
A |
241 |
242 |
0.08 |
|
|
|
| 45 |
A |
140 |
141 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32185.4 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 32236.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.133 |
|
|
|
| 2 |
O |
-0.200 |
|
|
|
| 3 |
H |
0.128 |
|
|
|
| 4 |
C |
0.037 |
|
|
|
| 5 |
C |
0.037 |
|
|
|
| 6 |
C |
0.077 |
|
|
|
| 7 |
C |
0.077 |
|
|
|
| 8 |
H |
0.014 |
|
|
|
| 9 |
H |
0.014 |
|
|
|
| 10 |
H |
-0.039 |
|
|
|
| 11 |
H |
-0.039 |
|
|
|
| 12 |
H |
-0.001 |
|
|
|
| 13 |
H |
-0.001 |
|
|
|
| 14 |
H |
0.014 |
|
|
|
| 15 |
H |
0.014 |
|
|
|
| 16 |
H |
0.000 |
|
|
|
| 17 |
H |
0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.352 |
-1.447 |
0.000 |
1.980 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.232 |
1.997 |
0.000 |
| y |
1.997 |
-38.715 |
0.000 |
| z |
0.000 |
0.000 |
-38.390 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.679 |
1.997 |
0.000 |
| y |
1.997 |
0.597 |
0.000 |
| z |
0.000 |
0.000 |
1.083 |
|
| Polar |
|---|
| 3z2-r2 | 2.166 |
|---|
| x2-y2 | -1.517 |
|---|
| xy | 1.997 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.708 |
0.160 |
0.000 |
| y |
0.160 |
8.392 |
0.000 |
| z |
0.000 |
0.000 |
11.010 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |