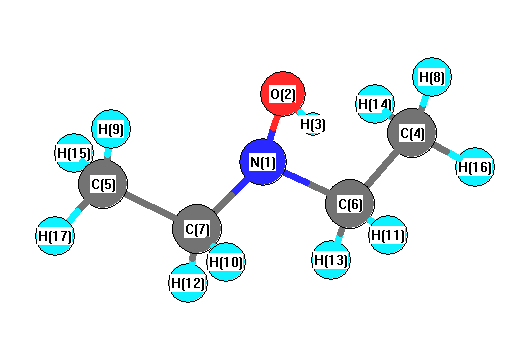
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3540 |
3416 |
6.56 |
|
|
|
| 2 |
A |
3157 |
3046 |
29.74 |
|
|
|
| 3 |
A |
3142 |
3032 |
29.97 |
|
|
|
| 4 |
A |
3086 |
2978 |
36.48 |
|
|
|
| 5 |
A |
3056 |
2949 |
4.67 |
|
|
|
| 6 |
A |
2900 |
2798 |
162.25 |
|
|
|
| 7 |
A |
1485 |
1433 |
11.36 |
|
|
|
| 8 |
A |
1460 |
1409 |
3.45 |
|
|
|
| 9 |
A |
1453 |
1402 |
23.84 |
|
|
|
| 10 |
A |
1421 |
1371 |
33.81 |
|
|
|
| 11 |
A |
1406 |
1357 |
19.62 |
|
|
|
| 12 |
A |
1368 |
1321 |
0.39 |
|
|
|
| 13 |
A |
1292 |
1246 |
3.29 |
|
|
|
| 14 |
A |
1183 |
1141 |
3.05 |
|
|
|
| 15 |
A |
1156 |
1116 |
13.38 |
|
|
|
| 16 |
A |
1096 |
1058 |
2.55 |
|
|
|
| 17 |
A |
963 |
929 |
17.42 |
|
|
|
| 18 |
A |
825 |
796 |
2.61 |
|
|
|
| 19 |
A |
789 |
761 |
18.28 |
|
|
|
| 20 |
A |
453 |
437 |
4.32 |
|
|
|
| 21 |
A |
426 |
411 |
0.27 |
|
|
|
| 22 |
A |
213 |
206 |
0.14 |
|
|
|
| 23 |
A |
209 |
202 |
2.22 |
|
|
|
| 24 |
A |
111 |
107 |
0.31 |
|
|
|
| 25 |
A |
3156 |
3045 |
0.44 |
|
|
|
| 26 |
A |
3142 |
3032 |
11.88 |
|
|
|
| 27 |
A |
3076 |
2969 |
0.17 |
|
|
|
| 28 |
A |
3056 |
2949 |
43.38 |
|
|
|
| 29 |
A |
2887 |
2786 |
6.87 |
|
|
|
| 30 |
A |
1482 |
1430 |
0.19 |
|
|
|
| 31 |
A |
1452 |
1402 |
1.28 |
|
|
|
| 32 |
A |
1447 |
1396 |
1.05 |
|
|
|
| 33 |
A |
1389 |
1340 |
39.69 |
|
|
|
| 34 |
A |
1364 |
1316 |
15.90 |
|
|
|
| 35 |
A |
1287 |
1242 |
2.59 |
|
|
|
| 36 |
A |
1204 |
1161 |
1.88 |
|
|
|
| 37 |
A |
1094 |
1055 |
3.28 |
|
|
|
| 38 |
A |
1072 |
1034 |
19.48 |
|
|
|
| 39 |
A |
964 |
931 |
0.42 |
|
|
|
| 40 |
A |
804 |
776 |
0.01 |
|
|
|
| 41 |
A |
558 |
538 |
33.69 |
|
|
|
| 42 |
A |
421 |
406 |
87.04 |
|
|
|
| 43 |
A |
345 |
333 |
1.99 |
|
|
|
| 44 |
A |
247 |
238 |
0.06 |
|
|
|
| 45 |
A |
140 |
135 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33387.8 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 32219.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.178 |
|
|
|
| 2 |
O |
-0.216 |
|
|
|
| 3 |
H |
0.144 |
|
|
|
| 4 |
C |
-0.058 |
|
|
|
| 5 |
C |
-0.058 |
|
|
|
| 6 |
C |
0.004 |
|
|
|
| 7 |
C |
0.004 |
|
|
|
| 8 |
H |
0.052 |
|
|
|
| 9 |
H |
0.052 |
|
|
|
| 10 |
H |
-0.002 |
|
|
|
| 11 |
H |
-0.002 |
|
|
|
| 12 |
H |
0.040 |
|
|
|
| 13 |
H |
0.040 |
|
|
|
| 14 |
H |
0.051 |
|
|
|
| 15 |
H |
0.051 |
|
|
|
| 16 |
H |
0.038 |
|
|
|
| 17 |
H |
0.038 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.326 |
-1.617 |
0.000 |
2.091 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.437 |
2.167 |
0.000 |
| y |
2.167 |
-38.096 |
0.000 |
| z |
0.000 |
0.000 |
-37.633 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.572 |
2.167 |
0.000 |
| y |
2.167 |
0.439 |
0.000 |
| z |
0.000 |
0.000 |
1.133 |
|
| Polar |
|---|
| 3z2-r2 | 2.266 |
|---|
| x2-y2 | -1.341 |
|---|
| xy | 2.167 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
210.630 |
| (<r2>)1/2 |
14.513 |