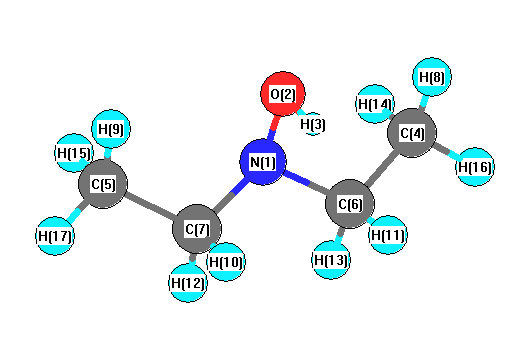
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3335 |
3315 |
17.18 |
|
|
|
| 2 |
A |
3082 |
3064 |
25.08 |
|
|
|
| 3 |
A |
3070 |
3053 |
28.88 |
|
|
|
| 4 |
A |
3003 |
2985 |
37.10 |
|
|
|
| 5 |
A |
2986 |
2968 |
5.63 |
|
|
|
| 6 |
A |
2807 |
2791 |
161.54 |
|
|
|
| 7 |
A |
1438 |
1429 |
9.06 |
|
|
|
| 8 |
A |
1408 |
1400 |
0.69 |
|
|
|
| 9 |
A |
1404 |
1396 |
29.48 |
|
|
|
| 10 |
A |
1374 |
1366 |
40.54 |
|
|
|
| 11 |
A |
1359 |
1351 |
7.81 |
|
|
|
| 12 |
A |
1322 |
1314 |
0.48 |
|
|
|
| 13 |
A |
1245 |
1238 |
2.47 |
|
|
|
| 14 |
A |
1129 |
1123 |
1.00 |
|
|
|
| 15 |
A |
1117 |
1111 |
13.06 |
|
|
|
| 16 |
A |
1059 |
1052 |
4.50 |
|
|
|
| 17 |
A |
908 |
902 |
13.36 |
|
|
|
| 18 |
A |
801 |
797 |
2.22 |
|
|
|
| 19 |
A |
750 |
745 |
20.36 |
|
|
|
| 20 |
A |
444 |
441 |
4.11 |
|
|
|
| 21 |
A |
416 |
413 |
0.31 |
|
|
|
| 22 |
A |
214 |
213 |
0.04 |
|
|
|
| 23 |
A |
206 |
205 |
1.97 |
|
|
|
| 24 |
A |
115 |
114 |
0.25 |
|
|
|
| 25 |
A |
3081 |
3064 |
1.21 |
|
|
|
| 26 |
A |
3070 |
3053 |
10.07 |
|
|
|
| 27 |
A |
2992 |
2975 |
0.15 |
|
|
|
| 28 |
A |
2985 |
2967 |
41.80 |
|
|
|
| 29 |
A |
2793 |
2777 |
8.30 |
|
|
|
| 30 |
A |
1435 |
1427 |
0.57 |
|
|
|
| 31 |
A |
1404 |
1396 |
0.81 |
|
|
|
| 32 |
A |
1393 |
1385 |
0.47 |
|
|
|
| 33 |
A |
1337 |
1329 |
31.51 |
|
|
|
| 34 |
A |
1312 |
1304 |
26.24 |
|
|
|
| 35 |
A |
1243 |
1236 |
2.63 |
|
|
|
| 36 |
A |
1160 |
1154 |
1.09 |
|
|
|
| 37 |
A |
1058 |
1052 |
0.67 |
|
|
|
| 38 |
A |
1031 |
1025 |
22.59 |
|
|
|
| 39 |
A |
938 |
933 |
0.33 |
|
|
|
| 40 |
A |
781 |
777 |
0.19 |
|
|
|
| 41 |
A |
543 |
540 |
34.21 |
|
|
|
| 42 |
A |
410 |
407 |
76.21 |
|
|
|
| 43 |
A |
337 |
335 |
1.38 |
|
|
|
| 44 |
A |
245 |
243 |
0.07 |
|
|
|
| 45 |
A |
146 |
145 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32341.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 32153.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.149 |
|
|
|
| 2 |
O |
-0.193 |
|
|
|
| 3 |
H |
0.134 |
|
|
|
| 4 |
C |
-0.033 |
|
|
|
| 5 |
C |
-0.033 |
|
|
|
| 6 |
C |
0.016 |
|
|
|
| 7 |
C |
0.016 |
|
|
|
| 8 |
H |
0.039 |
|
|
|
| 9 |
H |
0.039 |
|
|
|
| 10 |
H |
-0.014 |
|
|
|
| 11 |
H |
-0.014 |
|
|
|
| 12 |
H |
0.027 |
|
|
|
| 13 |
H |
0.027 |
|
|
|
| 14 |
H |
0.040 |
|
|
|
| 15 |
H |
0.040 |
|
|
|
| 16 |
H |
0.028 |
|
|
|
| 17 |
H |
0.028 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.195 |
1.940 |
0.000 |
1.950 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.146 |
0.462 |
0.000 |
| y |
0.462 |
-41.312 |
0.000 |
| z |
0.000 |
0.000 |
-38.079 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.550 |
0.462 |
0.000 |
| y |
0.462 |
-3.699 |
0.000 |
| z |
0.000 |
0.000 |
1.150 |
|
| Polar |
|---|
| 3z2-r2 | 2.299 |
|---|
| x2-y2 | 4.166 |
|---|
| xy | 0.462 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.617 |
-0.139 |
0.000 |
| y |
-0.139 |
8.378 |
0.000 |
| z |
0.000 |
0.000 |
10.901 |
<r2> (average value of r
2) Å
2
| <r2> |
212.576 |
| (<r2>)1/2 |
14.580 |