return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2FCH2OH (2-fluoroethanol)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-253.634574
Energy at 298.15K 
HF Energy-252.946869
Nuclear repulsion energy130.938486
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3868 3700 35.83      
2 A 3154 3017 44.89      
3 A 3134 2998 34.73      
4 A 3088 2954 27.72      
5 A 3042 2910 48.24      
6 A 1510 1445 2.65      
7 A 1505 1440 3.27      
8 A 1472 1408 47.29      
9 A 1434 1372 23.99      
10 A 1407 1346 1.74      
11 A 1285 1230 8.02      
12 A 1246 1192 10.61      
13 A 1163 1113 56.07      
14 A 1139 1089 32.33      
15 A 1098 1051 55.86      
16 A 920 880 15.44      
17 A 889 850 31.73      
18 A 527 504 8.63      
19 A 434 415 122.76      
20 A 321 307 13.12      
21 A 166 159 10.46      

Unscaled Zero Point Vibrational Energy (zpe) 16400.4 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 15690.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.51943 0.18555 0.15342

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.692 0.582 0.278
C2 -0.721 0.565 -0.276
O3 1.436 -0.525 -0.189
F4 -1.341 -0.606 0.159
H5 1.210 1.496 -0.063
H6 0.644 0.610 1.386
H7 -1.312 1.428 0.079
H8 -0.706 0.549 -1.380
H9 0.918 -1.304 0.052

Atom - Atom Distances (Å)
  C1 C2 O3 F4 H5 H6 H7 H8 H9
C11.51781.41322.35711.10411.11012.18412.16881.9130
C21.51782.41851.39412.15442.15221.10491.10432.5077
O31.41322.41852.79952.03702.09683.38212.67620.9661
F42.35711.39412.79953.31222.63182.03612.02632.3665
H51.10412.15442.03703.31221.79012.52672.51042.8171
H61.11012.15222.09682.63181.79012.49123.07942.3493
H72.18411.10493.38212.03612.52672.49121.80823.5271
H82.16881.10432.67622.02632.51043.07941.80822.8504
H91.91302.50770.96612.36652.81712.34933.52712.8504

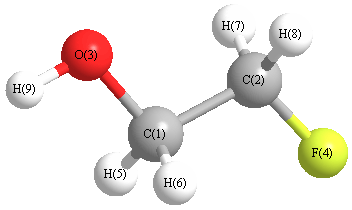
picture of 2-fluoroethanol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 F4 108.012 C1 C2 H7 111.800
C1 C2 H8 110.620 C1 O3 H9 105.488
C2 C1 O3 111.158 C2 C1 H5 109.497
C2 C1 H6 108.973 O3 C1 H5 107.401
O3 C1 H6 111.833
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability