return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (benzocyclobutadiene)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-307.438906
Energy at 298.15K-307.444429
HF Energy-306.356105
Nuclear repulsion energy309.589140
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3269 3136 9.32      
2 A1 3229 3097 12.17      
3 A1 3212 3082 13.11      
4 A1 1733 1662 1.91      
5 A1 1568 1504 1.63      
6 A1 1526 1464 2.70      
7 A1 1439 1381 9.43      
8 A1 1184 1136 0.08      
9 A1 1101 1057 0.19      
10 A1 1059 1016 0.24      
11 A1 967 928 3.35      
12 A1 820 787 0.38      
13 A1 543 521 0.18      
14 A2 948 910 0.00      
15 A2 888 851 0.00      
16 A2 858 823 0.00      
17 A2 665 638 0.00      
18 A2 555 533 0.00      
19 A2 290 279 0.00      
20 B1 899 862 6.20      
21 B1 750 720 84.10      
22 B1 716 687 7.30      
23 B1 345 331 1.82      
24 B1 234 225 11.38      
25 B2 3239 3108 1.13      
26 B2 3222 3091 27.52      
27 B2 3197 3067 0.21      
28 B2 1698 1629 0.33      
29 B2 1464 1404 3.81      
30 B2 1303 1251 4.26      
31 B2 1255 1204 10.40      
32 B2 1092 1048 1.54      
33 B2 1044 1002 0.53      
34 B2 877 841 0.62      
35 B2 651 625 0.79      
36 B2 418 401 3.47      

Unscaled Zero Point Vibrational Energy (zpe) 24128.8 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 23149.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.15785 0.07212 0.04950

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.457 -0.624
C2 0.000 0.691 -1.863
C3 0.000 -0.691 -1.863
C4 0.000 -1.457 -0.624
C5 0.000 -0.684 2.062
C6 0.000 0.684 2.062
C7 0.000 0.726 0.532
C8 0.000 -0.726 0.532
H9 0.000 2.553 -0.653
H10 0.000 1.233 -2.816
H11 0.000 -1.233 -2.816
H12 0.000 -2.553 -0.653
H13 0.000 -1.461 2.833
H14 0.000 1.461 2.833

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.45742.48072.91493.43492.79471.36762.47001.09632.20413.47094.01104.52403.4563
C21.45741.38242.48074.15923.92532.39532.78282.22131.09622.14743.46325.16594.7588
C32.48071.38241.45743.92534.15922.78282.39533.46322.14741.09622.22134.75885.1659
C42.91492.48071.45742.79473.43492.47001.36764.01103.47092.20411.09633.45634.5240
C53.43494.15923.92532.79471.36812.08061.53084.22485.24144.90903.29581.09472.2798
C62.79473.92534.15923.43491.36811.53082.08063.29584.90905.24144.22482.27981.0947
C71.36762.39532.78282.47002.08061.53081.45142.17783.38623.87893.48643.17462.4158
C82.47002.78282.39531.36761.53082.08061.45143.48643.87893.38622.17782.41583.1746
H91.09632.22133.46324.01104.22483.29582.17783.48642.53484.36125.10685.31653.6522
H102.20411.09622.14743.47095.24144.90903.38623.87892.53482.46634.36126.25875.6536
H113.47092.14741.09622.20414.90905.24143.87893.38624.36122.46632.53485.65366.2587
H124.01103.46322.22131.09633.29584.22483.48642.17785.10684.36122.53483.65225.3165
H134.52405.16594.75883.45631.09472.27983.17462.41585.31656.25875.65363.65222.9229
H143.45634.75885.16594.52402.27981.09472.41583.17463.65225.65366.25875.31652.9229

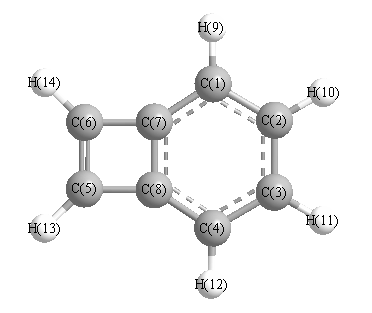
picture of benzocyclobutadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 121.719 C1 C2 H10 118.654
C1 C7 C6 149.209 C1 C7 N8 122.350
C2 C1 C7 115.931 C2 C1 H9 120.211
C2 C3 C4 121.719 C2 C3 H11 119.627
C3 C2 H10 119.627 C3 C4 N8 115.931
C3 C4 H12 120.211 C4 C3 H11 118.654
C4 N8 C5 149.209 C4 N8 C7 122.350
C5 C6 C7 91.559 C5 C6 H14 135.244
C5 N8 C7 88.441 C6 C5 N8 91.559
C6 C5 H13 135.244 C6 C7 N8 88.441
C7 C1 H9 123.858 C7 C6 H14 133.197
N8 C4 H12 123.858 N8 C5 H13 133.197
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability