Jump to
S1C2
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
Geometric Data calculated at PBEPBE/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -235.513253 |
| Energy at 298.15K | -235.525299 |
| HF Energy | -235.513253 |
| Nuclear repulsion energy | 224.811828 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3060 |
3043 |
45.70 |
|
|
|
| 2 |
A |
3051 |
3034 |
31.44 |
|
|
|
| 3 |
A |
3044 |
3026 |
15.14 |
|
|
|
| 4 |
A |
3041 |
3023 |
49.34 |
|
|
|
| 5 |
A |
3040 |
3022 |
4.90 |
|
|
|
| 6 |
A |
3010 |
2993 |
17.65 |
|
|
|
| 7 |
A |
3002 |
2985 |
22.88 |
|
|
|
| 8 |
A |
2980 |
2963 |
7.28 |
|
|
|
| 9 |
A |
2964 |
2947 |
34.09 |
|
|
|
| 10 |
A |
2957 |
2940 |
30.70 |
|
|
|
| 11 |
A |
2953 |
2936 |
38.19 |
|
|
|
| 12 |
A |
2928 |
2911 |
27.51 |
|
|
|
| 13 |
A |
1702 |
1692 |
0.60 |
|
|
|
| 14 |
A |
1432 |
1423 |
4.35 |
|
|
|
| 15 |
A |
1420 |
1412 |
5.39 |
|
|
|
| 16 |
A |
1416 |
1408 |
0.53 |
|
|
|
| 17 |
A |
1412 |
1404 |
10.12 |
|
|
|
| 18 |
A |
1399 |
1391 |
1.60 |
|
|
|
| 19 |
A |
1398 |
1390 |
5.75 |
|
|
|
| 20 |
A |
1347 |
1339 |
0.69 |
|
|
|
| 21 |
A |
1344 |
1337 |
1.24 |
|
|
|
| 22 |
A |
1328 |
1320 |
2.05 |
|
|
|
| 23 |
A |
1282 |
1275 |
0.73 |
|
|
|
| 24 |
A |
1278 |
1271 |
0.43 |
|
|
|
| 25 |
A |
1263 |
1255 |
0.05 |
|
|
|
| 26 |
A |
1229 |
1222 |
8.94 |
|
|
|
| 27 |
A |
1198 |
1191 |
0.99 |
|
|
|
| 28 |
A |
1138 |
1132 |
0.05 |
|
|
|
| 29 |
A |
1081 |
1075 |
6.12 |
|
|
|
| 30 |
A |
1076 |
1069 |
0.20 |
|
|
|
| 31 |
A |
1039 |
1033 |
0.58 |
|
|
|
| 32 |
A |
1012 |
1006 |
0.57 |
|
|
|
| 33 |
A |
1006 |
1000 |
7.20 |
|
|
|
| 34 |
A |
965 |
959 |
23.77 |
|
|
|
| 35 |
A |
900 |
894 |
8.52 |
|
|
|
| 36 |
A |
878 |
873 |
1.66 |
|
|
|
| 37 |
A |
843 |
838 |
0.52 |
|
|
|
| 38 |
A |
752 |
747 |
0.73 |
|
|
|
| 39 |
A |
729 |
724 |
2.87 |
|
|
|
| 40 |
A |
517 |
514 |
0.16 |
|
|
|
| 41 |
A |
379 |
377 |
1.12 |
|
|
|
| 42 |
A |
302 |
300 |
0.85 |
|
|
|
| 43 |
A |
278 |
277 |
1.45 |
|
|
|
| 44 |
A |
247 |
245 |
0.06 |
|
|
|
| 45 |
A |
210 |
209 |
0.96 |
|
|
|
| 46 |
A |
140 |
139 |
0.27 |
|
|
|
| 47 |
A |
98 |
98 |
0.06 |
|
|
|
| 48 |
A |
76 |
76 |
0.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35070.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 34867.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
3.201 |
0.210 |
-0.162 |
| C2 |
1.818 |
-0.372 |
-0.210 |
| C3 |
0.741 |
0.119 |
0.433 |
| C4 |
-0.646 |
-0.458 |
0.381 |
| C5 |
-1.696 |
0.518 |
-0.182 |
| C6 |
-3.112 |
-0.064 |
-0.189 |
| H7 |
3.241 |
1.120 |
0.469 |
| H8 |
3.934 |
-0.521 |
0.244 |
| H9 |
3.564 |
0.479 |
-1.177 |
| H10 |
1.690 |
-1.283 |
-0.825 |
| H11 |
0.866 |
1.033 |
1.046 |
| H12 |
-0.962 |
-0.755 |
1.409 |
| H13 |
-0.641 |
-1.392 |
-0.223 |
| H14 |
-1.399 |
0.810 |
-1.213 |
| H15 |
-1.679 |
1.456 |
0.416 |
| H16 |
-3.848 |
0.658 |
-0.597 |
| H17 |
-3.167 |
-0.984 |
-0.808 |
| H18 |
-3.441 |
-0.336 |
0.836 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.5005 | 2.5317 | 3.9413 | 4.9062 | 6.3182 | 1.1085 | 1.1120 | 1.1119 | 2.2249 | 2.7546 | 4.5525 | 4.1627 | 4.7561 | 5.0691 | 7.0763 | 6.5106 | 6.7386 |
C2 | 1.5005 | | 1.3472 | 2.5354 | 3.6253 | 4.9397 | 2.1710 | 2.1693 | 2.1694 | 1.1067 | 2.1120 | 3.2403 | 2.6626 | 3.5711 | 3.9955 | 5.7722 | 5.0581 | 5.3629 | C3 | 2.5317 | 1.3472 | | 1.5033 | 2.5451 | 3.9072 | 2.6930 | 3.2620 | 3.2693 | 2.1095 | 1.1075 | 2.1493 | 2.1509 | 2.7869 | 2.7648 | 4.7343 | 4.2464 | 4.2267 | C4 | 3.9413 | 2.5354 | 1.5033 | | 1.5405 | 2.5617 | 4.1960 | 4.5824 | 4.5855 | 2.7552 | 2.2250 | 1.1156 | 1.1123 | 2.1721 | 2.1757 | 3.5296 | 2.8367 | 2.8352 | C5 | 4.9062 | 3.6253 | 2.5451 | 1.5405 | | 1.5305 | 5.0161 | 5.7411 | 5.3532 | 3.8888 | 2.8875 | 2.1658 | 2.1821 | 1.1120 | 1.1130 | 2.1962 | 2.1935 | 2.1936 | C6 | 6.3182 | 4.9397 | 3.9072 | 2.5617 | 1.5305 | | 6.4956 | 7.0741 | 6.7700 | 4.9949 | 4.3069 | 2.7662 | 2.8052 | 2.1786 | 2.1747 | 1.1087 | 1.1104 | 1.1105 | H7 | 1.1085 | 2.1710 | 2.6930 | 4.1960 | 5.0161 | 6.4956 | | 1.7958 | 1.7961 | 3.1397 | 2.4460 | 4.6977 | 4.6758 | 4.9451 | 4.9316 | 7.1838 | 6.8646 | 6.8492 | H8 | 1.1120 | 2.1693 | 3.2620 | 4.5824 | 5.7411 | 7.0741 | 1.7958 | | 1.7769 | 2.6001 | 3.5318 | 5.0385 | 4.6811 | 5.6865 | 5.9535 | 7.9159 | 7.1936 | 7.4018 | H9 | 1.1119 | 2.1694 | 3.2693 | 4.5855 | 5.3532 | 6.7700 | 1.7961 | 1.7769 | | 2.5963 | 3.5399 | 5.3569 | 4.7003 | 4.9738 | 5.5658 | 7.4367 | 6.8979 | 7.3342 | H10 | 2.2249 | 1.1067 | 2.1095 | 2.7552 | 3.8888 | 4.9949 | 3.1397 | 2.6001 | 2.5963 | | 3.0897 | 3.5076 | 2.4101 | 3.7517 | 4.5160 | 5.8729 | 4.8663 | 5.4763 | H11 | 2.7546 | 2.1120 | 1.1075 | 2.2250 | 2.8875 | 4.3069 | 2.4460 | 3.5318 | 3.5399 | 3.0897 | | 2.5828 | 3.1247 | 3.2067 | 2.6554 | 5.0063 | 4.8754 | 4.5244 | H12 | 4.5525 | 3.2403 | 2.1493 | 1.1156 | 2.1658 | 2.7662 | 4.6977 | 5.0385 | 5.3569 | 3.5076 | 2.5828 | | 1.7811 | 3.0848 | 2.5277 | 3.7880 | 3.1346 | 2.5790 | H13 | 4.1627 | 2.6626 | 2.1509 | 1.1123 | 2.1821 | 2.8052 | 4.6758 | 4.6811 | 4.7003 | 2.4101 | 3.1247 | 1.7811 | | 2.5306 | 3.0980 | 3.8243 | 2.6243 | 3.1748 | H14 | 4.7561 | 3.5711 | 2.7869 | 2.1721 | 1.1120 | 2.1786 | 4.9451 | 5.6865 | 4.9738 | 3.7517 | 3.2067 | 3.0848 | 2.5306 | | 1.7748 | 2.5302 | 2.5518 | 3.1119 | H15 | 5.0691 | 3.9955 | 2.7648 | 2.1757 | 1.1130 | 2.1747 | 4.9316 | 5.9535 | 5.5658 | 4.5160 | 2.6554 | 2.5277 | 3.0980 | 1.7748 | | 2.5240 | 3.1093 | 2.5483 | H16 | 7.0763 | 5.7722 | 4.7343 | 3.5296 | 2.1962 | 1.1087 | 7.1838 | 7.9159 | 7.4367 | 5.8729 | 5.0063 | 3.7880 | 3.8243 | 2.5302 | 2.5240 | | 1.7900 | 1.7904 | H17 | 6.5106 | 5.0581 | 4.2464 | 2.8367 | 2.1935 | 1.1104 | 6.8646 | 7.1936 | 6.8979 | 4.8663 | 4.8754 | 3.1346 | 2.6243 | 2.5518 | 3.1093 | 1.7900 | | 1.7879 | H18 | 6.7386 | 5.3629 | 4.2267 | 2.8352 | 2.1936 | 1.1105 | 6.8492 | 7.4018 | 7.3342 | 5.4763 | 4.5244 | 2.5790 | 3.1748 | 3.1119 | 2.5483 | 1.7904 | 1.7879 | |
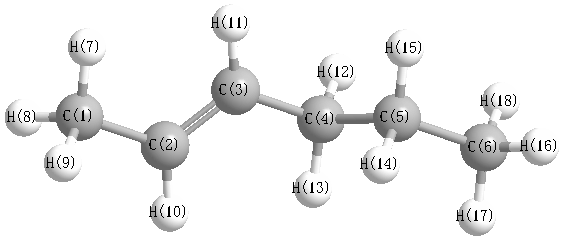 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.417 |
|
C1 |
C2 |
H10 |
116.356 |
| C2 |
C1 |
H7 |
111.756 |
|
C2 |
C1 |
H8 |
111.411 |
| C2 |
C1 |
H9 |
111.421 |
|
C2 |
C3 |
C4 |
125.515 |
| C2 |
C3 |
H11 |
118.393 |
|
C3 |
C2 |
H10 |
118.226 |
| C3 |
C4 |
C5 |
113.462 |
|
C3 |
C4 |
H12 |
109.414 |
| C3 |
C4 |
H13 |
109.729 |
|
C4 |
C3 |
H11 |
116.087 |
| C4 |
C5 |
C6 |
113.053 |
|
C4 |
C5 |
H14 |
108.872 |
| C4 |
C5 |
H15 |
109.088 |
|
C5 |
C4 |
H12 |
108.185 |
| C5 |
C4 |
H13 |
109.625 |
|
C5 |
C6 |
H16 |
111.646 |
| C5 |
C6 |
H17 |
111.319 |
|
C5 |
C6 |
H18 |
111.321 |
| C6 |
C5 |
H14 |
110.057 |
|
C6 |
C5 |
H15 |
109.695 |
| H7 |
C1 |
H8 |
107.947 |
|
H7 |
C1 |
H9 |
107.979 |
| H8 |
C1 |
H9 |
106.070 |
|
H12 |
C4 |
H13 |
106.155 |
| H14 |
C5 |
H15 |
105.814 |
|
H16 |
C6 |
H17 |
107.541 |
| H16 |
C6 |
H18 |
107.568 |
|
H17 |
C6 |
H18 |
107.225 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.015 |
|
|
|
| 2 |
C |
-0.084 |
|
|
|
| 3 |
C |
-0.030 |
|
|
|
| 4 |
C |
0.014 |
|
|
|
| 5 |
C |
-0.076 |
|
|
|
| 6 |
C |
-0.051 |
|
|
|
| 7 |
H |
0.027 |
|
|
|
| 8 |
H |
0.045 |
|
|
|
| 9 |
H |
0.044 |
|
|
|
| 10 |
H |
-0.021 |
|
|
|
| 11 |
H |
-0.024 |
|
|
|
| 12 |
H |
0.028 |
|
|
|
| 13 |
H |
0.008 |
|
|
|
| 14 |
H |
0.025 |
|
|
|
| 15 |
H |
0.021 |
|
|
|
| 16 |
H |
0.027 |
|
|
|
| 17 |
H |
0.031 |
|
|
|
| 18 |
H |
0.030 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.040 |
0.032 |
-0.050 |
0.072 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.713 |
-0.091 |
-0.670 |
| y |
-0.091 |
-39.982 |
1.052 |
| z |
-0.670 |
1.052 |
-40.207 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.382 |
-0.091 |
-0.670 |
| y |
-0.091 |
-0.522 |
1.052 |
| z |
-0.670 |
1.052 |
-0.860 |
|
| Polar |
|---|
| 3z2-r2 | -1.719 |
|---|
| x2-y2 | 1.270 |
|---|
| xy | -0.091 |
|---|
| xz | -0.670 |
|---|
| yz | 1.052 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.790 |
-0.140 |
-0.899 |
| y |
-0.140 |
9.091 |
0.706 |
| z |
-0.899 |
0.706 |
8.644 |
<r2> (average value of r
2) Å
2
| <r2> |
299.289 |
| (<r2>)1/2 |
17.300 |
