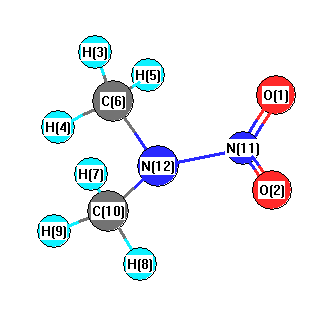
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3193 |
3081 |
2.33 |
|
|
|
| 2 |
A |
3190 |
3079 |
0.66 |
|
|
|
| 3 |
A |
3141 |
3031 |
27.75 |
|
|
|
| 4 |
A |
3138 |
3028 |
0.08 |
|
|
|
| 5 |
A |
3032 |
2926 |
44.67 |
|
|
|
| 6 |
A |
3026 |
2920 |
24.77 |
|
|
|
| 7 |
A |
1691 |
1631 |
287.23 |
|
|
|
| 8 |
A |
1490 |
1438 |
83.68 |
|
|
|
| 9 |
A |
1478 |
1426 |
31.06 |
|
|
|
| 10 |
A |
1469 |
1418 |
0.03 |
|
|
|
| 11 |
A |
1445 |
1394 |
11.07 |
|
|
|
| 12 |
A |
1444 |
1394 |
62.03 |
|
|
|
| 13 |
A |
1403 |
1354 |
6.84 |
|
|
|
| 14 |
A |
1364 |
1317 |
227.19 |
|
|
|
| 15 |
A |
1343 |
1296 |
1.73 |
|
|
|
| 16 |
A |
1274 |
1229 |
7.38 |
|
|
|
| 17 |
A |
1134 |
1094 |
38.26 |
|
|
|
| 18 |
A |
1107 |
1068 |
1.66 |
|
|
|
| 19 |
A |
1047 |
1011 |
11.27 |
|
|
|
| 20 |
A |
1018 |
983 |
65.33 |
|
|
|
| 21 |
A |
877 |
846 |
12.14 |
|
|
|
| 22 |
A |
796 |
768 |
18.23 |
|
|
|
| 23 |
A |
636 |
614 |
1.10 |
|
|
|
| 24 |
A |
625 |
603 |
8.95 |
|
|
|
| 25 |
A |
417 |
402 |
3.70 |
|
|
|
| 26 |
A |
361 |
349 |
2.31 |
|
|
|
| 27 |
A |
195 |
188 |
7.73 |
|
|
|
| 28 |
A |
136 |
131 |
0.20 |
|
|
|
| 29 |
A |
120 |
116 |
2.69 |
|
|
|
| 30 |
A |
108 |
104 |
0.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20849.2 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 20119.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.282 |
|
|
|
| 2 |
O |
-0.282 |
|
|
|
| 3 |
H |
0.060 |
|
|
|
| 4 |
H |
0.056 |
|
|
|
| 5 |
H |
0.085 |
|
|
|
| 6 |
C |
0.036 |
|
|
|
| 7 |
H |
0.060 |
|
|
|
| 8 |
H |
0.085 |
|
|
|
| 9 |
H |
0.056 |
|
|
|
| 10 |
C |
0.036 |
|
|
|
| 11 |
N |
0.343 |
|
|
|
| 12 |
N |
-0.252 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.367 |
0.000 |
0.106 |
4.368 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.759 |
0.000 |
-0.359 |
| y |
0.000 |
-35.377 |
-0.000 |
| z |
-0.359 |
-0.000 |
-34.441 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.850 |
0.000 |
-0.359 |
| y |
0.000 |
0.222 |
-0.000 |
| z |
-0.359 |
-0.000 |
1.628 |
|
| Polar |
|---|
| 3z2-r2 | 3.255 |
|---|
| x2-y2 | -1.382 |
|---|
| xy | 0.000 |
|---|
| xz | -0.359 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
148.354 |
| (<r2>)1/2 |
12.180 |