Jump to
S1C2
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -376.682729 |
| Energy at 298.15K | |
| HF Energy | -375.806145 |
| Nuclear repulsion energy | 189.980621 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3145 |
2998 |
|
|
|
|
| 2 |
A |
3107 |
2961 |
|
|
|
|
| 3 |
A |
3076 |
2932 |
|
|
|
|
| 4 |
A |
1491 |
1421 |
|
|
|
|
| 5 |
A |
1471 |
1402 |
|
|
|
|
| 6 |
A |
1413 |
1347 |
|
|
|
|
| 7 |
A |
1341 |
1279 |
|
|
|
|
| 8 |
A |
1262 |
1203 |
|
|
|
|
| 9 |
A |
1182 |
1126 |
|
|
|
|
| 10 |
A |
1149 |
1095 |
|
|
|
|
| 11 |
A |
1134 |
1081 |
|
|
|
|
| 12 |
A |
1105 |
1053 |
|
|
|
|
| 13 |
A |
916 |
873 |
|
|
|
|
| 14 |
A |
579 |
552 |
|
|
|
|
| 15 |
A |
472 |
450 |
|
|
|
|
| 16 |
A |
428 |
408 |
|
|
|
|
| 17 |
A |
238 |
227 |
|
|
|
|
| 18 |
A |
120 |
114 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11814.6 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 11260.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.774 |
-0.596 |
-0.284 |
| C2 |
0.470 |
0.022 |
0.330 |
| F3 |
-1.885 |
0.106 |
0.151 |
| F4 |
1.541 |
-0.760 |
-0.008 |
| F5 |
0.673 |
1.268 |
-0.183 |
| H6 |
-0.700 |
-0.537 |
-1.385 |
| H7 |
-0.854 |
-1.651 |
0.035 |
| H8 |
0.412 |
0.102 |
1.431 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5187 | 1.3841 | 2.3368 | 2.3617 | 1.1056 | 1.1053 | 2.1980 |
C2 | 1.5187 | | 2.3634 | 1.3676 | 1.3622 | 2.1499 | 2.1545 | 1.1054 | F3 | 1.3841 | 2.3634 | | 3.5367 | 2.8289 | 2.0437 | 2.0410 | 2.6289 | F4 | 2.3368 | 1.3676 | 3.5367 | | 2.2119 | 2.6394 | 2.5559 | 2.0214 | F5 | 2.3617 | 1.3622 | 2.8289 | 2.2119 | | 2.5664 | 3.3017 | 2.0079 | H6 | 1.1056 | 2.1499 | 2.0437 | 2.6394 | 2.5664 | | 1.8115 | 3.0938 | H7 | 1.1053 | 2.1545 | 2.0410 | 2.5559 | 3.3017 | 1.8115 | | 2.5738 | H8 | 2.1980 | 1.1054 | 2.6289 | 2.0214 | 2.0079 | 3.0938 | 2.5738 | |
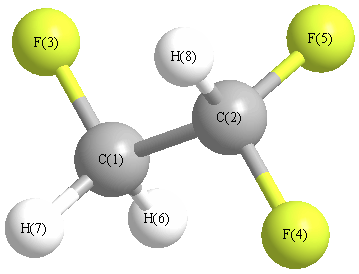 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.003 |
|
C1 |
C2 |
F5 |
110.004 |
| C1 |
C2 |
H8 |
112.827 |
|
C2 |
C1 |
F3 |
108.922 |
| C2 |
C1 |
H6 |
108.997 |
|
C2 |
C1 |
H7 |
109.373 |
| F3 |
C1 |
H6 |
109.842 |
|
F3 |
C1 |
H7 |
109.641 |
| F4 |
C2 |
F5 |
108.248 |
|
F4 |
C2 |
H8 |
109.188 |
| F5 |
C2 |
H8 |
108.473 |
|
H6 |
C1 |
H7 |
110.044 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -376.680146 |
| Energy at 298.15K | |
| HF Energy | -375.803121 |
| Nuclear repulsion energy | 192.794513 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3080 |
2935 |
|
|
|
|
| 2 |
A' |
3062 |
2918 |
|
|
|
|
| 3 |
A' |
1487 |
1417 |
|
|
|
|
| 4 |
A' |
1452 |
1384 |
|
|
|
|
| 5 |
A' |
1416 |
1349 |
|
|
|
|
| 6 |
A' |
1199 |
1143 |
|
|
|
|
| 7 |
A' |
1113 |
1061 |
|
|
|
|
| 8 |
A' |
886 |
845 |
|
|
|
|
| 9 |
A' |
759 |
724 |
|
|
|
|
| 10 |
A' |
517 |
493 |
|
|
|
|
| 11 |
A' |
230 |
220 |
|
|
|
|
| 12 |
A" |
3128 |
2982 |
|
|
|
|
| 13 |
A" |
1410 |
1344 |
|
|
|
|
| 14 |
A" |
1283 |
1222 |
|
|
|
|
| 15 |
A" |
1165 |
1111 |
|
|
|
|
| 16 |
A" |
966 |
921 |
|
|
|
|
| 17 |
A" |
366 |
349 |
|
|
|
|
| 18 |
A" |
115 |
109 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11817.2 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 11263.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.744 |
-0.834 |
0.000 |
| C2 |
0.357 |
0.634 |
0.000 |
| F3 |
-0.390 |
-1.621 |
0.000 |
| F4 |
-0.390 |
0.922 |
1.103 |
| F5 |
-0.390 |
0.922 |
-1.103 |
| H6 |
1.343 |
-1.049 |
-0.905 |
| H7 |
1.343 |
-1.049 |
0.905 |
| H8 |
1.254 |
1.284 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5177 | 1.3803 | 2.3639 | 2.3639 | 1.1063 | 1.1063 | 2.1786 |
C2 | 1.5177 | | 2.3749 | 1.3635 | 1.3635 | 2.1502 | 2.1502 | 1.1079 | F3 | 1.3803 | 2.3749 | | 2.7719 | 2.7719 | 2.0371 | 2.0371 | 3.3377 | F4 | 2.3639 | 1.3635 | 2.7719 | | 2.2067 | 3.3052 | 2.6329 | 2.0129 | F5 | 2.3639 | 1.3635 | 2.7719 | 2.2067 | | 2.6329 | 3.3052 | 2.0129 | H6 | 1.1063 | 2.1502 | 2.0371 | 3.3052 | 2.6329 | | 1.8092 | 2.5042 | H7 | 1.1063 | 2.1502 | 2.0371 | 2.6329 | 3.3052 | 1.8092 | | 2.5042 | H8 | 2.1786 | 1.1079 | 3.3377 | 2.0129 | 2.0129 | 2.5042 | 2.5042 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.144 |
|
C1 |
C2 |
F5 |
110.144 |
| C1 |
C2 |
H8 |
111.184 |
|
C2 |
C1 |
F3 |
109.980 |
| C2 |
C1 |
H6 |
109.049 |
|
C2 |
C1 |
H7 |
109.049 |
| F3 |
C1 |
H6 |
109.523 |
|
F3 |
C1 |
H7 |
109.523 |
| F4 |
C2 |
F5 |
108.036 |
|
F4 |
C2 |
H8 |
108.627 |
| F5 |
C2 |
H8 |
108.627 |
|
H6 |
C1 |
H7 |
109.700 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
