Jump to
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -377.160521 |
| Energy at 298.15K | |
| HF Energy | -377.160521 |
| Nuclear repulsion energy | 188.835684 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3033 |
3016 |
30.41 |
51.26 |
0.73 |
0.84 |
| 2 |
A |
2979 |
2962 |
40.10 |
107.80 |
0.32 |
0.49 |
| 3 |
A |
2969 |
2952 |
18.46 |
134.85 |
0.08 |
0.14 |
| 4 |
A |
1412 |
1404 |
1.88 |
9.79 |
0.71 |
0.83 |
| 5 |
A |
1385 |
1377 |
18.49 |
4.30 |
0.60 |
0.75 |
| 6 |
A |
1336 |
1329 |
21.92 |
6.11 |
0.73 |
0.84 |
| 7 |
A |
1270 |
1263 |
20.26 |
3.14 |
0.74 |
0.85 |
| 8 |
A |
1205 |
1198 |
9.58 |
8.82 |
0.73 |
0.85 |
| 9 |
A |
1116 |
1110 |
86.99 |
5.42 |
0.71 |
0.83 |
| 10 |
A |
1089 |
1083 |
6.61 |
1.74 |
0.40 |
0.57 |
| 11 |
A |
1077 |
1071 |
238.98 |
0.03 |
0.37 |
0.54 |
| 12 |
A |
1055 |
1049 |
13.01 |
6.06 |
0.57 |
0.73 |
| 13 |
A |
875 |
869 |
32.85 |
4.49 |
0.39 |
0.56 |
| 14 |
A |
554 |
551 |
3.02 |
2.11 |
0.47 |
0.64 |
| 15 |
A |
451 |
449 |
15.03 |
0.90 |
0.73 |
0.85 |
| 16 |
A |
409 |
407 |
4.52 |
1.80 |
0.56 |
0.72 |
| 17 |
A |
227 |
225 |
6.80 |
0.07 |
0.61 |
0.76 |
| 18 |
A |
109 |
108 |
6.20 |
0.02 |
0.73 |
0.84 |
Unscaled Zero Point Vibrational Energy (zpe) 11275.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 11210.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.779 |
-0.586 |
-0.288 |
| C2 |
0.470 |
0.021 |
0.330 |
| F3 |
-1.901 |
0.105 |
0.153 |
| F4 |
1.544 |
-0.774 |
-0.006 |
| F5 |
0.691 |
1.275 |
-0.182 |
| H6 |
-0.704 |
-0.517 |
-1.395 |
| H7 |
-0.854 |
-1.653 |
0.016 |
| H8 |
0.409 |
0.103 |
1.439 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5199 | 1.3897 | 2.3475 | 2.3746 | 1.1117 | 1.1113 | 2.2062 |
C2 | 1.5199 | | 2.3789 | 1.3782 | 1.3723 | 2.1545 | 2.1573 | 1.1134 | F3 | 1.3897 | 2.3789 | | 3.5590 | 2.8638 | 2.0533 | 2.0507 | 2.6440 | F4 | 2.3475 | 1.3782 | 3.5590 | | 2.2266 | 2.6547 | 2.5540 | 2.0355 | F5 | 2.3746 | 1.3723 | 2.8638 | 2.2266 | | 2.5752 | 3.3167 | 2.0197 | H6 | 1.1117 | 2.1545 | 2.0533 | 2.6547 | 2.5752 | | 1.8167 | 3.1067 | H7 | 1.1113 | 2.1573 | 2.0507 | 2.5540 | 3.3167 | 1.8167 | | 2.5892 | H8 | 2.2062 | 1.1134 | 2.6440 | 2.0355 | 2.0197 | 3.1067 | 2.5892 | |
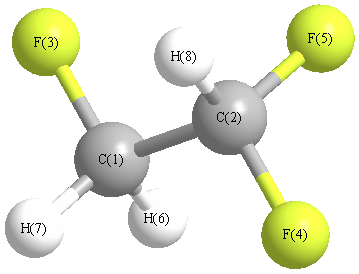 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.099 |
|
C1 |
C2 |
F5 |
110.273 |
| C1 |
C2 |
H8 |
112.913 |
|
C2 |
C1 |
F3 |
109.611 |
| C2 |
C1 |
H6 |
108.922 |
|
C2 |
C1 |
H7 |
109.162 |
| F3 |
C1 |
H6 |
109.850 |
|
F3 |
C1 |
H7 |
109.661 |
| F4 |
C2 |
F5 |
108.105 |
|
F4 |
C2 |
H8 |
109.103 |
| F5 |
C2 |
H8 |
108.238 |
|
H6 |
C1 |
H7 |
109.617 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.163 |
|
|
|
| 2 |
C |
0.281 |
|
|
|
| 3 |
F |
-0.205 |
|
|
|
| 4 |
F |
-0.179 |
|
|
|
| 5 |
F |
-0.170 |
|
|
|
| 6 |
H |
0.045 |
|
|
|
| 7 |
H |
0.038 |
|
|
|
| 8 |
H |
0.027 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.218 |
-1.227 |
0.297 |
1.281 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.320 |
1.226 |
1.100 |
| y |
1.226 |
-27.338 |
0.445 |
| z |
1.100 |
0.445 |
-24.814 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.244 |
1.226 |
1.100 |
| y |
1.226 |
0.729 |
0.445 |
| z |
1.100 |
0.445 |
4.515 |
|
| Polar |
|---|
| 3z2-r2 | 9.029 |
|---|
| x2-y2 | -3.982 |
|---|
| xy | 1.226 |
|---|
| xz | 1.100 |
|---|
| yz | 0.445 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.680 |
0.006 |
0.048 |
| y |
0.006 |
3.686 |
0.070 |
| z |
0.048 |
0.070 |
3.619 |
<r2> (average value of r
2) Å
2
| <r2> |
112.894 |
| (<r2>)1/2 |
10.625 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -377.158060 |
| Energy at 298.15K | |
| HF Energy | -377.158060 |
| Nuclear repulsion energy | 191.514580 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
2962 |
2945 |
53.12 |
150.78 |
0.02 |
0.04 |
| 2 |
A' |
2940 |
2923 |
30.04 |
68.91 |
0.60 |
0.75 |
| 3 |
A' |
1408 |
1400 |
4.87 |
8.94 |
0.72 |
0.84 |
| 4 |
A' |
1369 |
1361 |
27.02 |
3.71 |
0.45 |
0.62 |
| 5 |
A' |
1346 |
1338 |
28.43 |
4.40 |
0.66 |
0.80 |
| 6 |
A' |
1142 |
1135 |
115.60 |
4.36 |
0.43 |
0.60 |
| 7 |
A' |
1066 |
1060 |
22.86 |
3.17 |
0.71 |
0.83 |
| 8 |
A' |
847 |
843 |
30.64 |
6.20 |
0.22 |
0.36 |
| 9 |
A' |
734 |
729 |
43.42 |
3.00 |
0.57 |
0.73 |
| 10 |
A' |
496 |
493 |
9.57 |
1.81 |
0.75 |
0.86 |
| 11 |
A' |
217 |
216 |
1.58 |
0.15 |
0.42 |
0.59 |
| 12 |
A" |
3015 |
2997 |
27.28 |
67.18 |
0.75 |
0.86 |
| 13 |
A" |
1328 |
1320 |
32.22 |
2.07 |
0.75 |
0.86 |
| 14 |
A" |
1225 |
1218 |
12.55 |
16.50 |
0.75 |
0.86 |
| 15 |
A" |
1092 |
1085 |
102.71 |
1.22 |
0.75 |
0.86 |
| 16 |
A" |
912 |
907 |
60.54 |
3.10 |
0.75 |
0.86 |
| 17 |
A" |
351 |
349 |
0.11 |
0.29 |
0.75 |
0.86 |
| 18 |
A" |
103 |
103 |
1.47 |
0.02 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 11276.1 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 11210.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.739 |
-0.839 |
0.000 |
| C2 |
0.359 |
0.632 |
0.000 |
| F3 |
-0.391 |
-1.640 |
0.000 |
| F4 |
-0.391 |
0.935 |
1.110 |
| F5 |
-0.391 |
0.935 |
-1.110 |
| H6 |
1.347 |
-1.053 |
-0.907 |
| H7 |
1.347 |
-1.053 |
0.907 |
| H8 |
1.268 |
1.280 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5191 | 1.3855 | 2.3779 | 2.3779 | 1.1126 | 1.1126 | 2.1838 |
C2 | 1.5191 | | 2.3932 | 1.3739 | 1.3739 | 2.1536 | 2.1536 | 1.1162 | F3 | 1.3855 | 2.3932 | | 2.8041 | 2.8041 | 2.0467 | 2.0467 | 3.3589 | F4 | 2.3779 | 1.3739 | 2.8041 | | 2.2209 | 3.3228 | 2.6484 | 2.0263 | F5 | 2.3779 | 1.3739 | 2.8041 | 2.2209 | | 2.6484 | 3.3228 | 2.0263 | H6 | 1.1126 | 2.1536 | 2.0467 | 3.3228 | 2.6484 | | 1.8131 | 2.5043 | H7 | 1.1126 | 2.1536 | 2.0467 | 2.6484 | 3.3228 | 1.8131 | | 2.5043 | H8 | 2.1838 | 1.1162 | 3.3589 | 2.0263 | 2.0263 | 2.5043 | 2.5043 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.465 |
|
C1 |
C2 |
F5 |
110.465 |
| C1 |
C2 |
H8 |
111.007 |
|
C2 |
C1 |
F3 |
110.878 |
| C2 |
C1 |
H6 |
108.855 |
|
C2 |
C1 |
H7 |
108.855 |
| F3 |
C1 |
H6 |
109.543 |
|
F3 |
C1 |
H7 |
109.543 |
| F4 |
C2 |
F5 |
107.849 |
|
F4 |
C2 |
H8 |
108.481 |
| F5 |
C2 |
H8 |
108.481 |
|
H6 |
C1 |
H7 |
109.137 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.173 |
|
|
|
| 2 |
C |
0.285 |
|
|
|
| 3 |
F |
-0.198 |
|
|
|
| 4 |
F |
-0.172 |
|
|
|
| 5 |
F |
-0.172 |
|
|
|
| 6 |
H |
0.036 |
|
|
|
| 7 |
H |
0.036 |
|
|
|
| 8 |
H |
0.012 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.673 |
0.033 |
0.000 |
2.673 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.027 |
-0.245 |
0.000 |
| y |
-0.245 |
-28.426 |
0.000 |
| z |
0.000 |
0.000 |
-27.959 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.165 |
-0.245 |
0.000 |
| y |
-0.245 |
-1.932 |
0.000 |
| z |
0.000 |
0.000 |
-1.232 |
|
| Polar |
|---|
| 3z2-r2 | -2.465 |
|---|
| x2-y2 | 3.398 |
|---|
| xy | -0.245 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.677 |
0.000 |
0.000 |
| y |
0.000 |
3.709 |
0.000 |
| z |
0.000 |
0.000 |
3.639 |
<r2> (average value of r
2) Å
2
| <r2> |
104.620 |
| (<r2>)1/2 |
10.228 |
