return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O2 (2,3-Butanedione)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-305.660807
Energy at 298.15K-305.666956
Nuclear repulsion energy224.776363
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3196 3066 0.00      
2 Ag 3079 2954 0.00      
3 Ag 1825 1751 0.00      
4 Ag 1459 1400 0.00      
5 Ag 1413 1356 0.00      
6 Ag 1319 1266 0.00      
7 Ag 1019 978 0.00      
8 Ag 706 677 0.00      
9 Ag 535 513 0.00      
10 Ag 365 350 0.00      
11 Au 3161 3033 8.53      
12 Au 1466 1406 16.36      
13 Au 968 928 3.60      
14 Au 355 341 5.40      
15 Au 129 123 0.03      
16 Au 63 60 8.41      
17 Bg 3161 3032 0.00      
18 Bg 1470 1410 0.00      
19 Bg 1074 1031 0.00      
20 Bg 627 601 0.00      
21 Bg 126 121 0.00      
22 Bu 3197 3067 13.49      
23 Bu 3078 2953 4.14      
24 Bu 1806 1732 192.59      
25 Bu 1458 1399 23.65      
26 Bu 1395 1338 42.98      
27 Bu 1146 1100 74.82      
28 Bu 928 890 14.13      
29 Bu 540 518 41.54      
30 Bu 240 230 14.35      

Unscaled Zero Point Vibrational Energy (zpe) 20650.9 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 19812.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.17398 0.11175 0.06983

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.082 0.771 0.000
C2 0.082 -0.771 0.000
C3 1.200 1.578 0.000
C4 -1.200 -1.578 0.000
O5 -1.200 1.255 0.000
O6 1.200 -1.255 0.000
H7 0.959 2.651 0.000
H8 -0.959 -2.651 0.000
H9 1.806 1.319 0.886
H10 1.806 1.319 -0.886
H11 -1.806 -1.319 0.886
H12 -1.806 -1.319 -0.886

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 O6 H7 H8 H9 H10 H11 H12
C11.55021.51542.60121.21862.39772.14953.53262.15642.15642.84992.8499
C21.55022.60121.51542.39771.21863.53262.14952.84992.84992.15642.1564
C31.51542.60123.96512.42242.83301.10054.74851.10391.10394.26774.2677
C42.60121.51543.96512.83302.42244.74851.10054.26774.26771.10391.1039
O51.21862.39772.42242.83303.47352.57123.91403.13483.13482.78852.7885
O62.39771.21862.83302.42243.47353.91402.57122.78852.78853.13483.1348
H72.14953.53261.10054.74852.57123.91405.63881.81081.81084.91834.9183
H83.53262.14954.74851.10053.91402.57125.63884.91834.91831.81081.8108
H92.15642.84991.10394.26773.13482.78851.81084.91831.77154.47244.8105
H102.15642.84991.10394.26773.13482.78851.81084.91831.77154.81054.4724
H112.84992.15644.26771.10392.78853.13484.91831.81084.47244.81051.7715
H122.84992.15644.26771.10392.78853.13484.91831.81084.81054.47241.7715

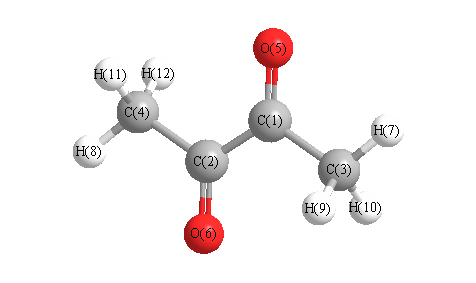
picture of 2,3-Butanedione state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 116.095 C1 C2 O6 119.509
C1 C3 H7 109.491 C1 C3 H9 109.836
C1 C3 H10 109.836 C2 C1 C3 116.095
C2 C1 O5 119.509 C2 C4 H8 109.491
C2 C4 H11 109.836 C2 C4 H12 109.836
C3 C1 O5 124.396 C4 C2 O6 124.396
H7 C3 H9 110.461 H7 C3 H10 110.461
H8 C4 H11 110.461 H8 C4 H12 110.461
H9 C3 H10 106.722 H11 C4 H12 106.722
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability