return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H11 (2-Methylbut-2-yl radical)

using model chemistry: ROHF/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at ROHF/cc-pVDZ
 hartrees
Energy at 0K-195.717896
Energy at 298.15K-195.729049
Nuclear repulsion energy181.608287
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at ROHF/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3262 2807 40.76      
2 A 3252 2799 81.03      
3 A 3246 2794 36.46      
4 A 3244 2792 34.19      
5 A 3218 2770 43.77      
6 A 3208 2761 31.41      
7 A 3191 2747 18.95      
8 A 3176 2733 35.51      
9 A 3132 2695 68.61      
10 A 3125 2690 33.79      
11 A 3106 2674 42.24      
12 A 1606 1383 0.79      
13 A 1598 1375 5.75      
14 A 1591 1369 16.32      
15 A 1591 1369 3.44      
16 A 1584 1363 1.66      
17 A 1577 1357 1.14      
18 A 1571 1352 0.63      
19 A 1528 1315 3.28      
20 A 1524 1311 2.00      
21 A 1514 1303 3.04      
22 A 1488 1281 0.12      
23 A 1403 1208 0.71      
24 A 1367 1177 5.84      
25 A 1328 1143 6.11      
26 A 1202 1035 2.12      
27 A 1140 981 0.21      
28 A 1117 961 3.01      
29 A 1066 917 0.64      
30 A 1053 906 1.09      
31 A 1002 863 0.19      
32 A 999 860 0.34      
33 A 827 712 0.86      
34 A 783 674 1.64      
35 A 478 411 0.41      
36 A 401 345 0.08      
37 A 348 300 1.66      
38 A 291 251 0.02      
39 A 234 201 0.53      
40 A 157 135 0.02      
41 A 141 121 0.04      
42 A 56 48 0.05      

Unscaled Zero Point Vibrational Energy (zpe) 33860.6 cm-1
Scaled (by 0.8607) Zero Point Vibrational Energy (zpe) 29143.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at ROHF/cc-pVDZ
ABC
0.24583 0.11357 0.08441

See section I.F.4 to change rotational constant units
Geometric Data calculated at ROHF/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.646 1.478 0.070
H2 0.208 2.043 -0.304
H3 -1.541 1.867 -0.421
H4 -0.740 1.701 1.142
C5 2.072 -0.033 -0.179
H6 2.210 0.958 0.257
H7 2.930 -0.641 0.118
H8 2.096 0.072 -1.266
C9 0.770 -0.688 0.282
H10 0.750 -1.722 -0.075
H11 0.774 -0.756 1.381
C12 -0.500 -0.002 -0.166
C13 -1.761 -0.822 -0.105
H14 -1.609 -1.823 -0.515
H15 -2.107 -0.946 0.931
H16 -2.574 -0.349 -0.661

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 C9 H10 H11 C12 C13 H14 H15 H16
C11.09101.09211.09953.12022.90904.15723.35892.59653.49442.95391.50592.56203.48792.95772.7538
H21.09101.76251.76292.79302.34463.84592.89392.84943.81113.31562.16943.48294.27763.97743.6862
H31.09211.76251.76414.08973.91865.15474.14323.51584.27203.93492.15462.71643.69173.17132.4558
H41.09951.76291.76413.55843.16814.47274.06182.95453.92722.89602.16132.99443.99052.98633.2885
C53.12022.79304.08973.55841.09171.09241.09301.52902.14722.15402.57273.91434.10714.41894.6813
H62.90902.34463.91863.16811.09171.75971.76612.18753.06992.50242.90624.36684.78694.76545.0428
H74.15723.84595.15474.47271.09241.75971.76652.16692.44042.50103.50054.69964.73255.11055.5658
H83.35892.89394.14324.06181.09301.76611.76652.17582.53953.07262.82094.12634.22884.85054.7275
C92.59652.84943.51582.95451.52902.18752.16692.17581.09401.10111.51102.56352.75332.95973.4900
H103.49443.81114.27203.92722.14723.06992.44042.53951.09401.74772.12832.66782.40203.12693.6437
H112.95393.31563.93492.89602.15402.50242.50103.07261.10111.74772.14102.93913.22692.92153.9420
C121.50592.16942.15462.16132.57272.90623.50052.82091.51102.12832.14101.50522.16022.16192.1594
C132.56203.48292.71642.99443.91434.36684.69964.12632.56352.66782.93911.50521.09241.09931.0925
H143.48794.27763.69173.99054.10714.78694.73254.22882.75332.40203.22692.16021.09241.76371.7681
H152.95773.97743.17132.98634.41894.76545.11054.85052.95973.12692.92152.16191.09931.76371.7634
H162.75383.68622.45583.28854.68135.04285.56584.72753.49003.64373.94202.15941.09251.76811.7634

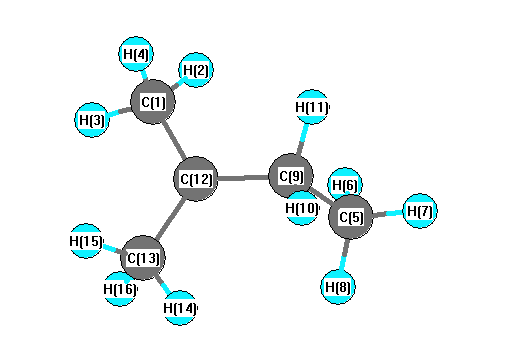
picture of 2-Methylbut-2-yl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C12 C9 118.770 C1 C12 C13 116.603
H2 C1 H3 107.678 H2 C1 H4 107.189
H2 C1 C12 112.324 H3 C1 H4 107.209
H3 C1 C12 111.052 H4 C1 C12 111.145
C5 C9 H10 108.758 C5 C9 H11 108.871
C5 C9 C12 115.612 H6 C5 H7 107.357
H6 C5 H8 107.880 H6 C5 C9 112.095
H7 C5 H8 107.865 H7 C5 C9 110.395
H8 C5 C9 111.071 C9 C12 C13 116.401
H10 C9 H11 105.537 H10 C9 C12 108.506
H11 C9 C12 109.084 C12 C13 H14 111.542
C12 C13 H15 111.256 C12 C13 H16 111.477
H14 C13 H15 107.165 H14 C13 H16 108.044
H15 C13 H16 107.134
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at ROHF/cc-pVDZ Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.028      
2 H 0.035      
3 H 0.030      
4 H 0.029      
5 C -0.034      
6 H 0.031      
7 H 0.029      
8 H 0.031      
9 C -0.029      
10 H 0.025      
11 H 0.024      
12 C -0.321      
13 C 0.032      
14 H 0.030      
15 H 0.029      
16 H 0.031      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å
Primitive
 xyz
x -34.713 -0.254 0.125
y -0.254 -34.435 0.086
z 0.125 0.086 -34.960
Traceless
 xyz
x -0.016 -0.254 0.125
y -0.254 0.402 0.086
z 0.125 0.086 -0.386
Polar
3z2-r2-0.773
x2-y2-0.278
xy-0.254
xz0.125
yz0.086


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 156.690
(<r2>)1/2 12.518