Jump to
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -154.171501 |
| Energy at 298.15K | |
| HF Energy | -154.171501 |
| Nuclear repulsion energy | 74.096747 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3632 |
3611 |
3.54 |
|
|
|
| 2 |
A' |
3079 |
3062 |
8.91 |
|
|
|
| 3 |
A' |
2880 |
2863 |
72.36 |
|
|
|
| 4 |
A' |
1444 |
1436 |
1.12 |
|
|
|
| 5 |
A' |
1411 |
1402 |
5.64 |
|
|
|
| 6 |
A' |
1368 |
1360 |
4.22 |
|
|
|
| 7 |
A' |
1194 |
1188 |
42.49 |
|
|
|
| 8 |
A' |
1030 |
1025 |
50.98 |
|
|
|
| 9 |
A' |
961 |
956 |
68.06 |
|
|
|
| 10 |
A' |
592 |
589 |
14.23 |
|
|
|
| 11 |
A' |
361 |
359 |
23.75 |
|
|
|
| 12 |
A" |
3192 |
3174 |
9.98 |
|
|
|
| 13 |
A" |
2911 |
2894 |
68.64 |
|
|
|
| 14 |
A" |
1219 |
1212 |
0.07 |
|
|
|
| 15 |
A" |
1116 |
1109 |
0.60 |
|
|
|
| 16 |
A" |
780 |
775 |
0.03 |
|
|
|
| 17 |
A" |
217 |
216 |
101.77 |
|
|
|
| 18 |
A" |
195i |
194i |
4.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13596.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 13517.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.122 |
-0.374 |
0.000 |
| C2 |
0.000 |
0.535 |
0.000 |
| C3 |
1.264 |
-0.255 |
0.000 |
| H4 |
-1.922 |
0.191 |
0.000 |
| H5 |
-0.025 |
1.199 |
0.900 |
| H6 |
-0.025 |
1.199 |
-0.900 |
| H7 |
1.683 |
-0.638 |
-0.942 |
| H8 |
1.683 |
-0.638 |
0.942 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4435 | 2.3887 | 0.9788 | 2.1176 | 2.1176 | 2.9709 | 2.9709 |
C2 | 1.4435 | | 1.4901 | 1.9525 | 1.1184 | 1.1184 | 2.2576 | 2.2576 | C3 | 2.3887 | 1.4901 | | 3.2167 | 2.1408 | 2.1408 | 1.1005 | 1.1005 | H4 | 0.9788 | 1.9525 | 3.2167 | | 2.3288 | 2.3288 | 3.8172 | 3.8172 | H5 | 2.1176 | 1.1184 | 2.1408 | 2.3288 | | 1.7993 | 3.1121 | 2.5087 | H6 | 2.1176 | 1.1184 | 2.1408 | 2.3288 | 1.7993 | | 2.5087 | 3.1121 | H7 | 2.9709 | 2.2576 | 1.1005 | 3.8172 | 3.1121 | 2.5087 | | 1.8849 | H8 | 2.9709 | 2.2576 | 1.1005 | 3.8172 | 2.5087 | 3.1121 | 1.8849 | |
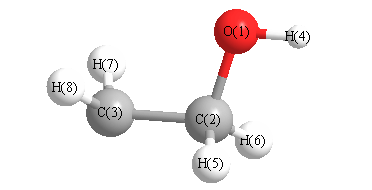 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
109.020 |
|
O1 |
C2 |
H5 |
110.858 |
| O1 |
C2 |
H6 |
110.858 |
|
C2 |
O1 |
H4 |
105.818 |
| C2 |
C3 |
H7 |
120.512 |
|
C2 |
C3 |
H8 |
120.512 |
| C3 |
C2 |
H5 |
109.485 |
|
C3 |
C2 |
H6 |
109.485 |
| H5 |
C2 |
H6 |
107.104 |
|
H7 |
C3 |
H8 |
117.831 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.252 |
|
|
|
| 2 |
C |
0.073 |
|
|
|
| 3 |
C |
-0.086 |
|
|
|
| 4 |
H |
0.135 |
|
|
|
| 5 |
H |
0.018 |
|
|
|
| 6 |
H |
0.018 |
|
|
|
| 7 |
H |
0.047 |
|
|
|
| 8 |
H |
0.047 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.246 |
1.347 |
0.000 |
1.369 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.775 |
-2.508 |
0.000 |
| y |
-2.508 |
-19.931 |
0.000 |
| z |
0.000 |
0.000 |
-18.831 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.605 |
-2.508 |
0.000 |
| y |
-2.508 |
-2.128 |
0.000 |
| z |
0.000 |
0.000 |
-0.478 |
|
| Polar |
|---|
| 3z2-r2 | -0.955 |
|---|
| x2-y2 | 3.156 |
|---|
| xy | -2.508 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.798 |
-0.577 |
0.000 |
| y |
-0.577 |
3.554 |
0.000 |
| z |
0.000 |
0.000 |
3.686 |
<r2> (average value of r
2) Å
2
| <r2> |
50.917 |
| (<r2>)1/2 |
7.136 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -154.174057 |
| Energy at 298.15K | -154.178628 |
| HF Energy | -154.174057 |
| Nuclear repulsion energy | 74.530816 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3675 |
3654 |
13.98 |
|
|
|
| 2 |
A |
3213 |
3194 |
4.53 |
|
|
|
| 3 |
A |
3092 |
3074 |
4.35 |
|
|
|
| 4 |
A |
2832 |
2815 |
75.15 |
|
|
|
| 5 |
A |
2763 |
2747 |
75.53 |
|
|
|
| 6 |
A |
1419 |
1411 |
10.58 |
|
|
|
| 7 |
A |
1392 |
1384 |
1.36 |
|
|
|
| 8 |
A |
1344 |
1336 |
0.62 |
|
|
|
| 9 |
A |
1232 |
1224 |
62.22 |
|
|
|
| 10 |
A |
1155 |
1148 |
2.67 |
|
|
|
| 11 |
A |
1097 |
1091 |
65.00 |
|
|
|
| 12 |
A |
1039 |
1033 |
9.51 |
|
|
|
| 13 |
A |
921 |
915 |
16.36 |
|
|
|
| 14 |
A |
825 |
820 |
13.65 |
|
|
|
| 15 |
A |
441 |
438 |
29.11 |
|
|
|
| 16 |
A |
391 |
389 |
25.74 |
|
|
|
| 17 |
A |
271 |
269 |
67.05 |
|
|
|
| 18 |
A |
148 |
147 |
30.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13623.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 13544.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.114 |
-0.382 |
-0.060 |
| C2 |
-0.010 |
0.520 |
0.034 |
| C3 |
1.252 |
-0.256 |
-0.017 |
| H4 |
-1.916 |
0.143 |
0.119 |
| H5 |
-0.054 |
1.109 |
0.992 |
| H6 |
-0.019 |
1.283 |
-0.788 |
| H7 |
2.207 |
0.248 |
-0.221 |
| H8 |
1.244 |
-1.316 |
0.277 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4289 | 2.3697 | 0.9747 | 2.1101 | 2.1212 | 3.3840 | 2.5585 |
C2 | 1.4289 | | 1.4823 | 1.9451 | 1.1258 | 1.1211 | 2.2476 | 2.2363 | C3 | 2.3697 | 1.4823 | | 3.1958 | 2.1419 | 2.1398 | 1.0992 | 1.0995 | H4 | 0.9747 | 1.9451 | 3.1958 | | 2.2720 | 2.3920 | 4.1381 | 3.4837 | H5 | 2.1101 | 1.1258 | 2.1419 | 2.2720 | | 1.7892 | 2.7063 | 2.8418 | H6 | 2.1212 | 1.1211 | 2.1398 | 2.3920 | 1.7892 | | 2.5191 | 3.0791 | H7 | 3.3840 | 2.2476 | 1.0992 | 4.1381 | 2.7063 | 2.5191 | | 1.9031 | H8 | 2.5585 | 2.2363 | 1.0995 | 3.4837 | 2.8418 | 3.0791 | 1.9031 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
108.965 |
|
O1 |
C2 |
H5 |
110.820 |
| O1 |
C2 |
H6 |
112.012 |
|
C2 |
O1 |
H4 |
106.513 |
| C2 |
C3 |
H7 |
120.350 |
|
C2 |
C3 |
H8 |
119.304 |
| C3 |
C2 |
H5 |
109.664 |
|
C3 |
C2 |
H6 |
109.777 |
| H5 |
C2 |
H6 |
105.553 |
|
H7 |
C3 |
H8 |
119.893 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.240 |
|
|
|
| 2 |
C |
0.065 |
|
|
|
| 3 |
C |
-0.093 |
|
|
|
| 4 |
H |
0.138 |
|
|
|
| 5 |
H |
0.027 |
|
|
|
| 6 |
H |
0.024 |
|
|
|
| 7 |
H |
0.034 |
|
|
|
| 8 |
H |
0.046 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.444 |
1.508 |
0.474 |
1.642 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.181 |
-1.557 |
-0.686 |
| y |
-1.557 |
-19.141 |
-0.360 |
| z |
-0.686 |
-0.360 |
-20.194 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.486 |
-1.557 |
-0.686 |
| y |
-1.557 |
-0.954 |
-0.360 |
| z |
-0.686 |
-0.360 |
-2.533 |
|
| Polar |
|---|
| 3z2-r2 | -5.065 |
|---|
| x2-y2 | 2.960 |
|---|
| xy | -1.557 |
|---|
| xz | -0.686 |
|---|
| yz | -0.360 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.977 |
-0.332 |
-0.211 |
| y |
-0.332 |
4.175 |
-0.113 |
| z |
-0.211 |
-0.113 |
3.002 |
<r2> (average value of r
2) Å
2
| <r2> |
50.679 |
| (<r2>)1/2 |
7.119 |
