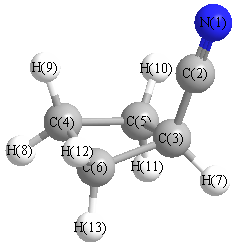
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3051 |
3056 |
47.58 |
|
|
|
| 2 |
A' |
3027 |
3031 |
9.13 |
|
|
|
| 3 |
A' |
2990 |
2995 |
1.97 |
|
|
|
| 4 |
A' |
2979 |
2984 |
23.46 |
|
|
|
| 5 |
A' |
2978 |
2982 |
13.62 |
|
|
|
| 6 |
A' |
2243 |
2247 |
11.29 |
|
|
|
| 7 |
A' |
1442 |
1444 |
0.65 |
|
|
|
| 8 |
A' |
1418 |
1420 |
2.07 |
|
|
|
| 9 |
A' |
1298 |
1300 |
1.47 |
|
|
|
| 10 |
A' |
1229 |
1231 |
1.12 |
|
|
|
| 11 |
A' |
1188 |
1190 |
0.07 |
|
|
|
| 12 |
A' |
1083 |
1085 |
1.41 |
|
|
|
| 13 |
A' |
1034 |
1036 |
0.01 |
|
|
|
| 14 |
A' |
925 |
927 |
0.31 |
|
|
|
| 15 |
A' |
858 |
860 |
0.84 |
|
|
|
| 16 |
A' |
726 |
727 |
1.76 |
|
|
|
| 17 |
A' |
579 |
580 |
0.84 |
|
|
|
| 18 |
A' |
508 |
509 |
0.73 |
|
|
|
| 19 |
A' |
246 |
246 |
1.20 |
|
|
|
| 20 |
A' |
112 |
112 |
2.08 |
|
|
|
| 21 |
A" |
3044 |
3048 |
10.51 |
|
|
|
| 22 |
A" |
2986 |
2991 |
52.94 |
|
|
|
| 23 |
A" |
1410 |
1413 |
1.05 |
|
|
|
| 24 |
A" |
1231 |
1233 |
0.27 |
|
|
|
| 25 |
A" |
1210 |
1212 |
0.07 |
|
|
|
| 26 |
A" |
1194 |
1196 |
0.07 |
|
|
|
| 27 |
A" |
1160 |
1162 |
0.15 |
|
|
|
| 28 |
A" |
989 |
991 |
0.06 |
|
|
|
| 29 |
A" |
912 |
914 |
0.11 |
|
|
|
| 30 |
A" |
905 |
906 |
1.35 |
|
|
|
| 31 |
A" |
763 |
764 |
0.45 |
|
|
|
| 32 |
A" |
517 |
518 |
0.15 |
|
|
|
| 33 |
A" |
175 |
176 |
3.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23205.1 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 23242.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.084 |
|
|
|
| 2 |
C |
-0.084 |
|
|
|
| 3 |
C |
-0.043 |
|
|
|
| 4 |
C |
0.008 |
|
|
|
| 5 |
C |
0.075 |
|
|
|
| 6 |
C |
0.075 |
|
|
|
| 7 |
H |
0.013 |
|
|
|
| 8 |
H |
0.004 |
|
|
|
| 9 |
H |
0.000 |
|
|
|
| 10 |
H |
0.012 |
|
|
|
| 11 |
H |
0.007 |
|
|
|
| 12 |
H |
0.012 |
|
|
|
| 13 |
H |
0.007 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.729 |
-2.822 |
0.000 |
3.925 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.522 |
-6.041 |
0.000 |
| y |
-6.041 |
-42.387 |
0.000 |
| z |
0.000 |
0.000 |
-35.758 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.450 |
-6.041 |
0.000 |
| y |
-6.041 |
-4.747 |
0.000 |
| z |
0.000 |
0.000 |
5.196 |
|
| Polar |
|---|
| 3z2-r2 | 10.393 |
|---|
| x2-y2 | 2.865 |
|---|
| xy | -6.041 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.421 |
1.841 |
0.000 |
| y |
1.841 |
9.309 |
0.000 |
| z |
0.000 |
0.000 |
7.463 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |