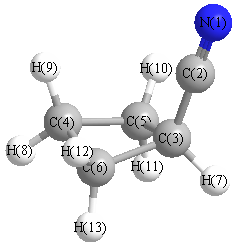
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3076 |
3059 |
32.25 |
|
|
|
| 2 |
A' |
3055 |
3037 |
8.39 |
|
|
|
| 3 |
A' |
3009 |
2991 |
1.87 |
|
|
|
| 4 |
A' |
2999 |
2982 |
20.52 |
|
|
|
| 5 |
A' |
2997 |
2980 |
9.68 |
|
|
|
| 6 |
A' |
2263 |
2250 |
13.46 |
|
|
|
| 7 |
A' |
1433 |
1425 |
0.91 |
|
|
|
| 8 |
A' |
1406 |
1397 |
3.51 |
|
|
|
| 9 |
A' |
1295 |
1288 |
1.11 |
|
|
|
| 10 |
A' |
1223 |
1216 |
0.60 |
|
|
|
| 11 |
A' |
1188 |
1181 |
0.08 |
|
|
|
| 12 |
A' |
1079 |
1072 |
1.61 |
|
|
|
| 13 |
A' |
1062 |
1055 |
0.02 |
|
|
|
| 14 |
A' |
949 |
944 |
0.51 |
|
|
|
| 15 |
A' |
872 |
867 |
0.85 |
|
|
|
| 16 |
A' |
731 |
727 |
1.73 |
|
|
|
| 17 |
A' |
570 |
567 |
1.31 |
|
|
|
| 18 |
A' |
516 |
513 |
0.68 |
|
|
|
| 19 |
A' |
252 |
251 |
1.00 |
|
|
|
| 20 |
A' |
118 |
117 |
2.25 |
|
|
|
| 21 |
A" |
3069 |
3052 |
11.70 |
|
|
|
| 22 |
A" |
3005 |
2988 |
41.67 |
|
|
|
| 23 |
A" |
1397 |
1389 |
2.27 |
|
|
|
| 24 |
A" |
1226 |
1218 |
0.95 |
|
|
|
| 25 |
A" |
1200 |
1193 |
0.29 |
|
|
|
| 26 |
A" |
1186 |
1179 |
0.12 |
|
|
|
| 27 |
A" |
1158 |
1151 |
0.25 |
|
|
|
| 28 |
A" |
1002 |
996 |
0.01 |
|
|
|
| 29 |
A" |
934 |
928 |
0.10 |
|
|
|
| 30 |
A" |
921 |
916 |
1.88 |
|
|
|
| 31 |
A" |
760 |
755 |
0.74 |
|
|
|
| 32 |
A" |
524 |
521 |
0.13 |
|
|
|
| 33 |
A" |
173 |
172 |
3.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23323.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 23188.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.118 |
|
|
|
| 2 |
C |
-0.031 |
|
|
|
| 3 |
C |
-0.111 |
|
|
|
| 4 |
C |
-0.053 |
|
|
|
| 5 |
C |
0.026 |
|
|
|
| 6 |
C |
0.026 |
|
|
|
| 7 |
H |
0.047 |
|
|
|
| 8 |
H |
0.034 |
|
|
|
| 9 |
H |
0.028 |
|
|
|
| 10 |
H |
0.042 |
|
|
|
| 11 |
H |
0.035 |
|
|
|
| 12 |
H |
0.042 |
|
|
|
| 13 |
H |
0.035 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.744 |
-2.856 |
0.000 |
3.961 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.249 |
-6.051 |
0.000 |
| y |
-6.051 |
-42.133 |
0.000 |
| z |
0.000 |
0.000 |
-35.521 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.422 |
-6.051 |
0.000 |
| y |
-6.051 |
-4.748 |
0.000 |
| z |
0.000 |
0.000 |
5.170 |
|
| Polar |
|---|
| 3z2-r2 | 10.341 |
|---|
| x2-y2 | 2.884 |
|---|
| xy | -6.051 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.408 |
1.860 |
0.000 |
| y |
1.860 |
9.270 |
0.000 |
| z |
0.000 |
0.000 |
7.431 |
<r2> (average value of r
2) Å
2
| <r2> |
158.783 |
| (<r2>)1/2 |
12.601 |