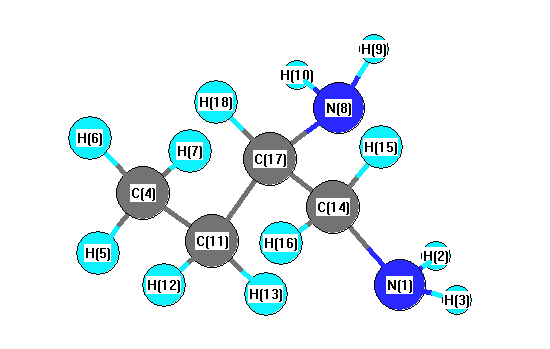
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3569 |
3444 |
2.21 |
|
|
|
| 2 |
A |
3568 |
3443 |
0.79 |
|
|
|
| 3 |
A |
3487 |
3365 |
1.70 |
|
|
|
| 4 |
A |
3479 |
3357 |
6.60 |
|
|
|
| 5 |
A |
3122 |
3013 |
38.85 |
|
|
|
| 6 |
A |
3118 |
3009 |
35.41 |
|
|
|
| 7 |
A |
3097 |
2989 |
5.96 |
|
|
|
| 8 |
A |
3054 |
2947 |
53.80 |
|
|
|
| 9 |
A |
3049 |
2942 |
7.98 |
|
|
|
| 10 |
A |
3031 |
2925 |
32.48 |
|
|
|
| 11 |
A |
2931 |
2829 |
127.65 |
|
|
|
| 12 |
A |
2910 |
2808 |
70.73 |
|
|
|
| 13 |
A |
1634 |
1577 |
31.09 |
|
|
|
| 14 |
A |
1615 |
1559 |
39.34 |
|
|
|
| 15 |
A |
1485 |
1433 |
4.41 |
|
|
|
| 16 |
A |
1477 |
1425 |
7.48 |
|
|
|
| 17 |
A |
1469 |
1418 |
5.07 |
|
|
|
| 18 |
A |
1453 |
1402 |
1.80 |
|
|
|
| 19 |
A |
1426 |
1376 |
0.99 |
|
|
|
| 20 |
A |
1415 |
1365 |
9.88 |
|
|
|
| 21 |
A |
1394 |
1345 |
3.01 |
|
|
|
| 22 |
A |
1374 |
1326 |
11.83 |
|
|
|
| 23 |
A |
1330 |
1284 |
1.65 |
|
|
|
| 24 |
A |
1293 |
1248 |
0.59 |
|
|
|
| 25 |
A |
1280 |
1236 |
1.98 |
|
|
|
| 26 |
A |
1256 |
1212 |
1.33 |
|
|
|
| 27 |
A |
1190 |
1148 |
4.57 |
|
|
|
| 28 |
A |
1158 |
1117 |
6.74 |
|
|
|
| 29 |
A |
1114 |
1075 |
2.33 |
|
|
|
| 30 |
A |
1075 |
1038 |
3.01 |
|
|
|
| 31 |
A |
1070 |
1033 |
3.09 |
|
|
|
| 32 |
A |
1012 |
977 |
1.35 |
|
|
|
| 33 |
A |
984 |
949 |
26.92 |
|
|
|
| 34 |
A |
936 |
904 |
28.32 |
|
|
|
| 35 |
A |
920 |
888 |
44.86 |
|
|
|
| 36 |
A |
874 |
843 |
120.91 |
|
|
|
| 37 |
A |
823 |
794 |
15.94 |
|
|
|
| 38 |
A |
767 |
740 |
0.68 |
|
|
|
| 39 |
A |
622 |
600 |
16.36 |
|
|
|
| 40 |
A |
446 |
431 |
5.27 |
|
|
|
| 41 |
A |
383 |
369 |
5.02 |
|
|
|
| 42 |
A |
341 |
330 |
35.11 |
|
|
|
| 43 |
A |
281 |
271 |
28.87 |
|
|
|
| 44 |
A |
263 |
254 |
24.08 |
|
|
|
| 45 |
A |
239 |
231 |
9.09 |
|
|
|
| 46 |
A |
216 |
209 |
4.56 |
|
|
|
| 47 |
A |
147 |
142 |
6.67 |
|
|
|
| 48 |
A |
82 |
79 |
1.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36629.3 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 35347.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.204 |
|
|
|
| 2 |
H |
0.088 |
|
|
|
| 3 |
H |
0.081 |
|
|
|
| 4 |
C |
-0.087 |
|
|
|
| 5 |
H |
0.043 |
|
|
|
| 6 |
H |
0.036 |
|
|
|
| 7 |
H |
0.040 |
|
|
|
| 8 |
N |
-0.178 |
|
|
|
| 9 |
H |
0.086 |
|
|
|
| 10 |
H |
0.082 |
|
|
|
| 11 |
C |
-0.031 |
|
|
|
| 12 |
H |
0.028 |
|
|
|
| 13 |
H |
0.041 |
|
|
|
| 14 |
C |
0.047 |
|
|
|
| 15 |
H |
0.003 |
|
|
|
| 16 |
H |
0.030 |
|
|
|
| 17 |
C |
-0.104 |
|
|
|
| 18 |
H |
-0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.066 |
1.390 |
1.085 |
1.765 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.638 |
0.350 |
0.637 |
| y |
0.350 |
-38.164 |
-0.353 |
| z |
0.637 |
-0.353 |
-40.580 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.734 |
0.350 |
0.637 |
| y |
0.350 |
-0.055 |
-0.353 |
| z |
0.637 |
-0.353 |
-3.679 |
|
| Polar |
|---|
| 3z2-r2 | -7.359 |
|---|
| x2-y2 | 2.526 |
|---|
| xy | 0.350 |
|---|
| xz | 0.637 |
|---|
| yz | -0.353 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
205.155 |
| (<r2>)1/2 |
14.323 |