return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH7N3 (triaminomethane)

using model chemistry: QCISD(T)=FULL/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C3 1A
Energy calculated at QCISD(T)=FULL/cc-pVDZ
 hartrees
Energy at 0K-206.030286
Energy at 298.15K-206.040516
HF Energy-205.294593
Nuclear repulsion energy137.198548
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3528 3383        
2 A 3432 3291        
3 A 3061 2936        
4 A 1641 1573        
5 A 1380 1323        
6 A 989 948        
7 A 887 851        
8 A 555 532        
9 A 305 292        
10 E 3528 3383        
10 E 3528 3383        
11 E 3436 3295        
11 E 3436 3295        
12 E 1654 1586        
12 E 1653 1586        
13 E 1424 1366        
13 E 1424 1366        
14 E 1242 1191        
14 E 1242 1191        
15 E 1077 1033        
15 E 1077 1033        
16 E 967 927        
16 E 966 926        
17 E 440 422        
17 E 439 421        
18 E 298 286        
18 E 298 285        

Unscaled Zero Point Vibrational Energy (zpe) 21951.2 cm-1
Scaled (by 0.959) Zero Point Vibrational Energy (zpe) 21051.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)=FULL/cc-pVDZ
ABC
0.28987 0.28987 0.16700

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)=FULL/cc-pVDZ

Point Group is C3

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.372
H2 0.000 0.000 1.479
N3 0.000 1.402 -0.055
N4 1.215 -0.701 -0.055
N5 -1.215 -0.701 -0.055
H6 0.899 1.807 0.233
H7 1.115 -1.682 0.233
H8 -2.014 -0.125 0.233
H9 0.038 1.409 -1.084
H10 1.201 -0.737 -1.084
H11 -1.239 -0.671 -1.084

Atom - Atom Distances (Å)
  C1 H2 N3 N4 N5 H6 H7 H8 H9 H10 H11
C11.10711.46591.46591.46592.02312.02312.02312.02562.02562.0256
H21.10712.07822.07822.07822.37182.37182.37182.92422.92422.9242
N31.46592.07822.42912.42911.02723.29272.54431.02932.66052.6254
N41.46592.07822.42912.42912.54431.02723.29272.62541.02932.6605
N51.46592.07822.42912.42913.29272.54431.02722.66052.62541.0293
H62.02312.37181.02722.54433.29273.49593.49591.62282.88063.5279
H72.02312.37183.29271.02722.54433.49593.49593.52791.62282.8806
H82.02312.37182.54433.29271.02723.49593.49592.88063.52791.6228
H92.02562.92421.02932.62542.66051.62283.52792.88062.44052.4405
H102.02562.92422.66051.02932.62542.88061.62283.52792.44052.4405
H112.02562.92422.62542.66051.02933.52792.88061.62282.44052.4405

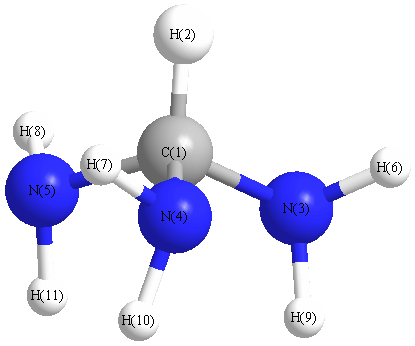
picture of triaminomethane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N3 H6 107.169 C1 N3 H9 107.244
C1 N4 H7 107.169 C1 N4 H10 107.244
C1 N5 H8 107.169 C1 N5 H11 107.244
H2 C1 N3 106.918 H2 C1 N4 106.918
H2 C1 N5 106.918 N3 C1 N4 111.899
N3 C1 N5 111.899 N4 C1 N5 111.899
H6 N3 H9 104.200 H7 N4 H10 104.200
H8 N5 H11 104.200
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability