Jump to
S1C2
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -217.790778 |
| Energy at 298.15K | -217.798656 |
| HF Energy | -217.126379 |
| Nuclear repulsion energy | 127.518485 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3160 |
3023 |
25.86 |
|
|
|
| 2 |
A' |
3082 |
2948 |
25.79 |
|
|
|
| 3 |
A' |
3068 |
2936 |
50.08 |
|
|
|
| 4 |
A' |
3064 |
2931 |
11.01 |
|
|
|
| 5 |
A' |
1527 |
1461 |
1.93 |
|
|
|
| 6 |
A' |
1512 |
1447 |
5.42 |
|
|
|
| 7 |
A' |
1496 |
1431 |
0.44 |
|
|
|
| 8 |
A' |
1460 |
1397 |
17.06 |
|
|
|
| 9 |
A' |
1426 |
1364 |
2.44 |
|
|
|
| 10 |
A' |
1343 |
1285 |
2.99 |
|
|
|
| 11 |
A' |
1154 |
1104 |
3.59 |
|
|
|
| 12 |
A' |
1125 |
1076 |
90.13 |
|
|
|
| 13 |
A' |
1073 |
1026 |
8.54 |
|
|
|
| 14 |
A' |
913 |
873 |
7.05 |
|
|
|
| 15 |
A' |
456 |
436 |
5.86 |
|
|
|
| 16 |
A' |
268 |
257 |
3.34 |
|
|
|
| 17 |
A" |
3151 |
3015 |
74.03 |
|
|
|
| 18 |
A" |
3132 |
2996 |
9.36 |
|
|
|
| 19 |
A" |
3112 |
2977 |
12.99 |
|
|
|
| 20 |
A" |
1502 |
1437 |
6.08 |
|
|
|
| 21 |
A" |
1324 |
1267 |
0.00 |
|
|
|
| 22 |
A" |
1282 |
1227 |
0.81 |
|
|
|
| 23 |
A" |
1206 |
1154 |
2.10 |
|
|
|
| 24 |
A" |
897 |
858 |
1.76 |
|
|
|
| 25 |
A" |
770 |
737 |
0.95 |
|
|
|
| 26 |
A" |
240 |
229 |
0.00 |
|
|
|
| 27 |
A" |
134 |
128 |
3.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21438.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 20509.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
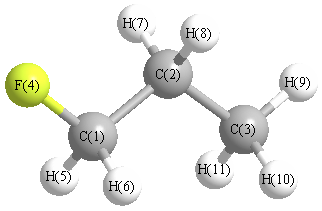
Geometric Data calculated at CCD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.131 |
-0.787 |
0.000 |
| C2 |
0.000 |
0.729 |
0.000 |
| C3 |
-1.468 |
1.173 |
0.000 |
| F4 |
1.469 |
-1.152 |
0.000 |
| H5 |
-0.347 |
-1.225 |
0.897 |
| H6 |
-0.347 |
-1.225 |
-0.897 |
| H7 |
0.524 |
1.127 |
-0.889 |
| H8 |
0.524 |
1.127 |
0.889 |
| H9 |
-1.549 |
2.274 |
0.000 |
| H10 |
-2.000 |
0.797 |
-0.893 |
| H11 |
-2.000 |
0.797 |
0.893 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5220 | 2.5296 | 1.3866 | 1.1066 | 1.1066 | 2.1469 | 2.1469 | 3.4912 | 2.8010 | 2.8010 |
C2 | 1.5220 | | 1.5336 | 2.3864 | 2.1779 | 2.1779 | 1.1057 | 1.1057 | 2.1871 | 2.1913 | 2.1913 | C3 | 2.5296 | 1.5336 | | 3.7455 | 2.7950 | 2.7950 | 2.1817 | 2.1817 | 1.1033 | 1.1057 | 1.1057 | F4 | 1.3866 | 2.3864 | 3.7455 | | 2.0263 | 2.0263 | 2.6221 | 2.6221 | 4.5648 | 4.0773 | 4.0773 | H5 | 1.1066 | 2.1779 | 2.7950 | 2.0263 | | 1.7946 | 3.0789 | 2.5081 | 3.8063 | 3.1662 | 2.6113 | H6 | 1.1066 | 2.1779 | 2.7950 | 2.0263 | 1.7946 | | 2.5081 | 3.0789 | 3.8063 | 2.6113 | 3.1662 | H7 | 2.1469 | 1.1057 | 2.1817 | 2.6221 | 3.0789 | 2.5081 | | 1.7771 | 2.5298 | 2.5454 | 3.1071 | H8 | 2.1469 | 1.1057 | 2.1817 | 2.6221 | 2.5081 | 3.0789 | 1.7771 | | 2.5298 | 3.1071 | 2.5454 | H9 | 3.4912 | 2.1871 | 1.1033 | 4.5648 | 3.8063 | 3.8063 | 2.5298 | 2.5298 | | 1.7840 | 1.7840 | H10 | 2.8010 | 2.1913 | 1.1057 | 4.0773 | 3.1662 | 2.6113 | 2.5454 | 3.1071 | 1.7840 | | 1.7866 | H11 | 2.8010 | 2.1913 | 1.1057 | 4.0773 | 2.6113 | 3.1662 | 3.1071 | 2.5454 | 1.7840 | 1.7866 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.755 |
|
C1 |
C2 |
H7 |
108.539 |
| C1 |
C2 |
H8 |
108.539 |
|
C2 |
C1 |
F4 |
110.173 |
| C2 |
C1 |
H5 |
110.911 |
|
C2 |
C1 |
H6 |
110.911 |
| C2 |
C3 |
H9 |
111.029 |
|
C2 |
C3 |
H10 |
111.213 |
| C2 |
C3 |
H11 |
111.213 |
|
C3 |
C2 |
H7 |
110.453 |
| C3 |
C2 |
H8 |
110.453 |
|
F4 |
C1 |
H5 |
108.200 |
| F4 |
C1 |
H6 |
108.200 |
|
H5 |
C1 |
H6 |
108.353 |
| H7 |
C2 |
H8 |
106.951 |
|
H9 |
C3 |
H10 |
107.719 |
| H9 |
C3 |
H11 |
107.719 |
|
H10 |
C3 |
H11 |
107.780 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -217.791521 |
| Energy at 298.15K | |
| HF Energy | -217.126897 |
| Nuclear repulsion energy | 130.005104 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3171 |
3034 |
21.29 |
|
|
|
| 2 |
A |
3153 |
3017 |
43.12 |
|
|
|
| 3 |
A |
3131 |
2995 |
48.72 |
|
|
|
| 4 |
A |
3118 |
2983 |
18.38 |
|
|
|
| 5 |
A |
3076 |
2943 |
20.29 |
|
|
|
| 6 |
A |
3070 |
2937 |
34.30 |
|
|
|
| 7 |
A |
3067 |
2935 |
26.64 |
|
|
|
| 8 |
A |
1522 |
1456 |
0.39 |
|
|
|
| 9 |
A |
1511 |
1445 |
7.82 |
|
|
|
| 10 |
A |
1497 |
1432 |
6.14 |
|
|
|
| 11 |
A |
1480 |
1416 |
1.70 |
|
|
|
| 12 |
A |
1449 |
1386 |
17.20 |
|
|
|
| 13 |
A |
1429 |
1367 |
2.31 |
|
|
|
| 14 |
A |
1388 |
1328 |
1.30 |
|
|
|
| 15 |
A |
1312 |
1255 |
1.10 |
|
|
|
| 16 |
A |
1287 |
1232 |
1.55 |
|
|
|
| 17 |
A |
1191 |
1140 |
2.01 |
|
|
|
| 18 |
A |
1149 |
1099 |
39.47 |
|
|
|
| 19 |
A |
1128 |
1079 |
23.04 |
|
|
|
| 20 |
A |
1003 |
960 |
31.97 |
|
|
|
| 21 |
A |
927 |
886 |
2.14 |
|
|
|
| 22 |
A |
899 |
860 |
4.30 |
|
|
|
| 23 |
A |
777 |
744 |
0.45 |
|
|
|
| 24 |
A |
484 |
463 |
3.97 |
|
|
|
| 25 |
A |
319 |
306 |
1.05 |
|
|
|
| 26 |
A |
232 |
222 |
1.99 |
|
|
|
| 27 |
A |
146 |
140 |
2.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21456.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 20527.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.806 |
0.512 |
0.294 |
| C2 |
-0.595 |
0.668 |
-0.280 |
| C3 |
-1.522 |
-0.487 |
0.119 |
| F4 |
1.380 |
-0.665 |
-0.172 |
| H5 |
1.460 |
1.352 |
-0.004 |
| H6 |
0.779 |
0.460 |
1.400 |
| H7 |
-0.516 |
0.730 |
-1.382 |
| H8 |
-1.004 |
1.635 |
0.070 |
| H9 |
-2.522 |
-0.371 |
-0.335 |
| H10 |
-1.099 |
-1.450 |
-0.213 |
| H11 |
-1.649 |
-0.531 |
1.216 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5223 | 2.5400 | 1.3900 | 1.1052 | 1.1067 | 2.1463 | 2.1421 | 3.5007 | 2.7816 | 2.8224 |
C2 | 1.5223 | | 1.5335 | 2.3846 | 2.1829 | 2.1795 | 1.1062 | 1.1071 | 2.1902 | 2.1776 | 2.1878 | C3 | 2.5400 | 1.5335 | | 2.9221 | 3.5055 | 2.7985 | 2.1777 | 2.1849 | 1.1037 | 1.1032 | 1.1059 | F4 | 1.3900 | 2.3846 | 2.9221 | | 2.0254 | 2.0243 | 2.6463 | 3.3212 | 3.9161 | 2.6002 | 3.3345 | H5 | 1.1052 | 2.1829 | 3.5055 | 2.0254 | | 1.7975 | 2.4877 | 2.4808 | 4.3513 | 3.8000 | 3.8339 | H6 | 1.1067 | 2.1795 | 2.7985 | 2.0243 | 1.7975 | | 3.0798 | 2.5154 | 3.8202 | 3.1263 | 2.6283 | H7 | 2.1463 | 1.1062 | 2.1777 | 2.6463 | 2.4877 | 3.0798 | | 1.7786 | 2.5160 | 2.5405 | 3.1018 | H8 | 2.1421 | 1.1071 | 2.1849 | 3.3212 | 2.4808 | 2.5154 | 1.7786 | | 2.5487 | 3.0994 | 2.5344 | H9 | 3.5007 | 2.1902 | 1.1037 | 3.9161 | 4.3513 | 3.8202 | 2.5160 | 2.5487 | | 1.7896 | 1.7874 | H10 | 2.7816 | 2.1776 | 1.1032 | 2.6002 | 3.8000 | 3.1263 | 2.5405 | 3.0994 | 1.7896 | | 1.7864 | H11 | 2.8224 | 2.1878 | 1.1059 | 3.3345 | 3.8339 | 2.6283 | 3.1018 | 2.5344 | 1.7874 | 1.7864 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.445 |
|
C1 |
C2 |
H7 |
108.438 |
| C1 |
C2 |
H8 |
108.068 |
|
C2 |
C1 |
F4 |
109.843 |
| C2 |
C1 |
H5 |
111.372 |
|
C2 |
C1 |
H6 |
111.010 |
| C2 |
C3 |
H9 |
111.249 |
|
C2 |
C3 |
H10 |
110.284 |
| C2 |
C3 |
H11 |
110.928 |
|
C3 |
C2 |
H7 |
110.118 |
| C3 |
C2 |
H8 |
110.634 |
|
F4 |
C1 |
H5 |
107.985 |
| F4 |
C1 |
H6 |
107.802 |
|
H5 |
C1 |
H6 |
108.712 |
| H7 |
C2 |
H8 |
106.950 |
|
H9 |
C3 |
H10 |
108.362 |
| H9 |
C3 |
H11 |
107.974 |
|
H10 |
C3 |
H11 |
107.928 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
