Jump to
S1C2
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -218.107102 |
| Energy at 298.15K | -218.114836 |
| HF Energy | -218.107102 |
| Nuclear repulsion energy | 127.031452 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3060 |
3042 |
21.58 |
|
|
|
| 2 |
A' |
2975 |
2957 |
34.00 |
|
|
|
| 3 |
A' |
2969 |
2952 |
19.84 |
|
|
|
| 4 |
A' |
2932 |
2915 |
41.53 |
|
|
|
| 5 |
A' |
1443 |
1434 |
3.44 |
|
|
|
| 6 |
A' |
1430 |
1421 |
3.14 |
|
|
|
| 7 |
A' |
1417 |
1409 |
0.40 |
|
|
|
| 8 |
A' |
1370 |
1362 |
14.44 |
|
|
|
| 9 |
A' |
1348 |
1340 |
1.03 |
|
|
|
| 10 |
A' |
1272 |
1264 |
2.29 |
|
|
|
| 11 |
A' |
1104 |
1098 |
1.35 |
|
|
|
| 12 |
A' |
1047 |
1041 |
62.98 |
|
|
|
| 13 |
A' |
1029 |
1023 |
39.82 |
|
|
|
| 14 |
A' |
878 |
873 |
9.03 |
|
|
|
| 15 |
A' |
438 |
436 |
5.70 |
|
|
|
| 16 |
A' |
255 |
254 |
2.47 |
|
|
|
| 17 |
A" |
3046 |
3029 |
50.98 |
|
|
|
| 18 |
A" |
3018 |
3001 |
13.00 |
|
|
|
| 19 |
A" |
2976 |
2958 |
29.70 |
|
|
|
| 20 |
A" |
1423 |
1415 |
6.98 |
|
|
|
| 21 |
A" |
1262 |
1255 |
0.06 |
|
|
|
| 22 |
A" |
1213 |
1206 |
0.35 |
|
|
|
| 23 |
A" |
1140 |
1133 |
0.93 |
|
|
|
| 24 |
A" |
860 |
855 |
1.91 |
|
|
|
| 25 |
A" |
749 |
745 |
2.80 |
|
|
|
| 26 |
A" |
232 |
230 |
0.00 |
|
|
|
| 27 |
A" |
128 |
127 |
2.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20505.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 20386.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
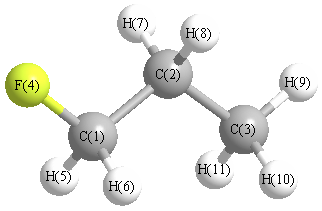
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.123 |
-0.787 |
0.000 |
| C2 |
0.000 |
0.729 |
0.000 |
| C3 |
-1.460 |
1.189 |
0.000 |
| F4 |
1.469 |
-1.169 |
0.000 |
| H5 |
-0.363 |
-1.226 |
0.902 |
| H6 |
-0.363 |
-1.226 |
-0.902 |
| H7 |
0.533 |
1.126 |
-0.891 |
| H8 |
0.533 |
1.126 |
0.891 |
| H9 |
-1.532 |
2.294 |
0.000 |
| H10 |
-2.004 |
0.820 |
-0.895 |
| H11 |
-2.004 |
0.820 |
0.895 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5206 | 2.5314 | 1.3995 | 1.1146 | 1.1146 | 2.1491 | 2.1491 | 3.4973 | 2.8118 | 2.8118 |
C2 | 1.5206 | | 1.5307 | 2.4002 | 2.1834 | 2.1834 | 1.1111 | 1.1111 | 2.1904 | 2.1966 | 2.1966 | C3 | 2.5314 | 1.5307 | | 3.7603 | 2.8015 | 2.8015 | 2.1837 | 2.1837 | 1.1077 | 1.1103 | 1.1103 | F4 | 1.3995 | 2.4002 | 3.7603 | | 2.0429 | 2.0429 | 2.6337 | 2.6337 | 4.5829 | 4.1013 | 4.1013 | H5 | 1.1146 | 2.1834 | 2.8015 | 2.0429 | | 1.8033 | 3.0897 | 2.5167 | 3.8174 | 3.1792 | 2.6229 | H6 | 1.1146 | 2.1834 | 2.8015 | 2.0429 | 1.8033 | | 2.5167 | 3.0897 | 3.8174 | 2.6229 | 3.1792 | H7 | 2.1491 | 1.1111 | 2.1837 | 2.6337 | 3.0897 | 2.5167 | | 1.7814 | 2.5344 | 2.5551 | 3.1172 | H8 | 2.1491 | 1.1111 | 2.1837 | 2.6337 | 2.5167 | 3.0897 | 1.7814 | | 2.5344 | 3.1172 | 2.5551 | H9 | 3.4973 | 2.1904 | 1.1077 | 4.5829 | 3.8174 | 3.8174 | 2.5344 | 2.5344 | | 1.7879 | 1.7879 | H10 | 2.8118 | 2.1966 | 1.1103 | 4.1013 | 3.1792 | 2.6229 | 2.5551 | 3.1172 | 1.7879 | | 1.7900 | H11 | 2.8118 | 2.1966 | 1.1103 | 4.1013 | 2.6229 | 3.1792 | 3.1172 | 2.5551 | 1.7879 | 1.7900 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.121 |
|
C1 |
C2 |
H7 |
108.492 |
| C1 |
C2 |
H8 |
108.492 |
|
C2 |
C1 |
F4 |
110.492 |
| C2 |
C1 |
H5 |
110.963 |
|
C2 |
C1 |
H6 |
110.963 |
| C2 |
C3 |
H9 |
111.224 |
|
C2 |
C3 |
H10 |
111.561 |
| C2 |
C3 |
H11 |
111.561 |
|
C3 |
C2 |
H7 |
110.488 |
| C3 |
C2 |
H8 |
110.488 |
|
F4 |
C1 |
H5 |
108.162 |
| F4 |
C1 |
H6 |
108.162 |
|
H5 |
C1 |
H6 |
107.989 |
| H7 |
C2 |
H8 |
106.570 |
|
H9 |
C3 |
H10 |
107.424 |
| H9 |
C3 |
H11 |
107.424 |
|
H10 |
C3 |
H11 |
107.423 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.183 |
|
|
|
| 2 |
C |
-0.097 |
|
|
|
| 3 |
C |
-0.084 |
|
|
|
| 4 |
F |
-0.223 |
|
|
|
| 5 |
H |
0.021 |
|
|
|
| 6 |
H |
0.021 |
|
|
|
| 7 |
H |
0.034 |
|
|
|
| 8 |
H |
0.034 |
|
|
|
| 9 |
H |
0.038 |
|
|
|
| 10 |
H |
0.037 |
|
|
|
| 11 |
H |
0.037 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.508 |
0.719 |
0.000 |
1.670 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.776 |
1.852 |
0.000 |
| y |
1.852 |
-25.845 |
0.000 |
| z |
0.000 |
0.000 |
-24.768 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.470 |
1.852 |
0.000 |
| y |
1.852 |
0.428 |
0.000 |
| z |
0.000 |
0.000 |
2.042 |
|
| Polar |
|---|
| 3z2-r2 | 4.085 |
|---|
| x2-y2 | -1.932 |
|---|
| xy | 1.852 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.627 |
-0.299 |
0.000 |
| y |
-0.299 |
5.621 |
0.000 |
| z |
0.000 |
0.000 |
5.054 |
<r2> (average value of r
2) Å
2
| <r2> |
104.483 |
| (<r2>)1/2 |
10.222 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -218.107901 |
| Energy at 298.15K | |
| HF Energy | -218.107901 |
| Nuclear repulsion energy | 129.474298 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3070 |
3052 |
16.76 |
|
|
|
| 2 |
A |
3053 |
3035 |
33.50 |
|
|
|
| 3 |
A |
3010 |
2993 |
29.80 |
|
|
|
| 4 |
A |
2992 |
2975 |
49.19 |
|
|
|
| 5 |
A |
2973 |
2956 |
22.78 |
|
|
|
| 6 |
A |
2964 |
2947 |
13.64 |
|
|
|
| 7 |
A |
2939 |
2922 |
45.35 |
|
|
|
| 8 |
A |
1433 |
1425 |
4.47 |
|
|
|
| 9 |
A |
1432 |
1423 |
4.50 |
|
|
|
| 10 |
A |
1414 |
1406 |
5.62 |
|
|
|
| 11 |
A |
1396 |
1388 |
2.50 |
|
|
|
| 12 |
A |
1364 |
1356 |
13.12 |
|
|
|
| 13 |
A |
1344 |
1337 |
4.22 |
|
|
|
| 14 |
A |
1314 |
1307 |
1.25 |
|
|
|
| 15 |
A |
1250 |
1243 |
0.93 |
|
|
|
| 16 |
A |
1217 |
1210 |
0.90 |
|
|
|
| 17 |
A |
1127 |
1120 |
1.13 |
|
|
|
| 18 |
A |
1085 |
1079 |
4.48 |
|
|
|
| 19 |
A |
1063 |
1057 |
46.57 |
|
|
|
| 20 |
A |
953 |
947 |
46.71 |
|
|
|
| 21 |
A |
887 |
881 |
2.81 |
|
|
|
| 22 |
A |
867 |
862 |
4.73 |
|
|
|
| 23 |
A |
745 |
741 |
0.67 |
|
|
|
| 24 |
A |
465 |
462 |
2.91 |
|
|
|
| 25 |
A |
310 |
308 |
1.03 |
|
|
|
| 26 |
A |
217 |
216 |
1.71 |
|
|
|
| 27 |
A |
146 |
146 |
2.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20514.9 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 20395.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.801 |
0.520 |
0.293 |
| C2 |
-0.600 |
0.663 |
-0.279 |
| C3 |
-1.526 |
-0.489 |
0.118 |
| F4 |
1.393 |
-0.667 |
-0.170 |
| H5 |
1.453 |
1.369 |
-0.012 |
| H6 |
0.777 |
0.474 |
1.406 |
| H7 |
-0.522 |
0.728 |
-1.386 |
| H8 |
-1.011 |
1.639 |
0.064 |
| H9 |
-2.522 |
-0.395 |
-0.357 |
| H10 |
-1.085 |
-1.457 |
-0.190 |
| H11 |
-1.677 |
-0.524 |
1.218 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5204 | 2.5423 | 1.4046 | 1.1126 | 1.1145 | 2.1478 | 2.1421 | 3.5076 | 2.7752 | 2.8438 |
C2 | 1.5204 | | 1.5304 | 2.3988 | 2.1877 | 2.1849 | 1.1117 | 1.1126 | 2.1953 | 2.1772 | 2.1934 | C3 | 2.5423 | 1.5304 | | 2.9385 | 3.5132 | 2.8089 | 2.1799 | 2.1894 | 1.1079 | 1.1080 | 1.1105 | F4 | 1.4046 | 2.3988 | 2.9385 | | 2.0426 | 2.0404 | 2.6631 | 3.3392 | 3.9290 | 2.6013 | 3.3720 | H5 | 1.1126 | 2.1877 | 3.5132 | 2.0426 | | 1.8073 | 2.4905 | 2.4804 | 4.3627 | 3.8029 | 3.8590 | H6 | 1.1145 | 2.1849 | 2.8089 | 2.0404 | 1.8073 | | 3.0903 | 2.5213 | 3.8406 | 3.1220 | 2.6561 | H7 | 2.1478 | 1.1117 | 2.1799 | 2.6631 | 2.4905 | 3.0903 | | 1.7810 | 2.5139 | 2.5540 | 3.1119 | H8 | 2.1421 | 1.1126 | 2.1894 | 3.3392 | 2.4804 | 2.5213 | 1.7810 | | 2.5681 | 3.1072 | 2.5402 | H9 | 3.5076 | 2.1953 | 1.1079 | 3.9290 | 4.3627 | 3.8406 | 2.5139 | 2.5681 | | 1.7948 | 1.7924 | H10 | 2.7752 | 2.1772 | 1.1080 | 2.6013 | 3.8029 | 3.1220 | 2.5540 | 3.1072 | 1.7948 | | 1.7899 | H11 | 2.8438 | 2.1934 | 1.1105 | 3.3720 | 3.8590 | 2.6561 | 3.1119 | 2.5402 | 1.7924 | 1.7899 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.883 |
|
C1 |
C2 |
H7 |
108.371 |
| C1 |
C2 |
H8 |
107.885 |
|
C2 |
C1 |
F4 |
110.131 |
| C2 |
C1 |
H5 |
111.437 |
|
C2 |
C1 |
H6 |
111.100 |
| C2 |
C3 |
H9 |
111.623 |
|
C2 |
C3 |
H10 |
110.190 |
| C2 |
C3 |
H11 |
111.313 |
|
C3 |
C2 |
H7 |
110.181 |
| C3 |
C2 |
H8 |
110.873 |
|
F4 |
C1 |
H5 |
107.918 |
| F4 |
C1 |
H6 |
107.634 |
|
H5 |
C1 |
H6 |
108.484 |
| H7 |
C2 |
H8 |
106.395 |
|
H9 |
C3 |
H10 |
108.193 |
| H9 |
C3 |
H11 |
107.798 |
|
H10 |
C3 |
H11 |
107.571 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.185 |
|
|
|
| 2 |
C |
-0.105 |
|
|
|
| 3 |
C |
-0.063 |
|
|
|
| 4 |
F |
-0.224 |
|
|
|
| 5 |
H |
0.021 |
|
|
|
| 6 |
H |
0.021 |
|
|
|
| 7 |
H |
0.035 |
|
|
|
| 8 |
H |
0.023 |
|
|
|
| 9 |
H |
0.031 |
|
|
|
| 10 |
H |
0.044 |
|
|
|
| 11 |
H |
0.032 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.835 |
1.272 |
0.383 |
1.569 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.711 |
1.464 |
0.531 |
| y |
1.464 |
-25.375 |
0.051 |
| z |
0.531 |
0.051 |
-24.886 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.580 |
1.464 |
0.531 |
| y |
1.464 |
0.424 |
0.051 |
| z |
0.531 |
0.051 |
1.156 |
|
| Polar |
|---|
| 3z2-r2 | 2.312 |
|---|
| x2-y2 | -1.336 |
|---|
| xy | 1.464 |
|---|
| xz | 0.531 |
|---|
| yz | 0.051 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.790 |
0.303 |
0.076 |
| y |
0.303 |
5.366 |
-0.045 |
| z |
0.076 |
-0.045 |
5.167 |
<r2> (average value of r
2) Å
2
| <r2> |
92.275 |
| (<r2>)1/2 |
9.606 |
