return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2FCH2CH2F (1,3-difluoropropane)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-316.816484
Energy at 298.15K 
HF Energy-315.981956
Nuclear repulsion energy187.786745
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3156 3019 35.30      
2 A 3098 2964 4.61      
3 A 3081 2947 6.72      
4 A 1519 1454 0.81      
5 A 1466 1403 17.24      
6 A 1456 1392 4.95      
7 A 1322 1265 0.51      
8 A 1256 1202 0.83      
9 A 1153 1103 31.56      
10 A 1041 996 30.24      
11 A 898 859 0.11      
12 A 541 517 3.84      
13 A 248 237 3.55      
14 A 86 82 3.60      
15 B 3158 3021 37.18      
16 B 3137 3001 15.85      
17 B 3093 2960 68.25      
18 B 1523 1457 2.79      
19 B 1444 1381 17.06      
20 B 1402 1341 4.50      
21 B 1276 1221 8.22      
22 B 1141 1092 11.88      
23 B 1138 1089 98.25      
24 B 982 939 17.96      
25 B 791 757 2.69      
26 B 419 401 7.43      
27 B 196 188 10.88      

Unscaled Zero Point Vibrational Energy (zpe) 20009.8 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 19143.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.30196 0.09932 0.09274

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.988
C2 0.000 1.262 0.136
C3 0.000 -1.262 0.136
F4 1.163 1.301 -0.625
F5 -1.163 -1.301 -0.625
H6 0.894 -0.003 1.640
H7 -0.894 0.003 1.640
H8 -0.862 1.263 -0.554
H9 -0.033 2.170 0.763
H10 0.862 -1.263 -0.554
H11 0.033 -2.170 0.763

Atom - Atom Distances (Å)
  C1 C2 C3 F4 F5 H6 H7 H8 H9 H10 H11
C11.52271.52272.37692.37691.10571.10572.17182.18152.17182.1815
C21.52272.52311.39022.91552.15872.15481.10361.10422.75533.4882
C31.52272.52312.91551.39022.15482.15872.75533.48821.10361.1042
F42.37691.39022.91553.49042.62763.32322.02602.02772.58303.9053
F52.37692.91551.39023.49043.32322.62762.58303.90532.02602.0277
H61.10572.15872.15482.62763.32321.78743.08162.51972.52972.4903
H71.10572.15482.15873.32322.62761.78742.52972.49033.08162.5197
H82.17181.10362.75532.02602.58303.08162.52971.80083.05783.7839
H92.18151.10423.48822.02773.90532.51972.49031.80083.78394.3397
H102.17182.75531.10362.58302.02602.52973.08163.05783.78391.8008
H112.18153.48821.10423.90532.02772.49032.51973.78394.33971.8008

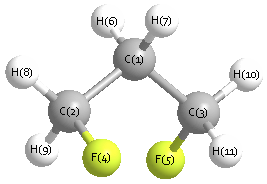
picture of 1,3-difluoropropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 F4 109.286 C1 C2 H8 110.552
C1 C2 H9 111.293 C1 C3 F5 109.286
C1 C3 H10 110.552 C1 C3 H11 111.293
C2 C1 C3 111.890 C2 C1 H6 109.406
C2 C1 H7 109.103 C3 C1 H6 109.103
C3 C1 H7 109.406 F4 C2 H8 108.108
F4 C2 H9 108.212 F5 C3 H10 108.108
F5 C3 H11 108.212 H6 C1 H7 107.845
H8 C2 H9 109.303 H10 C3 H11 109.303
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability