return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4 (Benzyne)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-230.220354
Energy at 298.15K-230.223375
Nuclear repulsion energy185.727654
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3243 3111 5.09      
2 A1 3216 3085 6.35      
3 A1 1960 1880 2.90      
4 A1 1492 1431 0.03      
5 A1 1325 1271 0.24      
6 A1 1159 1112 0.01      
7 A1 1059 1016 21.13      
8 A1 1010 969 4.89      
9 A1 612 587 0.08      
10 A2 958 919 0.00      
11 A2 869 834 0.00      
12 A2 568 545 0.00      
13 A2 418 401 0.00      
14 B1 926 888 0.06      
15 B1 752 722 65.90      
16 B1 388 372 4.66      
17 B2 3240 3108 24.52      
18 B2 3199 3069 1.66      
19 B2 1528 1466 8.36      
20 B2 1433 1375 6.67      
21 B2 1264 1213 0.00      
22 B2 1119 1074 1.03      
23 B2 846 812 27.73      
24 B2 425 408 85.69      

Unscaled Zero Point Vibrational Energy (zpe) 16503.5 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 15833.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.22971 0.18695 0.10307

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.634 -1.253
C2 0.000 -0.634 -1.253
C3 0.000 1.472 -0.128
C4 0.000 -1.472 -0.128
C5 0.000 0.711 1.065
C6 0.000 -0.711 1.065
H7 0.000 2.566 -0.129
H8 0.000 -2.566 -0.129
H9 0.000 1.236 2.028
H10 0.000 -1.236 2.028

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10
C11.26771.40272.38762.31852.67942.23533.39173.33523.7760
C21.26772.38761.40272.67942.31853.39172.23533.77603.3352
C31.40272.38762.94481.41492.48831.09404.03882.16883.4618
C42.38761.40272.94482.48831.41494.03881.09403.46182.1688
C52.31852.67941.41492.48831.42272.20603.48841.09692.1727
C62.67942.31852.48831.41491.42273.48842.20602.17271.0969
H72.23533.39171.09404.03882.20603.48845.13282.53424.3718
H83.39172.23534.03881.09403.48842.20605.13284.37182.5342
H93.33523.77602.16883.46181.09692.17272.53424.37182.4723
H103.77603.33523.46182.16882.17271.09694.37182.53422.4723

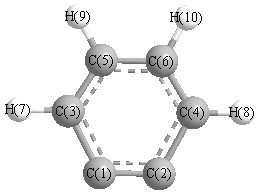
picture of Benzyne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 126.713 C1 C3 C5 110.748
C1 C3 H7 126.655 C2 C1 C3 126.713
C2 C4 C6 110.748 C2 C4 H8 126.655
C3 C5 C6 122.539 C3 C5 H9 118.876
C4 C6 C5 122.539 C4 C6 H10 118.876
C5 C3 H7 122.597 C5 C6 H10 118.585
C6 C4 H8 122.597 C6 C5 H9 118.585
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability