Jump to
S1C2
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -245.100138 |
| Energy at 298.15K | |
| HF Energy | -245.100138 |
| Nuclear repulsion energy | 121.817384 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3641 |
3647 |
47.92 |
|
|
|
| 2 |
A' |
3601 |
3607 |
48.99 |
|
|
|
| 3 |
A' |
3496 |
3501 |
34.60 |
|
|
|
| 4 |
A' |
1778 |
1781 |
401.51 |
|
|
|
| 5 |
A' |
1530 |
1532 |
109.36 |
|
|
|
| 6 |
A' |
1373 |
1375 |
85.49 |
|
|
|
| 7 |
A' |
1190 |
1192 |
191.12 |
|
|
|
| 8 |
A' |
1029 |
1031 |
37.25 |
|
|
|
| 9 |
A' |
902 |
904 |
50.82 |
|
|
|
| 10 |
A' |
556 |
557 |
27.57 |
|
|
|
| 11 |
A' |
468 |
469 |
6.20 |
|
|
|
| 12 |
A" |
738 |
739 |
17.72 |
|
|
|
| 13 |
A" |
578 |
579 |
51.42 |
|
|
|
| 14 |
A" |
461 |
461 |
55.29 |
|
|
|
| 15 |
A" |
279i |
279i |
187.78 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10530.3 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 10547.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.135 |
0.000 |
| O2 |
-0.074 |
1.360 |
0.000 |
| N3 |
1.156 |
-0.604 |
0.000 |
| O4 |
-1.102 |
-0.705 |
0.000 |
| H5 |
2.044 |
-0.108 |
0.000 |
| H6 |
1.137 |
-1.622 |
0.000 |
| H7 |
-1.867 |
-0.088 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2269 | 1.3721 | 1.3859 | 2.0579 | 2.0924 | 1.8800 |
O2 | 1.2269 | | 2.3170 | 2.3069 | 2.5760 | 3.2177 | 2.3046 | N3 | 1.3721 | 2.3170 | | 2.2605 | 1.0168 | 1.0179 | 3.0666 | O4 | 1.3859 | 2.3069 | 2.2605 | | 3.2019 | 2.4191 | 0.9827 | H5 | 2.0579 | 2.5760 | 1.0168 | 3.2019 | | 1.7649 | 3.9104 | H6 | 2.0924 | 3.2177 | 1.0179 | 2.4191 | 1.7649 | | 3.3723 | H7 | 1.8800 | 2.3046 | 3.0666 | 0.9827 | 3.9104 | 3.3723 | |
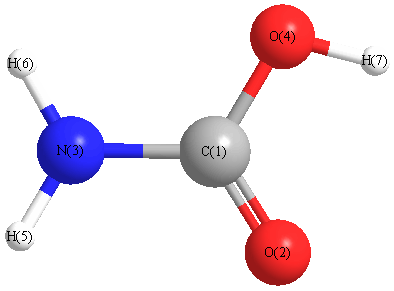 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.199 |
|
C1 |
N3 |
H6 |
121.491 |
| C1 |
O4 |
H7 |
103.768 |
|
O2 |
C1 |
N3 |
126.030 |
| O2 |
C1 |
O4 |
123.879 |
|
N3 |
C1 |
O4 |
110.091 |
| H5 |
N3 |
H6 |
120.310 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.130 |
|
|
|
| 2 |
O |
-0.251 |
|
|
|
| 3 |
N |
-0.046 |
|
|
|
| 4 |
O |
-0.175 |
|
|
|
| 5 |
H |
0.098 |
|
|
|
| 6 |
H |
0.093 |
|
|
|
| 7 |
H |
0.150 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.897 |
-2.198 |
0.000 |
2.374 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.359 |
-2.319 |
0.000 |
| y |
-2.319 |
-25.383 |
0.000 |
| z |
0.000 |
0.000 |
-23.366 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
9.015 |
-2.319 |
0.000 |
| y |
-2.319 |
-6.020 |
0.000 |
| z |
0.000 |
0.000 |
-2.995 |
|
| Polar |
|---|
| 3z2-r2 | -5.990 |
|---|
| x2-y2 | 10.024 |
|---|
| xy | -2.319 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.606 |
-0.338 |
0.000 |
| y |
-0.338 |
4.206 |
0.000 |
| z |
0.000 |
0.000 |
1.821 |
<r2> (average value of r
2) Å
2
| <r2> |
65.497 |
| (<r2>)1/2 |
8.093 |
Jump to
S1C1
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -245.100470 |
| Energy at 298.15K | -245.105210 |
| HF Energy | -245.100470 |
| Nuclear repulsion energy | 121.736353 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3600 |
3606 |
51.31 |
|
|
|
| 2 |
A |
3591 |
3597 |
32.55 |
|
|
|
| 3 |
A |
3462 |
3468 |
22.98 |
|
|
|
| 4 |
A |
1778 |
1781 |
379.25 |
|
|
|
| 5 |
A |
1543 |
1546 |
85.40 |
|
|
|
| 6 |
A |
1372 |
1374 |
93.29 |
|
|
|
| 7 |
A |
1197 |
1199 |
148.85 |
|
|
|
| 8 |
A |
1046 |
1047 |
82.41 |
|
|
|
| 9 |
A |
905 |
907 |
46.65 |
|
|
|
| 10 |
A |
736 |
737 |
29.04 |
|
|
|
| 11 |
A |
566 |
567 |
64.07 |
|
|
|
| 12 |
A |
553 |
554 |
41.89 |
|
|
|
| 13 |
A |
482 |
482 |
15.98 |
|
|
|
| 14 |
A |
438 |
438 |
17.02 |
|
|
|
| 15 |
A |
385 |
385 |
221.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10826.6 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 10844.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.040 |
0.129 |
-0.002 |
| O2 |
-0.469 |
1.277 |
0.008 |
| N3 |
1.290 |
-0.241 |
-0.067 |
| O4 |
-0.847 |
-0.996 |
0.003 |
| H5 |
1.956 |
0.492 |
0.175 |
| H6 |
1.537 |
-1.193 |
0.207 |
| H7 |
-1.760 |
-0.634 |
0.006 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2255 | 1.3816 | 1.3848 | 2.0367 | 2.0684 | 1.8822 |
O2 | 1.2255 | | 2.3242 | 2.3046 | 2.5544 | 3.1883 | 2.3067 | N3 | 1.3816 | 2.3242 | | 2.2672 | 1.0202 | 1.0214 | 3.0762 | O4 | 1.3848 | 2.3046 | 2.2672 | | 3.1787 | 2.4003 | 0.9827 | H5 | 2.0367 | 2.5544 | 1.0202 | 3.1787 | | 1.7377 | 3.8874 | H6 | 2.0684 | 3.1883 | 1.0214 | 2.4003 | 1.7377 | | 3.3502 | H7 | 1.8822 | 2.3067 | 3.0762 | 0.9827 | 3.8874 | 3.3502 | |
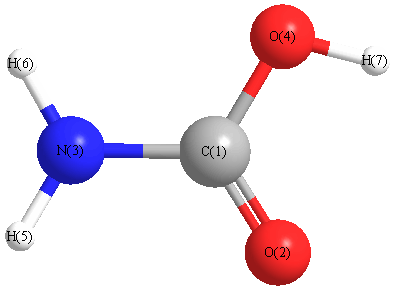 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
115.163 |
|
C1 |
N3 |
H6 |
118.028 |
| C1 |
O4 |
H7 |
104.016 |
|
O2 |
C1 |
N3 |
126.017 |
| O2 |
C1 |
O4 |
123.867 |
|
N3 |
C1 |
O4 |
110.077 |
| H5 |
N3 |
H6 |
116.666 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.127 |
|
|
|
| 2 |
O |
-0.245 |
|
|
|
| 3 |
N |
-0.054 |
|
|
|
| 4 |
O |
-0.172 |
|
|
|
| 5 |
H |
0.099 |
|
|
|
| 6 |
H |
0.095 |
|
|
|
| 7 |
H |
0.150 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.269 |
-1.760 |
0.701 |
2.280 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.437 |
1.017 |
1.381 |
| y |
1.017 |
-25.896 |
-0.356 |
| z |
1.381 |
-0.356 |
-23.246 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
9.134 |
1.017 |
1.381 |
| y |
1.017 |
-6.555 |
-0.356 |
| z |
1.381 |
-0.356 |
-2.579 |
|
| Polar |
|---|
| 3z2-r2 | -5.158 |
|---|
| x2-y2 | 10.459 |
|---|
| xy | 1.017 |
|---|
| xz | 1.381 |
|---|
| yz | -0.356 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.759 |
-0.154 |
0.045 |
| y |
-0.154 |
4.066 |
-0.012 |
| z |
0.045 |
-0.012 |
1.878 |
<r2> (average value of r
2) Å
2
| <r2> |
65.549 |
| (<r2>)1/2 |
8.096 |
