Jump to
S1C2
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -245.066165 |
| Energy at 298.15K | |
| HF Energy | -245.066165 |
| Nuclear repulsion energy | 123.462676 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3790 |
3657 |
86.54 |
|
|
|
| 2 |
A' |
3778 |
3646 |
70.78 |
|
|
|
| 3 |
A' |
3629 |
3502 |
54.84 |
|
|
|
| 4 |
A' |
1874 |
1808 |
484.65 |
|
|
|
| 5 |
A' |
1588 |
1533 |
170.90 |
|
|
|
| 6 |
A' |
1445 |
1394 |
104.30 |
|
|
|
| 7 |
A' |
1235 |
1192 |
193.96 |
|
|
|
| 8 |
A' |
1071 |
1033 |
26.86 |
|
|
|
| 9 |
A' |
971 |
937 |
36.71 |
|
|
|
| 10 |
A' |
585 |
564 |
35.77 |
|
|
|
| 11 |
A' |
485 |
468 |
6.32 |
|
|
|
| 12 |
A" |
787 |
760 |
27.96 |
|
|
|
| 13 |
A" |
594 |
573 |
58.53 |
|
|
|
| 14 |
A" |
485 |
468 |
59.23 |
|
|
|
| 15 |
A" |
244i |
235i |
213.40 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11036.0 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 10649.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.129 |
0.000 |
| O2 |
-0.054 |
1.341 |
0.000 |
| N3 |
1.134 |
-0.615 |
0.000 |
| O4 |
-1.097 |
-0.673 |
0.000 |
| H5 |
2.020 |
-0.133 |
0.000 |
| H6 |
1.100 |
-1.624 |
0.000 |
| H7 |
-1.847 |
-0.057 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2130 | 1.3566 | 1.3591 | 2.0370 | 2.0693 | 1.8561 |
O2 | 1.2130 | | 2.2889 | 2.2678 | 2.5448 | 3.1813 | 2.2729 | N3 | 1.3566 | 2.2889 | | 2.2322 | 1.0084 | 1.0090 | 3.0329 | O4 | 1.3591 | 2.2678 | 2.2322 | | 3.1636 | 2.3941 | 0.9704 | H5 | 2.0370 | 2.5448 | 1.0084 | 3.1636 | | 1.7515 | 3.8676 | H6 | 2.0693 | 3.1813 | 1.0090 | 2.3941 | 1.7515 | | 3.3375 | H7 | 1.8561 | 2.2729 | 3.0329 | 0.9704 | 3.8676 | 3.3375 | |
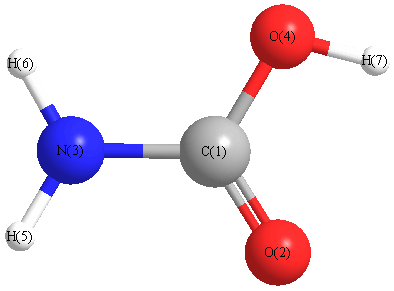 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.177 |
|
C1 |
N3 |
H6 |
121.327 |
| C1 |
O4 |
H7 |
104.415 |
|
O2 |
C1 |
N3 |
125.842 |
| O2 |
C1 |
O4 |
123.595 |
|
N3 |
C1 |
O4 |
110.563 |
| H5 |
N3 |
H6 |
120.496 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.236 |
|
|
|
| 2 |
O |
-0.303 |
|
|
|
| 3 |
N |
-0.140 |
|
|
|
| 4 |
O |
-0.212 |
|
|
|
| 5 |
H |
0.127 |
|
|
|
| 6 |
H |
0.123 |
|
|
|
| 7 |
H |
0.170 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.807 |
-2.315 |
0.000 |
2.451 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.077 |
-2.487 |
0.000 |
| y |
-2.487 |
-25.312 |
0.000 |
| z |
0.000 |
0.000 |
-23.245 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
9.202 |
-2.487 |
0.000 |
| y |
-2.487 |
-6.151 |
0.000 |
| z |
0.000 |
0.000 |
-3.051 |
|
| Polar |
|---|
| 3z2-r2 | -6.102 |
|---|
| x2-y2 | 10.235 |
|---|
| xy | -2.487 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.150 |
-0.320 |
0.000 |
| y |
-0.320 |
3.974 |
0.000 |
| z |
0.000 |
0.000 |
1.803 |
<r2> (average value of r
2) Å
2
| <r2> |
64.022 |
| (<r2>)1/2 |
8.001 |
Jump to
S1C1
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -245.066345 |
| Energy at 298.15K | -245.071145 |
| HF Energy | -245.066345 |
| Nuclear repulsion energy | 123.403895 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3789 |
3657 |
85.62 |
|
|
|
| 2 |
A |
3745 |
3614 |
59.09 |
|
|
|
| 3 |
A |
3608 |
3482 |
43.99 |
|
|
|
| 4 |
A |
1876 |
1810 |
466.15 |
|
|
|
| 5 |
A |
1596 |
1540 |
142.36 |
|
|
|
| 6 |
A |
1445 |
1394 |
119.13 |
|
|
|
| 7 |
A |
1239 |
1195 |
168.84 |
|
|
|
| 8 |
A |
1085 |
1047 |
53.89 |
|
|
|
| 9 |
A |
973 |
939 |
34.96 |
|
|
|
| 10 |
A |
786 |
759 |
38.20 |
|
|
|
| 11 |
A |
590 |
569 |
55.63 |
|
|
|
| 12 |
A |
577 |
557 |
61.90 |
|
|
|
| 13 |
A |
499 |
481 |
19.98 |
|
|
|
| 14 |
A |
462 |
446 |
21.09 |
|
|
|
| 15 |
A |
335 |
324 |
238.73 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11301.3 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 10905.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.041 |
0.123 |
-0.003 |
| O2 |
-0.475 |
1.254 |
0.007 |
| N3 |
1.276 |
-0.226 |
-0.056 |
| O4 |
-0.828 |
-0.985 |
0.003 |
| H5 |
1.938 |
0.510 |
0.147 |
| H6 |
1.539 |
-1.174 |
0.177 |
| H7 |
-1.735 |
-0.640 |
0.007 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2120 | 1.3627 | 1.3590 | 2.0216 | 2.0517 | 1.8583 |
O2 | 1.2120 | | 2.2935 | 2.2665 | 2.5290 | 3.1595 | 2.2748 | N3 | 1.3627 | 2.2935 | | 2.2372 | 1.0106 | 1.0113 | 3.0398 | O4 | 1.3590 | 2.2665 | 2.2372 | | 3.1474 | 2.3810 | 0.9704 | H5 | 2.0216 | 2.5290 | 1.0106 | 3.1474 | | 1.7311 | 3.8514 | H6 | 2.0517 | 3.1595 | 1.0113 | 2.3810 | 1.7311 | | 3.3218 | H7 | 1.8583 | 2.2748 | 3.0398 | 0.9704 | 3.8514 | 3.3218 | |
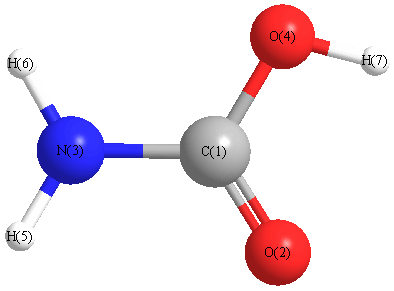 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
116.022 |
|
C1 |
N3 |
H6 |
118.846 |
| C1 |
O4 |
H7 |
104.600 |
|
O2 |
C1 |
N3 |
125.842 |
| O2 |
C1 |
O4 |
123.568 |
|
N3 |
C1 |
O4 |
110.569 |
| H5 |
N3 |
H6 |
117.777 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.232 |
|
|
|
| 2 |
O |
-0.299 |
|
|
|
| 3 |
N |
-0.144 |
|
|
|
| 4 |
O |
-0.210 |
|
|
|
| 5 |
H |
0.127 |
|
|
|
| 6 |
H |
0.123 |
|
|
|
| 7 |
H |
0.170 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.330 |
-1.878 |
0.636 |
2.388 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.989 |
1.172 |
1.244 |
| y |
1.172 |
-25.842 |
-0.321 |
| z |
1.244 |
-0.321 |
-23.150 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
9.507 |
1.172 |
1.244 |
| y |
1.172 |
-6.773 |
-0.321 |
| z |
1.244 |
-0.321 |
-2.735 |
|
| Polar |
|---|
| 3z2-r2 | -5.469 |
|---|
| x2-y2 | 10.853 |
|---|
| xy | 1.172 |
|---|
| xz | 1.244 |
|---|
| yz | -0.321 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.324 |
-0.194 |
0.022 |
| y |
-0.194 |
3.814 |
-0.006 |
| z |
0.022 |
-0.006 |
1.838 |
<r2> (average value of r
2) Å
2
| <r2> |
64.061 |
| (<r2>)1/2 |
8.004 |
