Jump to
S1C2
Energy calculated at QCISD(T)=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -244.554940 |
| Energy at 298.15K | |
| HF Energy | -243.845427 |
| Nuclear repulsion energy | 122.943868 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3815 |
3659 |
|
|
|
|
| 2 |
A' |
3802 |
3647 |
|
|
|
|
| 3 |
A' |
3652 |
3503 |
|
|
|
|
| 4 |
A' |
1875 |
1798 |
|
|
|
|
| 5 |
A' |
1612 |
1546 |
|
|
|
|
| 6 |
A' |
1456 |
1396 |
|
|
|
|
| 7 |
A' |
1256 |
1204 |
|
|
|
|
| 8 |
A' |
1080 |
1035 |
|
|
|
|
| 9 |
A' |
964 |
925 |
|
|
|
|
| 10 |
A' |
585 |
561 |
|
|
|
|
| 11 |
A' |
486 |
466 |
|
|
|
|
| 12 |
A" |
782 |
750 |
|
|
|
|
| 13 |
A" |
593 |
569 |
|
|
|
|
| 14 |
A" |
479 |
459 |
|
|
|
|
| 15 |
A" |
374i |
359i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11031.8 cm
-1
Scaled (by 0.959) Zero Point Vibrational Energy (zpe) 10579.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)=FULL/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.134 |
0.000 |
| O2 |
-0.048 |
1.350 |
0.000 |
| N3 |
1.134 |
-0.625 |
0.000 |
| O4 |
-1.103 |
-0.673 |
0.000 |
| H5 |
2.027 |
-0.152 |
0.000 |
| H6 |
1.086 |
-1.634 |
0.000 |
| H7 |
-1.848 |
-0.051 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2171 | 1.3646 | 1.3668 | 2.0466 | 2.0748 | 1.8569 |
O2 | 1.2171 | | 2.3014 | 2.2820 | 2.5606 | 3.1921 | 2.2810 | N3 | 1.3646 | 2.3014 | | 2.2379 | 1.0100 | 1.0103 | 3.0367 | O4 | 1.3668 | 2.2820 | 2.2379 | | 3.1729 | 2.3909 | 0.9704 | H5 | 2.0466 | 2.5606 | 1.0100 | 3.1729 | | 1.7556 | 3.8756 | H6 | 2.0748 | 3.1921 | 1.0103 | 2.3909 | 1.7556 | | 3.3338 | H7 | 1.8569 | 2.2810 | 3.0367 | 0.9704 | 3.8756 | 3.3338 | |
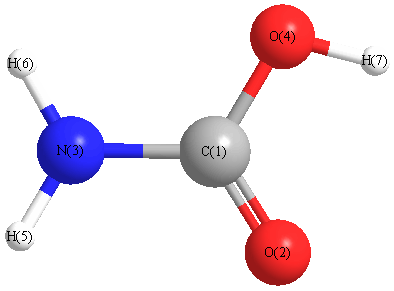 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.282 |
|
C1 |
N3 |
H6 |
121.048 |
| C1 |
O4 |
H7 |
103.923 |
|
O2 |
C1 |
N3 |
126.012 |
| O2 |
C1 |
O4 |
123.948 |
|
N3 |
C1 |
O4 |
110.039 |
| H5 |
N3 |
H6 |
120.670 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -244.555892 |
| Energy at 298.15K | -244.560832 |
| HF Energy | -243.845210 |
| Nuclear repulsion energy | 122.804147 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3810 |
3654 |
|
|
|
|
| 2 |
A |
3726 |
3574 |
|
|
|
|
| 3 |
A |
3599 |
3451 |
|
|
|
|
| 4 |
A |
1876 |
1799 |
|
|
|
|
| 5 |
A |
1625 |
1559 |
|
|
|
|
| 6 |
A |
1454 |
1394 |
|
|
|
|
| 7 |
A |
1266 |
1214 |
|
|
|
|
| 8 |
A |
1111 |
1066 |
|
|
|
|
| 9 |
A |
964 |
925 |
|
|
|
|
| 10 |
A |
785 |
753 |
|
|
|
|
| 11 |
A |
591 |
567 |
|
|
|
|
| 12 |
A |
570 |
547 |
|
|
|
|
| 13 |
A |
512 |
491 |
|
|
|
|
| 14 |
A |
502 |
481 |
|
|
|
|
| 15 |
A |
431 |
414 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11411.4 cm
-1
Scaled (by 0.959) Zero Point Vibrational Energy (zpe) 10943.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)=FULL/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.045 |
0.125 |
-0.004 |
| O2 |
-0.488 |
1.257 |
0.010 |
| N3 |
1.289 |
-0.221 |
-0.082 |
| O4 |
-0.820 |
-0.998 |
0.003 |
| H5 |
1.915 |
0.522 |
0.213 |
| H6 |
1.525 |
-1.145 |
0.267 |
| H7 |
-1.729 |
-0.658 |
0.011 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2154 | 1.3806 | 1.3645 | 2.0115 | 2.0374 | 1.8577 |
O2 | 1.2154 | | 2.3134 | 2.2787 | 2.5213 | 3.1445 | 2.2816 | N3 | 1.3806 | 2.3134 | | 2.2495 | 1.0153 | 1.0157 | 3.0515 | O4 | 1.3645 | 2.2787 | 2.2495 | | 3.1362 | 2.3642 | 0.9708 | H5 | 2.0115 | 2.5213 | 1.0153 | 3.1362 | | 1.7135 | 3.8361 | H6 | 2.0374 | 3.1445 | 1.0157 | 2.3642 | 1.7135 | | 3.3003 | H7 | 1.8577 | 2.2816 | 3.0515 | 0.9708 | 3.8361 | 3.3003 | |
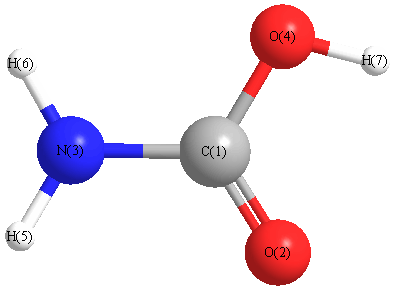 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
113.309 |
|
C1 |
N3 |
H6 |
115.634 |
| C1 |
O4 |
H7 |
104.126 |
|
O2 |
C1 |
N3 |
125.922 |
| O2 |
C1 |
O4 |
123.971 |
|
N3 |
C1 |
O4 |
110.059 |
| H5 |
N3 |
H6 |
115.058 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
