Jump to
S1C2
Energy calculated at MP3=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -244.524982 |
| Energy at 298.15K | |
| HF Energy | -243.847446 |
| Nuclear repulsion energy | 123.681817 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3901 |
3669 |
89.33 |
|
|
|
| 2 |
A' |
3854 |
3624 |
74.44 |
|
|
|
| 3 |
A' |
3708 |
3486 |
66.02 |
|
|
|
| 4 |
A' |
1936 |
1821 |
550.32 |
|
|
|
| 5 |
A' |
1640 |
1542 |
180.17 |
|
|
|
| 6 |
A' |
1489 |
1400 |
112.48 |
|
|
|
| 7 |
A' |
1286 |
1209 |
215.44 |
|
|
|
| 8 |
A' |
1105 |
1039 |
25.66 |
|
|
|
| 9 |
A' |
998 |
938 |
32.03 |
|
|
|
| 10 |
A' |
605 |
569 |
38.94 |
|
|
|
| 11 |
A' |
499 |
469 |
4.97 |
|
|
|
| 12 |
A" |
810 |
762 |
33.00 |
|
|
|
| 13 |
A" |
599 |
564 |
59.61 |
|
|
|
| 14 |
A" |
492 |
463 |
64.90 |
|
|
|
| 15 |
A" |
336i |
316i |
234.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11292.3 cm
-1
Scaled (by 0.9403) Zero Point Vibrational Energy (zpe) 10618.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3=FULL/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.132 |
0.000 |
| O2 |
-0.041 |
1.338 |
0.000 |
| N3 |
1.128 |
-0.627 |
0.000 |
| O4 |
-1.102 |
-0.659 |
0.000 |
| H5 |
2.019 |
-0.158 |
0.000 |
| H6 |
1.076 |
-1.632 |
0.000 |
| H7 |
-1.844 |
-0.042 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2075 | 1.3592 | 1.3565 | 2.0394 | 2.0664 | 1.8522 |
O2 | 1.2075 | | 2.2869 | 2.2620 | 2.5462 | 3.1741 | 2.2707 | N3 | 1.3592 | 2.2869 | | 2.2299 | 1.0068 | 1.0068 | 3.0288 | O4 | 1.3565 | 2.2620 | 2.2299 | | 3.1606 | 2.3853 | 0.9653 | H5 | 2.0394 | 2.5462 | 1.0068 | 3.1606 | | 1.7501 | 3.8644 | H6 | 2.0664 | 3.1741 | 1.0068 | 2.3853 | 1.7501 | | 3.3249 | H7 | 1.8522 | 2.2707 | 3.0288 | 0.9653 | 3.8644 | 3.3249 | |
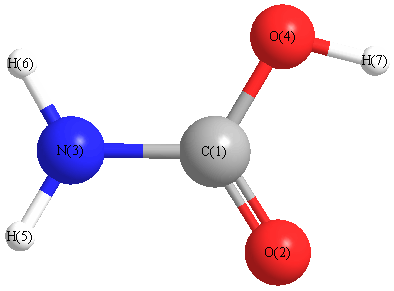 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.310 |
|
C1 |
N3 |
H6 |
120.979 |
| C1 |
O4 |
H7 |
104.571 |
|
O2 |
C1 |
N3 |
125.890 |
| O2 |
C1 |
O4 |
123.719 |
|
N3 |
C1 |
O4 |
110.391 |
| H5 |
N3 |
H6 |
120.711 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP3=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -244.525606 |
| Energy at 298.15K | -244.530572 |
| HF Energy | -243.847486 |
| Nuclear repulsion energy | 123.579674 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3899 |
3666 |
88.68 |
|
|
|
| 2 |
A |
3793 |
3567 |
50.24 |
|
|
|
| 3 |
A |
3666 |
3447 |
41.80 |
|
|
|
| 4 |
A |
1941 |
1825 |
504.21 |
|
|
|
| 5 |
A |
1650 |
1551 |
131.44 |
|
|
|
| 6 |
A |
1488 |
1399 |
131.60 |
|
|
|
| 7 |
A |
1293 |
1216 |
170.91 |
|
|
|
| 8 |
A |
1131 |
1064 |
72.18 |
|
|
|
| 9 |
A |
999 |
940 |
28.03 |
|
|
|
| 10 |
A |
811 |
763 |
57.87 |
|
|
|
| 11 |
A |
609 |
573 |
57.61 |
|
|
|
| 12 |
A |
578 |
544 |
85.02 |
|
|
|
| 13 |
A |
514 |
483 |
13.33 |
|
|
|
| 14 |
A |
481 |
452 |
89.83 |
|
|
|
| 15 |
A |
435 |
409 |
194.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11644.5 cm
-1
Scaled (by 0.9403) Zero Point Vibrational Energy (zpe) 10949.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3=FULL/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.044 |
0.124 |
-0.004 |
| O2 |
-0.475 |
1.250 |
0.009 |
| N3 |
1.280 |
-0.227 |
-0.074 |
| O4 |
-0.826 |
-0.982 |
0.003 |
| H5 |
1.920 |
0.509 |
0.192 |
| H6 |
1.519 |
-1.158 |
0.240 |
| H7 |
-1.730 |
-0.645 |
0.010 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2058 | 1.3720 | 1.3546 | 2.0107 | 2.0364 | 1.8533 |
O2 | 1.2058 | | 2.2959 | 2.2596 | 2.5136 | 3.1352 | 2.2728 | N3 | 1.3720 | 2.2959 | | 2.2386 | 1.0109 | 1.0111 | 3.0405 | O4 | 1.3546 | 2.2596 | 2.2386 | | 3.1300 | 2.3632 | 0.9655 | H5 | 2.0107 | 2.5136 | 1.0109 | 3.1300 | | 1.7154 | 3.8323 | H6 | 2.0364 | 3.1352 | 1.0111 | 2.3632 | 1.7154 | | 3.2976 | H7 | 1.8533 | 2.2728 | 3.0405 | 0.9655 | 3.8323 | 3.2976 | |
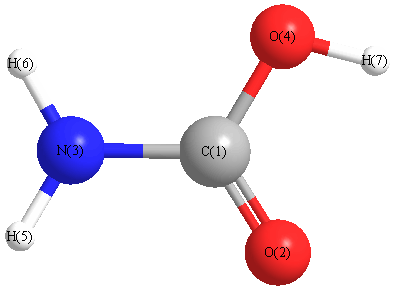 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
114.238 |
|
C1 |
N3 |
H6 |
116.596 |
| C1 |
O4 |
H7 |
104.786 |
|
O2 |
C1 |
N3 |
125.796 |
| O2 |
C1 |
O4 |
123.796 |
|
N3 |
C1 |
O4 |
110.373 |
| H5 |
N3 |
H6 |
116.066 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
