Jump to
S1C2
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -358.355674 |
| Energy at 298.15K | -358.360692 |
| Nuclear repulsion energy | 232.421469 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3739 |
3608 |
89.17 |
|
|
|
| 2 |
A' |
3733 |
3602 |
78.96 |
|
|
|
| 3 |
A' |
3585 |
3460 |
57.89 |
|
|
|
| 4 |
A' |
1851 |
1787 |
56.89 |
|
|
|
| 5 |
A' |
1831 |
1767 |
520.46 |
|
|
|
| 6 |
A' |
1570 |
1516 |
113.58 |
|
|
|
| 7 |
A' |
1435 |
1384 |
39.84 |
|
|
|
| 8 |
A' |
1325 |
1279 |
49.85 |
|
|
|
| 9 |
A' |
1205 |
1163 |
261.46 |
|
|
|
| 10 |
A' |
1089 |
1051 |
2.02 |
|
|
|
| 11 |
A' |
785 |
757 |
9.09 |
|
|
|
| 12 |
A' |
611 |
590 |
69.66 |
|
|
|
| 13 |
A' |
529 |
510 |
0.24 |
|
|
|
| 14 |
A' |
413 |
399 |
4.30 |
|
|
|
| 15 |
A' |
265 |
256 |
14.86 |
|
|
|
| 16 |
A" |
843 |
814 |
5.29 |
|
|
|
| 17 |
A" |
688 |
664 |
119.43 |
|
|
|
| 18 |
A" |
643 |
620 |
17.52 |
|
|
|
| 19 |
A" |
436 |
421 |
4.95 |
|
|
|
| 20 |
A" |
292 |
282 |
196.50 |
|
|
|
| 21 |
A" |
75 |
72 |
1.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13471.1 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 12999.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.751 |
0.000 |
| C2 |
-0.058 |
-0.789 |
0.000 |
| O3 |
-1.099 |
-1.413 |
0.000 |
| O4 |
1.036 |
1.378 |
0.000 |
| O5 |
-1.214 |
1.296 |
0.000 |
| N6 |
1.191 |
-1.305 |
0.000 |
| H7 |
1.316 |
-2.308 |
0.000 |
| H8 |
1.985 |
-0.676 |
0.000 |
| H9 |
-1.074 |
2.260 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5407 | 2.4267 | 1.2114 | 1.3309 | 2.3760 | 3.3301 | 2.4446 | 1.8523 |
C2 | 1.5407 | | 1.2140 | 2.4276 | 2.3839 | 1.3515 | 2.0484 | 2.0458 | 3.2136 | O3 | 2.4267 | 1.2140 | | 3.5143 | 2.7110 | 2.2930 | 2.5758 | 3.1707 | 3.6727 | O4 | 1.2114 | 2.4276 | 3.5143 | | 2.2519 | 2.6878 | 3.6973 | 2.2630 | 2.2871 | O5 | 1.3309 | 2.3839 | 2.7110 | 2.2519 | | 3.5427 | 4.4036 | 3.7581 | 0.9742 | N6 | 2.3760 | 1.3515 | 2.2930 | 2.6878 | 3.5427 | | 1.0110 | 1.0125 | 4.2238 | H7 | 3.3301 | 2.0484 | 2.5758 | 3.6973 | 4.4036 | 1.0110 | | 1.7639 | 5.1557 | H8 | 2.4446 | 2.0458 | 3.1707 | 2.2630 | 3.7581 | 1.0125 | 1.7639 | | 4.2401 | H9 | 1.8523 | 3.2136 | 3.6727 | 2.2871 | 0.9742 | 4.2238 | 5.1557 | 4.2401 | |
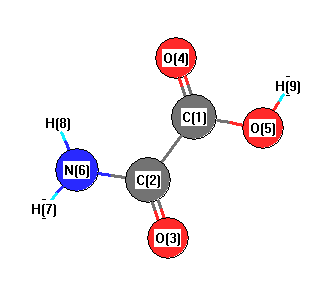 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
123.079 |
|
C1 |
C2 |
N6 |
110.303 |
| C1 |
O5 |
H9 |
105.909 |
|
C2 |
C1 |
O4 |
123.351 |
| C2 |
C1 |
O5 |
112.023 |
|
C2 |
N6 |
H7 |
119.532 |
| C2 |
N6 |
H8 |
119.157 |
|
O3 |
C2 |
N6 |
126.618 |
| O4 |
C1 |
O5 |
124.626 |
|
H7 |
N6 |
H8 |
121.311 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.195 |
|
|
|
| 2 |
C |
0.096 |
|
|
|
| 3 |
O |
-0.236 |
|
|
|
| 4 |
O |
-0.254 |
|
|
|
| 5 |
O |
-0.131 |
|
|
|
| 6 |
N |
-0.092 |
|
|
|
| 7 |
H |
0.125 |
|
|
|
| 8 |
H |
0.129 |
|
|
|
| 9 |
H |
0.168 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.130 |
0.652 |
0.000 |
2.227 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.191 |
7.799 |
0.008 |
| y |
7.799 |
-37.881 |
0.004 |
| z |
0.008 |
0.004 |
-32.791 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
11.146 |
7.799 |
0.008 |
| y |
7.799 |
-9.390 |
0.004 |
| z |
0.008 |
0.004 |
-1.755 |
|
| Polar |
|---|
| 3z2-r2 | -3.510 |
|---|
| x2-y2 | 13.691 |
|---|
| xy | 7.799 |
|---|
| xz | 0.008 |
|---|
| yz | 0.004 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
141.111 |
| (<r2>)1/2 |
11.879 |
Jump to
S1C1
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -358.363180 |
| Energy at 298.15K | -358.368483 |
| Nuclear repulsion energy | 234.144271 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3729 |
3598 |
95.73 |
|
|
|
| 2 |
A' |
3577 |
3452 |
69.65 |
|
|
|
| 3 |
A' |
3498 |
3375 |
177.44 |
|
|
|
| 4 |
A' |
1888 |
1822 |
234.75 |
|
|
|
| 5 |
A' |
1819 |
1756 |
300.77 |
|
|
|
| 6 |
A' |
1580 |
1525 |
51.23 |
|
|
|
| 7 |
A' |
1463 |
1412 |
323.59 |
|
|
|
| 8 |
A' |
1362 |
1314 |
252.11 |
|
|
|
| 9 |
A' |
1229 |
1186 |
3.91 |
|
|
|
| 10 |
A' |
1096 |
1057 |
5.75 |
|
|
|
| 11 |
A' |
815 |
786 |
8.60 |
|
|
|
| 12 |
A' |
634 |
611 |
11.52 |
|
|
|
| 13 |
A' |
548 |
529 |
1.66 |
|
|
|
| 14 |
A' |
400 |
386 |
8.88 |
|
|
|
| 15 |
A' |
266 |
257 |
44.93 |
|
|
|
| 16 |
A" |
840 |
810 |
1.78 |
|
|
|
| 17 |
A" |
796 |
768 |
92.96 |
|
|
|
| 18 |
A" |
672 |
649 |
2.56 |
|
|
|
| 19 |
A" |
467 |
451 |
80.66 |
|
|
|
| 20 |
A" |
359 |
347 |
140.40 |
|
|
|
| 21 |
A" |
128 |
123 |
4.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13581.7 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 13106.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.014 |
-0.796 |
0.000 |
| C2 |
0.000 |
0.744 |
0.000 |
| O3 |
-1.078 |
1.328 |
0.000 |
| O4 |
1.023 |
-1.456 |
0.000 |
| O5 |
-1.223 |
-1.273 |
0.000 |
| N6 |
1.223 |
1.289 |
0.000 |
| H7 |
1.339 |
2.294 |
0.000 |
| H8 |
2.022 |
0.666 |
0.000 |
| H9 |
-1.786 |
-0.462 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5394 | 2.3882 | 1.2060 | 1.3256 | 2.4096 | 3.3616 | 2.4842 | 1.8304 |
C2 | 1.5394 | | 1.2261 | 2.4260 | 2.3585 | 1.3386 | 2.0484 | 2.0239 | 2.1549 | O3 | 2.3882 | 1.2261 | | 3.4880 | 2.6055 | 2.3007 | 2.6026 | 3.1701 | 1.9256 | O4 | 1.2060 | 2.4260 | 3.4880 | | 2.2532 | 2.7521 | 3.7629 | 2.3456 | 2.9796 | O5 | 1.3256 | 2.3585 | 2.6055 | 2.2532 | | 3.5416 | 4.3915 | 3.7804 | 0.9873 | N6 | 2.4096 | 1.3386 | 2.3007 | 2.7521 | 3.5416 | | 1.0115 | 1.0137 | 3.4810 | H7 | 3.3616 | 2.0484 | 2.6026 | 3.7629 | 4.3915 | 1.0115 | | 1.7651 | 4.1667 | H8 | 2.4842 | 2.0239 | 3.1701 | 2.3456 | 3.7804 | 1.0137 | 1.7651 | | 3.9719 | H9 | 1.8304 | 2.1549 | 1.9256 | 2.9796 | 0.9873 | 3.4810 | 4.1667 | 3.9719 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
118.996 |
|
C1 |
C2 |
N6 |
113.517 |
| C1 |
O5 |
H9 |
103.670 |
|
C2 |
C1 |
O4 |
123.713 |
| C2 |
C1 |
O5 |
110.595 |
|
C2 |
N6 |
H7 |
120.654 |
| C2 |
N6 |
H8 |
118.062 |
|
O3 |
C2 |
N6 |
127.487 |
| O4 |
C1 |
O5 |
125.692 |
|
H7 |
N6 |
H8 |
121.284 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.193 |
|
|
|
| 2 |
C |
0.098 |
|
|
|
| 3 |
O |
-0.284 |
|
|
|
| 4 |
O |
-0.236 |
|
|
|
| 5 |
O |
-0.137 |
|
|
|
| 6 |
N |
-0.077 |
|
|
|
| 7 |
H |
0.130 |
|
|
|
| 8 |
H |
0.137 |
|
|
|
| 9 |
H |
0.176 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.600 |
2.635 |
0.000 |
3.082 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.931 |
7.070 |
0.000 |
| y |
7.070 |
-36.382 |
0.000 |
| z |
0.000 |
0.000 |
-32.742 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.632 |
7.070 |
0.000 |
| y |
7.070 |
-5.046 |
0.000 |
| z |
0.000 |
0.000 |
0.414 |
|
| Polar |
|---|
| 3z2-r2 | 0.828 |
|---|
| x2-y2 | 6.452 |
|---|
| xy | 7.070 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.830 |
0.073 |
0.000 |
| y |
0.073 |
5.476 |
0.000 |
| z |
0.000 |
0.000 |
2.572 |
<r2> (average value of r
2) Å
2
| <r2> |
138.492 |
| (<r2>)1/2 |
11.768 |
