Jump to
S1C2
Vibrational Frequencies calculated at CCD/cc-pVDZ
Geometric Data calculated at CCD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -357.561097 |
| Energy at 298.15K | -357.566279 |
| Nuclear repulsion energy | 233.539910 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3793 |
3628 |
91.95 |
|
|
|
| 2 |
A' |
3719 |
3558 |
144.62 |
|
|
|
| 3 |
A' |
3644 |
3486 |
82.65 |
|
|
|
| 4 |
A' |
1939 |
1855 |
195.38 |
|
|
|
| 5 |
A' |
1871 |
1790 |
373.99 |
|
|
|
| 6 |
A' |
1631 |
1561 |
58.43 |
|
|
|
| 7 |
A' |
1502 |
1437 |
246.15 |
|
|
|
| 8 |
A' |
1386 |
1326 |
340.57 |
|
|
|
| 9 |
A' |
1253 |
1199 |
13.16 |
|
|
|
| 10 |
A' |
1119 |
1071 |
2.59 |
|
|
|
| 11 |
A' |
831 |
795 |
9.98 |
|
|
|
| 12 |
A' |
643 |
615 |
15.93 |
|
|
|
| 13 |
A' |
558 |
534 |
2.24 |
|
|
|
| 14 |
A' |
414 |
396 |
6.35 |
|
|
|
| 15 |
A' |
277 |
265 |
39.53 |
|
|
|
| 16 |
A" |
850 |
813 |
0.00 |
|
|
|
| 17 |
A" |
729 |
698 |
107.96 |
|
|
|
| 18 |
A" |
661 |
633 |
6.22 |
|
|
|
| 19 |
A" |
464 |
444 |
39.32 |
|
|
|
| 20 |
A" |
263 |
252 |
206.94 |
|
|
|
| 21 |
A" |
109 |
105 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13828.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 13229.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.006 |
-0.796 |
0.000 |
| C2 |
0.000 |
0.749 |
0.000 |
| O3 |
-1.066 |
1.347 |
0.000 |
| O4 |
1.015 |
-1.452 |
0.000 |
| O5 |
-1.230 |
-1.292 |
0.000 |
| N6 |
1.234 |
1.287 |
0.000 |
| H7 |
1.349 |
2.292 |
0.000 |
| H8 |
2.036 |
0.669 |
0.000 |
| H9 |
-1.812 |
-0.508 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5456 | 2.3958 | 1.2037 | 1.3311 | 2.4179 | 3.3670 | 2.5037 | 1.8400 |
C2 | 1.5456 | | 1.2218 | 2.4241 | 2.3832 | 1.3458 | 2.0486 | 2.0375 | 2.2054 | O3 | 2.3958 | 1.2218 | | 3.4875 | 2.6439 | 2.3007 | 2.5928 | 3.1749 | 1.9992 | O4 | 1.2037 | 2.4241 | 3.4875 | | 2.2502 | 2.7473 | 3.7584 | 2.3540 | 2.9800 | O5 | 1.3311 | 2.3832 | 2.6439 | 2.2502 | | 3.5663 | 4.4147 | 3.8091 | 0.9765 | N6 | 2.4179 | 1.3458 | 2.3007 | 2.7473 | 3.5663 | | 1.0115 | 1.0121 | 3.5352 | H7 | 3.3670 | 2.0486 | 2.5928 | 3.7584 | 4.4147 | 1.0115 | | 1.7621 | 4.2221 | H8 | 2.5037 | 2.0375 | 3.1749 | 2.3540 | 3.8091 | 1.0121 | 1.7621 | | 4.0236 | H9 | 1.8400 | 2.2054 | 1.9992 | 2.9800 | 0.9765 | 3.5352 | 4.2221 | 4.0236 | |
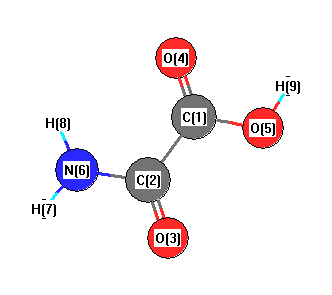 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
119.473 |
|
C1 |
C2 |
N6 |
113.309 |
| C1 |
O5 |
H9 |
104.717 |
|
C2 |
C1 |
O4 |
123.231 |
| C2 |
C1 |
O5 |
111.668 |
|
C2 |
N6 |
H7 |
120.022 |
| C2 |
N6 |
H8 |
118.882 |
|
O3 |
C2 |
N6 |
127.218 |
| O4 |
C1 |
O5 |
125.101 |
|
H7 |
N6 |
H8 |
121.096 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
