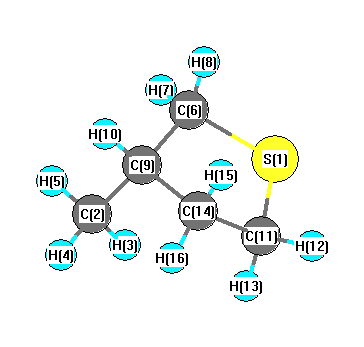
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3056 |
3038 |
18.94 |
|
|
|
| 2 |
A |
3052 |
3034 |
21.00 |
|
|
|
| 3 |
A |
3049 |
3031 |
21.70 |
|
|
|
| 4 |
A |
3042 |
3024 |
10.49 |
|
|
|
| 5 |
A |
3022 |
3004 |
21.01 |
|
|
|
| 6 |
A |
2990 |
2973 |
9.34 |
|
|
|
| 7 |
A |
2986 |
2969 |
61.30 |
|
|
|
| 8 |
A |
2975 |
2958 |
27.42 |
|
|
|
| 9 |
A |
2970 |
2953 |
25.56 |
|
|
|
| 10 |
A |
2961 |
2944 |
6.80 |
|
|
|
| 11 |
A |
1432 |
1423 |
4.45 |
|
|
|
| 12 |
A |
1427 |
1418 |
1.30 |
|
|
|
| 13 |
A |
1412 |
1404 |
5.61 |
|
|
|
| 14 |
A |
1405 |
1397 |
2.08 |
|
|
|
| 15 |
A |
1396 |
1388 |
4.08 |
|
|
|
| 16 |
A |
1345 |
1337 |
9.62 |
|
|
|
| 17 |
A |
1310 |
1302 |
4.22 |
|
|
|
| 18 |
A |
1284 |
1276 |
0.73 |
|
|
|
| 19 |
A |
1256 |
1249 |
2.62 |
|
|
|
| 20 |
A |
1236 |
1229 |
5.11 |
|
|
|
| 21 |
A |
1196 |
1189 |
3.51 |
|
|
|
| 22 |
A |
1177 |
1170 |
12.91 |
|
|
|
| 23 |
A |
1133 |
1126 |
0.72 |
|
|
|
| 24 |
A |
1087 |
1081 |
0.05 |
|
|
|
| 25 |
A |
1038 |
1032 |
0.34 |
|
|
|
| 26 |
A |
1020 |
1014 |
1.15 |
|
|
|
| 27 |
A |
1007 |
1001 |
0.57 |
|
|
|
| 28 |
A |
970 |
965 |
2.00 |
|
|
|
| 29 |
A |
928 |
923 |
0.37 |
|
|
|
| 30 |
A |
908 |
903 |
0.34 |
|
|
|
| 31 |
A |
863 |
858 |
2.94 |
|
|
|
| 32 |
A |
826 |
821 |
0.70 |
|
|
|
| 33 |
A |
788 |
784 |
1.56 |
|
|
|
| 34 |
A |
697 |
693 |
4.74 |
|
|
|
| 35 |
A |
667 |
663 |
2.20 |
|
|
|
| 36 |
A |
589 |
585 |
1.05 |
|
|
|
| 37 |
A |
480 |
477 |
0.19 |
|
|
|
| 38 |
A |
369 |
367 |
0.20 |
|
|
|
| 39 |
A |
343 |
341 |
0.10 |
|
|
|
| 40 |
A |
225 |
224 |
0.12 |
|
|
|
| 41 |
A |
188 |
187 |
0.18 |
|
|
|
| 42 |
A |
110 |
109 |
2.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30105.5 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 29930.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.075 |
|
|
|
| 2 |
C |
-0.007 |
|
|
|
| 3 |
H |
0.042 |
|
|
|
| 4 |
H |
0.028 |
|
|
|
| 5 |
H |
0.032 |
|
|
|
| 6 |
C |
-0.057 |
|
|
|
| 7 |
H |
0.049 |
|
|
|
| 8 |
H |
0.056 |
|
|
|
| 9 |
C |
-0.143 |
|
|
|
| 10 |
H |
0.003 |
|
|
|
| 11 |
C |
-0.110 |
|
|
|
| 12 |
H |
0.057 |
|
|
|
| 13 |
H |
0.060 |
|
|
|
| 14 |
C |
0.017 |
|
|
|
| 15 |
H |
0.028 |
|
|
|
| 16 |
H |
0.020 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.714 |
0.738 |
0.649 |
1.976 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.431 |
-1.748 |
-0.085 |
| y |
-1.748 |
-43.597 |
-1.086 |
| z |
-0.085 |
-1.086 |
-45.935 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.665 |
-1.748 |
-0.085 |
| y |
-1.748 |
3.586 |
-1.086 |
| z |
-0.085 |
-1.086 |
0.079 |
|
| Polar |
|---|
| 3z2-r2 | 0.159 |
|---|
| x2-y2 | -4.834 |
|---|
| xy | -1.748 |
|---|
| xz | -0.085 |
|---|
| yz | -1.086 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.216 |
0.012 |
0.387 |
| y |
0.012 |
10.975 |
-0.289 |
| z |
0.387 |
-0.289 |
9.191 |
<r2> (average value of r
2) Å
2
| <r2> |
196.427 |
| (<r2>)1/2 |
14.015 |