return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CSCH3 (Thioacetone)

using model chemistry: QCISD(T)/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD(T)/cc-pVDZ
 hartrees
Energy at 0K-515.246169
Energy at 298.15K-515.251953
HF Energy-514.620089
Nuclear repulsion energy151.072425
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3172 3023        
2 A1 3034 2892        
3 A1 1479 1410        
4 A1 1389 1324        
5 A1 1305 1243        
6 A1 1016 968        
7 A1 703 670        
8 A1 366 349        
9 A2 3098 2953        
10 A2 1458 1390        
11 A2 914 871        
12 A2 88 84        
13 B1 3104 2958        
14 B1 1480 1410        
15 B1 1062 1012        
16 B1 427 407        
17 B1 147 140        
18 B2 3170 3022        
19 B2 3028 2886        
20 B2 1452 1384        
21 B2 1383 1318        
22 B2 1224 1167        
23 B2 931 887        
24 B2 379 361        

Unscaled Zero Point Vibrational Energy (zpe) 17902.0 cm-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 17062.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/cc-pVDZ
ABC
0.28732 0.15971 0.10681

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 -0.257
S2 0.000 0.000 1.392
C3 0.000 1.278 -1.077
C4 0.000 -1.278 -1.077
H5 0.000 2.171 -0.433
H6 0.000 -2.171 -0.433
H7 0.891 1.299 -1.737
H8 -0.891 1.299 -1.737
H9 -0.891 -1.299 -1.737
H10 0.891 -1.299 -1.737

Atom - Atom Distances (Å)
  C1 S2 C3 C4 H5 H6 H7 H8 H9 H10
C11.64911.51831.51832.17832.17832.16152.16152.16152.1615
S21.64912.78032.78032.83642.83643.50353.50353.50353.5035
C31.51832.78032.55521.10163.50841.10891.10892.80472.8047
C41.51832.78032.55523.50841.10162.80472.80471.10891.1089
H52.17832.83641.10163.50844.34241.80451.80453.81253.8125
H62.17832.83643.50841.10164.34243.81253.81251.80451.8045
H72.16153.50351.10892.80471.80453.81251.78103.14942.5974
H82.16153.50351.10892.80471.80453.81251.78102.59743.1494
H92.16153.50352.80471.10893.81251.80453.14942.59741.7810
H102.16153.50352.80471.10893.81251.80452.59743.14941.7810

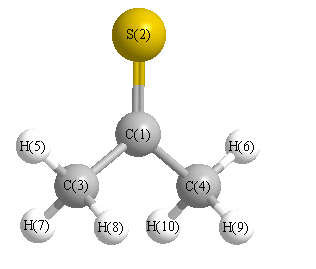
picture of Thioacetone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 H5 111.509 C1 C3 H7 109.743
C1 C3 H8 109.743 C1 C4 H6 111.509
C1 C4 H9 109.743 C1 C4 H10 109.743
S2 C1 C3 122.703 S2 C1 C4 122.703
C3 C1 C4 114.594 H5 C3 H7 109.440
H5 C3 H8 109.440 H6 C4 H9 109.440
H6 C4 H10 109.440 H7 C3 H8 106.853
H9 C4 H10 106.853
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability