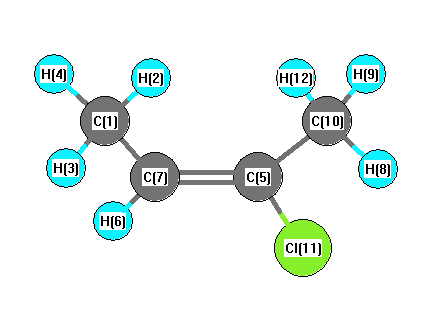
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3106 |
3088 |
7.82 |
|
|
|
| 2 |
A' |
3076 |
3058 |
12.91 |
|
|
|
| 3 |
A' |
3067 |
3049 |
12.11 |
|
|
|
| 4 |
A' |
2972 |
2955 |
14.58 |
|
|
|
| 5 |
A' |
2960 |
2943 |
31.02 |
|
|
|
| 6 |
A' |
1692 |
1682 |
13.74 |
|
|
|
| 7 |
A' |
1413 |
1405 |
6.88 |
|
|
|
| 8 |
A' |
1402 |
1394 |
3.57 |
|
|
|
| 9 |
A' |
1348 |
1340 |
8.96 |
|
|
|
| 10 |
A' |
1344 |
1336 |
0.36 |
|
|
|
| 11 |
A' |
1285 |
1278 |
1.48 |
|
|
|
| 12 |
A' |
1119 |
1112 |
21.35 |
|
|
|
| 13 |
A' |
1045 |
1039 |
53.73 |
|
|
|
| 14 |
A' |
979 |
974 |
11.71 |
|
|
|
| 15 |
A' |
894 |
889 |
14.25 |
|
|
|
| 16 |
A' |
662 |
658 |
20.89 |
|
|
|
| 17 |
A' |
428 |
426 |
9.27 |
|
|
|
| 18 |
A' |
337 |
336 |
0.58 |
|
|
|
| 19 |
A' |
260 |
259 |
0.34 |
|
|
|
| 20 |
A" |
3040 |
3022 |
7.16 |
|
|
|
| 21 |
A" |
3017 |
2999 |
13.12 |
|
|
|
| 22 |
A" |
1400 |
1392 |
8.47 |
|
|
|
| 23 |
A" |
1389 |
1381 |
5.86 |
|
|
|
| 24 |
A" |
1012 |
1007 |
0.02 |
|
|
|
| 25 |
A" |
1003 |
997 |
0.01 |
|
|
|
| 26 |
A" |
817 |
812 |
16.41 |
|
|
|
| 27 |
A" |
431 |
428 |
3.73 |
|
|
|
| 28 |
A" |
227 |
225 |
0.40 |
|
|
|
| 29 |
A" |
176 |
175 |
0.38 |
|
|
|
| 30 |
A" |
141 |
140 |
0.91 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21020.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 20898.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.019 |
|
|
|
| 2 |
H |
0.032 |
|
|
|
| 3 |
H |
0.054 |
|
|
|
| 4 |
H |
0.054 |
|
|
|
| 5 |
C |
-0.235 |
|
|
|
| 6 |
H |
0.002 |
|
|
|
| 7 |
C |
0.009 |
|
|
|
| 8 |
H |
0.061 |
|
|
|
| 9 |
H |
0.061 |
|
|
|
| 10 |
C |
0.034 |
|
|
|
| 11 |
Cl |
-0.093 |
|
|
|
| 12 |
H |
0.039 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.971 |
0.211 |
0.000 |
1.982 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.069 |
0.414 |
0.000 |
| y |
0.414 |
-35.443 |
0.000 |
| z |
0.000 |
0.000 |
-39.225 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.735 |
0.414 |
0.000 |
| y |
0.414 |
3.204 |
0.000 |
| z |
0.000 |
0.000 |
-2.470 |
|
| Polar |
|---|
| 3z2-r2 | -4.939 |
|---|
| x2-y2 | -2.626 |
|---|
| xy | 0.414 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.941 |
1.478 |
0.000 |
| y |
1.478 |
9.362 |
0.000 |
| z |
0.000 |
0.000 |
5.259 |
<r2> (average value of r
2) Å
2
| <r2> |
186.080 |
| (<r2>)1/2 |
13.641 |