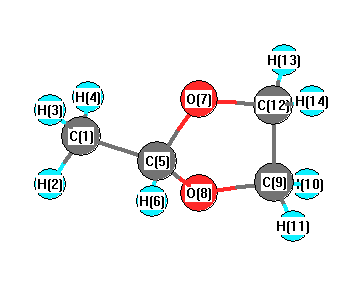
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3090 |
3072 |
11.59 |
|
|
|
| 2 |
A |
3086 |
3068 |
11.51 |
|
|
|
| 3 |
A |
3048 |
3031 |
26.17 |
|
|
|
| 4 |
A |
2992 |
2974 |
10.46 |
|
|
|
| 5 |
A |
2986 |
2968 |
50.90 |
|
|
|
| 6 |
A |
2943 |
2926 |
69.62 |
|
|
|
| 7 |
A |
2904 |
2887 |
62.43 |
|
|
|
| 8 |
A |
2822 |
2806 |
114.17 |
|
|
|
| 9 |
A |
1451 |
1443 |
0.40 |
|
|
|
| 10 |
A |
1431 |
1422 |
6.22 |
|
|
|
| 11 |
A |
1407 |
1399 |
7.40 |
|
|
|
| 12 |
A |
1405 |
1397 |
5.29 |
|
|
|
| 13 |
A |
1373 |
1365 |
68.68 |
|
|
|
| 14 |
A |
1325 |
1317 |
9.41 |
|
|
|
| 15 |
A |
1320 |
1313 |
3.68 |
|
|
|
| 16 |
A |
1314 |
1307 |
4.79 |
|
|
|
| 17 |
A |
1267 |
1260 |
0.53 |
|
|
|
| 18 |
A |
1198 |
1191 |
19.76 |
|
|
|
| 19 |
A |
1170 |
1163 |
0.81 |
|
|
|
| 20 |
A |
1139 |
1133 |
85.87 |
|
|
|
| 21 |
A |
1112 |
1105 |
32.89 |
|
|
|
| 22 |
A |
1086 |
1080 |
59.52 |
|
|
|
| 23 |
A |
1061 |
1055 |
53.52 |
|
|
|
| 24 |
A |
1034 |
1028 |
37.91 |
|
|
|
| 25 |
A |
1013 |
1007 |
33.59 |
|
|
|
| 26 |
A |
929 |
924 |
10.37 |
|
|
|
| 27 |
A |
857 |
852 |
4.12 |
|
|
|
| 28 |
A |
843 |
838 |
35.29 |
|
|
|
| 29 |
A |
818 |
813 |
19.43 |
|
|
|
| 30 |
A |
685 |
681 |
1.14 |
|
|
|
| 31 |
A |
653 |
650 |
1.82 |
|
|
|
| 32 |
A |
489 |
486 |
3.87 |
|
|
|
| 33 |
A |
319 |
317 |
4.94 |
|
|
|
| 34 |
A |
220 |
218 |
1.49 |
|
|
|
| 35 |
A |
205 |
204 |
2.12 |
|
|
|
| 36 |
A |
53 |
53 |
2.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25524.1 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 25376.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.007 |
|
|
|
| 2 |
H |
0.040 |
|
|
|
| 3 |
H |
0.044 |
|
|
|
| 4 |
H |
0.038 |
|
|
|
| 5 |
C |
0.161 |
|
|
|
| 6 |
H |
0.000 |
|
|
|
| 7 |
O |
-0.296 |
|
|
|
| 8 |
O |
-0.268 |
|
|
|
| 9 |
C |
0.065 |
|
|
|
| 10 |
H |
0.033 |
|
|
|
| 11 |
H |
0.022 |
|
|
|
| 12 |
C |
0.112 |
|
|
|
| 13 |
H |
0.031 |
|
|
|
| 14 |
H |
0.024 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.894 |
-0.257 |
0.307 |
0.979 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.709 |
-0.198 |
-0.700 |
| y |
-0.198 |
-40.277 |
-0.537 |
| z |
-0.700 |
-0.537 |
-36.123 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.491 |
-0.198 |
-0.700 |
| y |
-0.198 |
-6.361 |
-0.537 |
| z |
-0.700 |
-0.537 |
-0.130 |
|
| Polar |
|---|
| 3z2-r2 | -0.260 |
|---|
| x2-y2 | 8.568 |
|---|
| xy | -0.198 |
|---|
| xz | -0.700 |
|---|
| yz | -0.537 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.356 |
0.050 |
-0.117 |
| y |
0.050 |
6.820 |
-0.158 |
| z |
-0.117 |
-0.158 |
6.973 |
<r2> (average value of r
2) Å
2
| <r2> |
151.334 |
| (<r2>)1/2 |
12.302 |