return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C7H10 (Norbornene)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-271.906428
Energy at 298.15K-271.918665
HF Energy-270.879322
Nuclear repulsion energy306.255494
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3252 3111 15.22      
2 A' 3156 3019 25.48      
3 A' 3151 3015 48.62      
4 A' 3148 3012 11.12      
5 A' 3094 2960 47.59      
6 A' 3083 2949 34.85      
7 A' 1665 1593 0.84      
8 A' 1515 1449 0.77      
9 A' 1493 1429 2.43      
10 A' 1347 1289 0.33      
11 A' 1334 1276 0.83      
12 A' 1218 1165 0.37      
13 A' 1155 1105 3.44      
14 A' 1116 1067 0.74      
15 A' 1053 1008 1.12      
16 A' 990 947 0.42      
17 A' 981 939 0.11      
18 A' 937 897 0.61      
19 A' 906 867 4.26      
20 A' 825 789 0.27      
21 A' 791 757 4.70      
22 A' 729 697 35.77      
23 A' 478 458 0.75      
24 A' 383 366 2.10      
25 A" 3225 3085 7.02      
26 A" 3146 3010 65.78      
27 A" 3134 2998 2.31      
28 A" 3080 2947 25.67      
29 A" 1487 1423 0.47      
30 A" 1386 1326 5.84      
31 A" 1327 1269 1.74      
32 A" 1295 1239 0.09      
33 A" 1290 1234 0.40      
34 A" 1253 1198 0.19      
35 A" 1218 1165 0.10      
36 A" 1146 1096 0.16      
37 A" 1058 1012 0.45      
38 A" 975 933 0.01      
39 A" 961 919 0.00      
40 A" 934 893 3.14      
41 A" 857 820 2.63      
42 A" 816 780 1.21      
43 A" 675 646 0.36      
44 A" 481 461 0.08      
45 A" 251 240 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 33896.2 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 32428.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.13010 0.11342 0.09946

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.132 0.795 0.000
C2 0.225 0.248 1.131
C3 0.225 0.248 -1.131
C4 -1.131 0.790 0.676
C5 -1.131 0.790 -0.676
C6 0.225 -1.278 0.782
C7 0.225 -1.278 -0.782
H8 1.202 1.895 0.000
H9 2.144 0.350 0.000
H10 0.515 0.480 2.168
H11 0.515 0.480 -2.168
H12 -1.974 1.013 1.338
H13 -1.974 1.013 -1.338
H14 1.134 -1.764 1.180
H15 1.134 -1.764 -1.180
H16 -0.650 -1.796 1.211
H17 -0.650 -1.796 -1.211

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17
C11.54951.54952.36192.36192.39362.39361.10291.10572.27542.27543.38883.38882.81742.81743.36923.3692
C21.54952.26281.52942.32341.56572.44782.22422.23021.10103.31992.33713.39372.20883.19652.22523.2299
C31.54952.26282.32341.52942.44781.56572.22422.23023.31991.10103.39372.33713.19652.20883.22992.2252
C42.36191.52942.32341.35202.47472.87052.66913.37322.24343.30061.09472.19473.45033.88522.68413.2372
C52.36192.32341.52941.35202.87052.47472.66913.37323.30062.24342.19471.09473.88523.45033.23722.6841
C62.39361.56572.44782.47472.87051.56463.41102.63512.25713.44643.22373.81831.10482.21641.10312.2375
C72.39362.44781.56572.87052.47471.56463.41102.63513.44642.25713.81833.22372.21641.10482.23751.1031
H81.10292.22422.22422.66912.66913.41103.41101.81012.67812.67813.55743.55743.84523.84524.30364.3036
H91.10572.23022.23023.37323.37322.63512.63511.81012.71442.71444.38054.38052.62282.62283.72483.7248
H102.27541.10103.31992.24343.30062.25713.44642.67812.71444.33532.67774.33272.52874.07732.73034.2374
H112.27543.31991.10103.30062.24343.44642.25712.67812.71444.33534.33272.67774.07732.52874.23742.7303
H123.38882.33713.39371.09472.19473.22373.81833.55744.38052.67774.33272.67624.17094.86943.10864.0182
H133.38883.39372.33712.19471.09473.81833.22373.55744.38054.33272.67772.67624.86944.17094.01823.1086
H142.81742.20883.19653.45033.88521.10482.21643.84522.62282.52874.07734.17094.86942.35961.78382.9828
H152.81743.19652.20883.88523.45032.21641.10483.84522.62284.07732.52874.86944.17092.35962.98281.7838
H163.36922.22523.22992.68413.23721.10312.23754.30363.72482.73034.23743.10864.01821.78382.98282.4221
H173.36923.22992.22523.23722.68412.23751.10314.30363.72484.23742.73034.01823.10862.98281.78382.4221

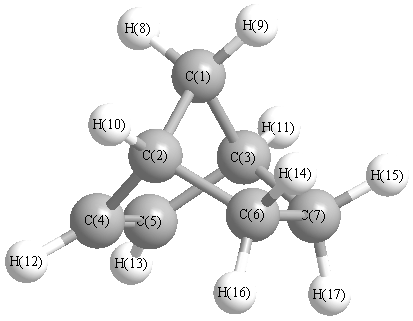
picture of Norbornene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 100.192 C1 C2 C6 100.413
C1 C2 H10 117.287 C1 C3 C5 100.192
C1 C3 C7 100.413 C1 C3 H11 117.287
C2 C1 C3 93.800 C2 C1 H8 112.893
C2 C1 H9 113.206 C2 C4 C5 107.323
C2 C4 H12 125.080 C2 C6 C7 102.883
C2 C6 H14 110.409 C2 C6 H16 111.804
C3 C1 H8 112.893 C3 C1 H9 113.206
C3 C5 C4 107.323 C3 C5 H13 125.080
C3 C7 C6 102.883 C3 C7 H15 110.409
C3 C7 H17 111.804 C4 C2 C6 106.170
C4 C2 H10 116.104 C4 C5 H13 127.213
C5 C3 C7 106.170 C5 C3 H11 116.104
C5 C4 H12 127.213 C6 C2 H10 114.522
C6 C7 H15 111.086 C6 C7 H17 112.873
C7 C3 H11 114.522 C7 C6 H14 111.086
C7 C6 H16 112.873 H8 C1 H9 110.078
H14 C6 H16 107.786 H15 C7 H17 107.786
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability