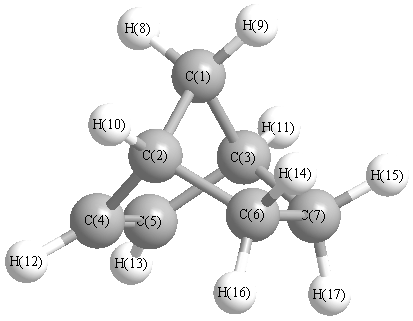
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3150 |
3132 |
17.40 |
|
|
|
| 2 |
A' |
3050 |
3032 |
19.55 |
|
|
|
| 3 |
A' |
3048 |
3030 |
50.78 |
|
|
|
| 4 |
A' |
3044 |
3027 |
9.47 |
|
|
|
| 5 |
A' |
2992 |
2975 |
47.09 |
|
|
|
| 6 |
A' |
2977 |
2959 |
37.37 |
|
|
|
| 7 |
A' |
1598 |
1589 |
1.76 |
|
|
|
| 8 |
A' |
1432 |
1424 |
0.73 |
|
|
|
| 9 |
A' |
1406 |
1398 |
3.69 |
|
|
|
| 10 |
A' |
1272 |
1265 |
0.77 |
|
|
|
| 11 |
A' |
1260 |
1253 |
0.60 |
|
|
|
| 12 |
A' |
1139 |
1132 |
0.12 |
|
|
|
| 13 |
A' |
1103 |
1096 |
5.02 |
|
|
|
| 14 |
A' |
1066 |
1060 |
0.97 |
|
|
|
| 15 |
A' |
1001 |
995 |
1.41 |
|
|
|
| 16 |
A' |
956 |
950 |
0.16 |
|
|
|
| 17 |
A' |
935 |
929 |
0.48 |
|
|
|
| 18 |
A' |
898 |
893 |
0.56 |
|
|
|
| 19 |
A' |
870 |
865 |
5.49 |
|
|
|
| 20 |
A' |
791 |
786 |
0.80 |
|
|
|
| 21 |
A' |
758 |
753 |
3.84 |
|
|
|
| 22 |
A' |
698 |
694 |
34.26 |
|
|
|
| 23 |
A' |
463 |
460 |
0.87 |
|
|
|
| 24 |
A' |
366 |
364 |
1.91 |
|
|
|
| 25 |
A" |
3124 |
3106 |
8.56 |
|
|
|
| 26 |
A" |
3043 |
3026 |
67.51 |
|
|
|
| 27 |
A" |
3031 |
3014 |
2.69 |
|
|
|
| 28 |
A" |
2980 |
2963 |
25.26 |
|
|
|
| 29 |
A" |
1406 |
1398 |
0.58 |
|
|
|
| 30 |
A" |
1314 |
1306 |
6.37 |
|
|
|
| 31 |
A" |
1249 |
1241 |
1.30 |
|
|
|
| 32 |
A" |
1226 |
1219 |
0.00 |
|
|
|
| 33 |
A" |
1220 |
1213 |
0.61 |
|
|
|
| 34 |
A" |
1184 |
1177 |
0.30 |
|
|
|
| 35 |
A" |
1146 |
1139 |
0.24 |
|
|
|
| 36 |
A" |
1093 |
1086 |
0.18 |
|
|
|
| 37 |
A" |
1010 |
1004 |
0.62 |
|
|
|
| 38 |
A" |
936 |
931 |
0.04 |
|
|
|
| 39 |
A" |
919 |
914 |
0.00 |
|
|
|
| 40 |
A" |
893 |
888 |
4.84 |
|
|
|
| 41 |
A" |
828 |
823 |
3.33 |
|
|
|
| 42 |
A" |
781 |
776 |
1.60 |
|
|
|
| 43 |
A" |
654 |
650 |
0.66 |
|
|
|
| 44 |
A" |
471 |
468 |
0.20 |
|
|
|
| 45 |
A" |
245 |
243 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32511.0 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 32322.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.092 |
|
|
|
| 2 |
C |
-0.094 |
|
|
|
| 3 |
C |
-0.094 |
|
|
|
| 4 |
C |
0.036 |
|
|
|
| 5 |
C |
0.036 |
|
|
|
| 6 |
C |
0.035 |
|
|
|
| 7 |
C |
0.035 |
|
|
|
| 8 |
H |
0.014 |
|
|
|
| 9 |
H |
0.004 |
|
|
|
| 10 |
H |
-0.035 |
|
|
|
| 11 |
H |
-0.035 |
|
|
|
| 12 |
H |
-0.028 |
|
|
|
| 13 |
H |
-0.028 |
|
|
|
| 14 |
H |
0.017 |
|
|
|
| 15 |
H |
0.017 |
|
|
|
| 16 |
H |
0.014 |
|
|
|
| 17 |
H |
0.014 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.150 |
-0.287 |
0.000 |
0.324 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.721 |
-0.316 |
0.000 |
| y |
-0.316 |
-45.078 |
0.000 |
| z |
0.000 |
0.000 |
-42.431 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.034 |
-0.316 |
0.000 |
| y |
-0.316 |
-2.503 |
0.000 |
| z |
0.000 |
0.000 |
1.469 |
|
| Polar |
|---|
| 3z2-r2 | 2.937 |
|---|
| x2-y2 | 2.358 |
|---|
| xy | -0.316 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.517 |
-0.713 |
0.000 |
| y |
-0.713 |
9.945 |
0.000 |
| z |
0.000 |
0.000 |
11.291 |
<r2> (average value of r
2) Å
2
| <r2> |
166.477 |
| (<r2>)1/2 |
12.903 |