Jump to
S1C2
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -192.584856 |
| Energy at 298.15K | |
| HF Energy | -191.919875 |
| Nuclear repulsion energy | 127.267061 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3090 |
2945 |
|
|
|
|
| 2 |
A1 |
3036 |
2894 |
|
|
|
|
| 3 |
A1 |
1537 |
1465 |
|
|
|
|
| 4 |
A1 |
1477 |
1408 |
|
|
|
|
| 5 |
A1 |
1382 |
1317 |
|
|
|
|
| 6 |
A1 |
1054 |
1005 |
|
|
|
|
| 7 |
A1 |
938 |
894 |
|
|
|
|
| 8 |
A1 |
823 |
785 |
|
|
|
|
| 9 |
A2 |
3079 |
2934 |
|
|
|
|
| 10 |
A2 |
1225 |
1168 |
|
|
|
|
| 11 |
A2 |
1164 |
1109 |
|
|
|
|
| 12 |
A2 |
832 |
793 |
|
|
|
|
| 13 |
B1 |
3153 |
3005 |
|
|
|
|
| 14 |
B1 |
3077 |
2933 |
|
|
|
|
| 15 |
B1 |
1186 |
1130 |
|
|
|
|
| 16 |
B1 |
1143 |
1090 |
|
|
|
|
| 17 |
B1 |
755 |
720 |
|
|
|
|
| 18 |
B1 |
37i |
35i |
|
|
|
|
| 19 |
B2 |
3026 |
2885 |
|
|
|
|
| 20 |
B2 |
1507 |
1436 |
|
|
|
|
| 21 |
B2 |
1307 |
1245 |
|
|
|
|
| 22 |
B2 |
1256 |
1197 |
|
|
|
|
| 23 |
B2 |
1052 |
1002 |
|
|
|
|
| 24 |
B2 |
951 |
906 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19006.5 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 18115.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.076 |
| C2 |
0.000 |
0.000 |
-1.083 |
| C3 |
0.000 |
1.041 |
0.067 |
| C4 |
0.000 |
-1.041 |
0.067 |
| H5 |
0.901 |
0.000 |
-1.719 |
| H6 |
-0.901 |
0.000 |
-1.719 |
| H7 |
0.902 |
1.682 |
0.132 |
| H8 |
-0.902 |
1.682 |
0.132 |
| H9 |
-0.902 |
-1.682 |
0.132 |
| H10 |
0.902 |
-1.682 |
0.132 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1587 | 1.4495 | 1.4495 | 2.9370 | 2.9370 | 2.1293 | 2.1293 | 2.1293 | 2.1293 |
C2 | 2.1587 | | 1.5505 | 1.5505 | 1.1034 | 1.1034 | 2.2620 | 2.2620 | 2.2620 | 2.2620 | C3 | 1.4495 | 1.5505 | | 2.0810 | 2.2550 | 2.2550 | 1.1085 | 1.1085 | 2.8686 | 2.8686 | C4 | 1.4495 | 1.5505 | 2.0810 | | 2.2550 | 2.2550 | 2.8686 | 2.8686 | 1.1085 | 1.1085 | H5 | 2.9370 | 1.1034 | 2.2550 | 2.2550 | | 1.8025 | 2.5009 | 3.0831 | 3.0831 | 2.5009 | H6 | 2.9370 | 1.1034 | 2.2550 | 2.2550 | 1.8025 | | 3.0831 | 2.5009 | 2.5009 | 3.0831 | H7 | 2.1293 | 2.2620 | 1.1085 | 2.8686 | 2.5009 | 3.0831 | | 1.8037 | 3.8168 | 3.3638 | H8 | 2.1293 | 2.2620 | 1.1085 | 2.8686 | 3.0831 | 2.5009 | 1.8037 | | 3.3638 | 3.8168 | H9 | 2.1293 | 2.2620 | 2.8686 | 1.1085 | 3.0831 | 2.5009 | 3.8168 | 3.3638 | | 1.8037 | H10 | 2.1293 | 2.2620 | 2.8686 | 1.1085 | 2.5009 | 3.0831 | 3.3638 | 3.8168 | 1.8037 | |
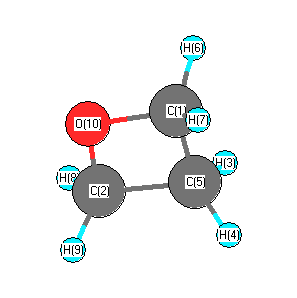 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
91.974 |
|
O1 |
C3 |
H7 |
112.004 |
| O1 |
C3 |
H8 |
112.004 |
|
O1 |
C4 |
C2 |
91.974 |
| O1 |
C4 |
H9 |
112.004 |
|
O1 |
C4 |
H10 |
112.004 |
| C2 |
C3 |
H7 |
115.566 |
|
C2 |
C3 |
H8 |
115.566 |
| C2 |
C4 |
H9 |
115.566 |
|
C2 |
C4 |
H10 |
115.566 |
| C3 |
O1 |
C4 |
91.753 |
|
C3 |
C2 |
C4 |
84.298 |
| C3 |
C2 |
H5 |
115.323 |
|
C3 |
C2 |
H6 |
115.323 |
| C4 |
C2 |
H5 |
115.323 |
|
C4 |
C2 |
H6 |
115.323 |
| H5 |
C2 |
H6 |
109.533 |
|
H7 |
C3 |
H8 |
108.887 |
| H9 |
C4 |
H10 |
108.887 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -192.584857 |
| Energy at 298.15K | -192.591994 |
| HF Energy | -191.919846 |
| Nuclear repulsion energy | 127.288255 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3154 |
3006 |
|
|
|
|
| 2 |
A' |
3092 |
2947 |
|
|
|
|
| 3 |
A' |
3079 |
2935 |
|
|
|
|
| 4 |
A' |
3035 |
2893 |
|
|
|
|
| 5 |
A' |
1537 |
1465 |
|
|
|
|
| 6 |
A' |
1477 |
1407 |
|
|
|
|
| 7 |
A' |
1381 |
1316 |
|
|
|
|
| 8 |
A' |
1186 |
1131 |
|
|
|
|
| 9 |
A' |
1144 |
1090 |
|
|
|
|
| 10 |
A' |
1054 |
1005 |
|
|
|
|
| 11 |
A' |
937 |
893 |
|
|
|
|
| 12 |
A' |
830 |
791 |
|
|
|
|
| 13 |
A' |
747 |
712 |
|
|
|
|
| 14 |
A' |
20 |
19 |
|
|
|
|
| 15 |
A" |
3081 |
2936 |
|
|
|
|
| 16 |
A" |
3026 |
2884 |
|
|
|
|
| 17 |
A" |
1507 |
1436 |
|
|
|
|
| 18 |
A" |
1306 |
1245 |
|
|
|
|
| 19 |
A" |
1256 |
1197 |
|
|
|
|
| 20 |
A" |
1223 |
1166 |
|
|
|
|
| 21 |
A" |
1162 |
1108 |
|
|
|
|
| 22 |
A" |
1050 |
1001 |
|
|
|
|
| 23 |
A" |
951 |
906 |
|
|
|
|
| 24 |
A" |
831 |
792 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19033.3 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 18140.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.071 |
-1.073 |
0.000 |
| C2 |
-0.019 |
1.082 |
0.000 |
| C3 |
-0.019 |
-0.068 |
1.040 |
| C4 |
-0.019 |
-0.068 |
-1.040 |
| H5 |
0.894 |
1.702 |
0.000 |
| H6 |
-0.909 |
1.735 |
0.000 |
| H7 |
0.848 |
-0.103 |
1.728 |
| H8 |
-0.952 |
-0.165 |
1.632 |
| H9 |
0.848 |
-0.103 |
-1.728 |
| H10 |
-0.952 |
-0.165 |
-1.632 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1576 | 1.4495 | 1.4495 | 2.8951 | 2.9738 | 2.1293 | 2.1294 | 2.1293 | 2.1294 |
C2 | 2.1576 | | 1.5505 | 1.5505 | 1.1033 | 1.1032 | 2.2681 | 2.2558 | 2.2681 | 2.2558 | C3 | 1.4495 | 1.5505 | | 2.0799 | 2.2466 | 2.2630 | 1.1078 | 1.1091 | 2.9012 | 2.8316 | C4 | 1.4495 | 1.5505 | 2.0799 | | 2.2466 | 2.2630 | 2.9012 | 2.8316 | 1.1078 | 1.1091 | H5 | 2.8951 | 1.1033 | 2.2466 | 2.2466 | | 1.8028 | 2.4996 | 3.0912 | 2.4996 | 3.0912 | H6 | 2.9738 | 1.1032 | 2.2630 | 2.2630 | 1.8028 | | 3.0741 | 2.5044 | 3.0741 | 2.5044 | H7 | 2.1293 | 2.2681 | 1.1078 | 2.9012 | 2.4996 | 3.0741 | | 1.8038 | 3.4568 | 3.8125 | H8 | 2.1294 | 2.2558 | 1.1091 | 2.8316 | 3.0912 | 2.5044 | 1.8038 | | 3.8125 | 3.2636 | H9 | 2.1293 | 2.2681 | 2.9012 | 1.1078 | 2.4996 | 3.0741 | 3.4568 | 3.8125 | | 1.8038 | H10 | 2.1294 | 2.2558 | 2.8316 | 1.1091 | 3.0912 | 2.5044 | 3.8125 | 3.2636 | 1.8038 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
91.914 |
|
O1 |
C3 |
H7 |
112.045 |
| O1 |
C3 |
H8 |
111.972 |
|
O1 |
C4 |
C2 |
91.914 |
| O1 |
C4 |
H9 |
112.045 |
|
O1 |
C4 |
H10 |
111.972 |
| C2 |
C3 |
H7 |
116.126 |
|
C2 |
C3 |
H8 |
115.019 |
| C2 |
C4 |
H9 |
116.126 |
|
C2 |
C4 |
H10 |
115.019 |
| C3 |
O1 |
C4 |
91.689 |
|
C3 |
C2 |
C4 |
84.248 |
| C3 |
C2 |
H5 |
114.630 |
|
C3 |
C2 |
H6 |
116.003 |
| C4 |
C2 |
H5 |
114.630 |
|
C4 |
C2 |
H6 |
116.003 |
| H5 |
C2 |
H6 |
109.578 |
|
H7 |
C3 |
H8 |
108.911 |
| H9 |
C4 |
H10 |
108.911 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
