Jump to
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -192.869446 |
| Energy at 298.15K | -192.876470 |
| HF Energy | -192.869446 |
| Nuclear repulsion energy | 127.236452 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3015 |
2997 |
27.03 |
|
|
|
| 2 |
A1 |
2935 |
2918 |
1.90 |
|
|
|
| 3 |
A1 |
1470 |
1462 |
0.00 |
|
|
|
| 4 |
A1 |
1412 |
1403 |
2.35 |
|
|
|
| 5 |
A1 |
1325 |
1317 |
3.55 |
|
|
|
| 6 |
A1 |
1037 |
1030 |
4.84 |
|
|
|
| 7 |
A1 |
912 |
907 |
24.94 |
|
|
|
| 8 |
A1 |
789 |
784 |
3.72 |
|
|
|
| 9 |
A2 |
2967 |
2950 |
0.00 |
|
|
|
| 10 |
A2 |
1184 |
1177 |
0.00 |
|
|
|
| 11 |
A2 |
1117 |
1111 |
0.00 |
|
|
|
| 12 |
A2 |
811 |
806 |
0.00 |
|
|
|
| 13 |
B1 |
3075 |
3057 |
24.80 |
|
|
|
| 14 |
B1 |
2966 |
2949 |
104.70 |
|
|
|
| 15 |
B1 |
1138 |
1131 |
0.87 |
|
|
|
| 16 |
B1 |
1093 |
1086 |
0.86 |
|
|
|
| 17 |
B1 |
738 |
734 |
0.50 |
|
|
|
| 18 |
B1 |
3 |
3 |
1.31 |
|
|
|
| 19 |
B2 |
2922 |
2905 |
179.49 |
|
|
|
| 20 |
B2 |
1437 |
1429 |
4.28 |
|
|
|
| 21 |
B2 |
1251 |
1244 |
0.02 |
|
|
|
| 22 |
B2 |
1209 |
1202 |
5.04 |
|
|
|
| 23 |
B2 |
1021 |
1015 |
95.47 |
|
|
|
| 24 |
B2 |
939 |
934 |
10.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18381.4 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 18274.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.079 |
| C2 |
0.000 |
0.000 |
-1.078 |
| C3 |
0.000 |
1.038 |
0.063 |
| C4 |
0.000 |
-1.038 |
0.063 |
| H5 |
0.901 |
0.000 |
-1.720 |
| H6 |
-0.901 |
0.000 |
-1.720 |
| H7 |
0.904 |
1.687 |
0.129 |
| H8 |
-0.904 |
1.687 |
0.129 |
| H9 |
-0.904 |
-1.687 |
0.129 |
| H10 |
0.904 |
-1.687 |
0.129 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1571 | 1.4525 | 1.4525 | 2.9401 | 2.9401 | 2.1361 | 2.1361 | 2.1361 | 2.1361 |
C2 | 2.1571 | | 1.5433 | 1.5433 | 1.1062 | 1.1062 | 2.2627 | 2.2627 | 2.2627 | 2.2627 | C3 | 1.4525 | 1.5433 | | 2.0769 | 2.2516 | 2.2516 | 1.1141 | 1.1141 | 2.8718 | 2.8718 | C4 | 1.4525 | 1.5433 | 2.0769 | | 2.2516 | 2.2516 | 2.8718 | 2.8718 | 1.1141 | 1.1141 | H5 | 2.9401 | 1.1062 | 2.2516 | 2.2516 | | 1.8024 | 2.5028 | 3.0857 | 3.0857 | 2.5028 | H6 | 2.9401 | 1.1062 | 2.2516 | 2.2516 | 1.8024 | | 3.0857 | 2.5028 | 2.5028 | 3.0857 | H7 | 2.1361 | 2.2627 | 1.1141 | 2.8718 | 2.5028 | 3.0857 | | 1.8073 | 3.8270 | 3.3734 | H8 | 2.1361 | 2.2627 | 1.1141 | 2.8718 | 3.0857 | 2.5028 | 1.8073 | | 3.3734 | 3.8270 | H9 | 2.1361 | 2.2627 | 2.8718 | 1.1141 | 3.0857 | 2.5028 | 3.8270 | 3.3734 | | 1.8073 | H10 | 2.1361 | 2.2627 | 2.8718 | 1.1141 | 2.5028 | 3.0857 | 3.3734 | 3.8270 | 1.8073 | |
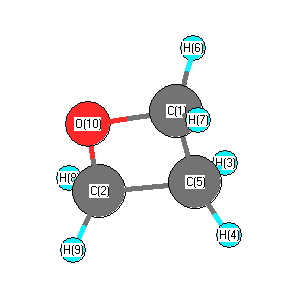 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
92.070 |
|
O1 |
C3 |
H7 |
111.996 |
| O1 |
C3 |
H8 |
111.996 |
|
O1 |
C4 |
C2 |
92.070 |
| O1 |
C4 |
H9 |
111.996 |
|
O1 |
C4 |
H10 |
111.996 |
| C2 |
C3 |
H7 |
115.811 |
|
C2 |
C3 |
H8 |
115.811 |
| C2 |
C4 |
H9 |
115.811 |
|
C2 |
C4 |
H10 |
115.811 |
| C3 |
O1 |
C4 |
91.279 |
|
C3 |
C2 |
C4 |
84.582 |
| C3 |
C2 |
H5 |
115.402 |
|
C3 |
C2 |
H6 |
115.402 |
| C4 |
C2 |
H5 |
115.402 |
|
C4 |
C2 |
H6 |
115.402 |
| H5 |
C2 |
H6 |
109.115 |
|
H7 |
C3 |
H8 |
108.410 |
| H9 |
C4 |
H10 |
108.410 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.309 |
|
|
|
| 2 |
C |
-0.124 |
|
|
|
| 3 |
C |
0.138 |
|
|
|
| 4 |
C |
0.138 |
|
|
|
| 5 |
H |
0.032 |
|
|
|
| 6 |
H |
0.032 |
|
|
|
| 7 |
H |
0.024 |
|
|
|
| 8 |
H |
0.024 |
|
|
|
| 9 |
H |
0.024 |
|
|
|
| 10 |
H |
0.024 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.721 |
1.721 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.289 |
0.000 |
0.000 |
| y |
0.000 |
-23.293 |
0.000 |
| z |
0.000 |
0.000 |
-27.401 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.058 |
0.000 |
0.000 |
| y |
0.000 |
2.552 |
0.000 |
| z |
0.000 |
0.000 |
-3.610 |
|
| Polar |
|---|
| 3z2-r2 | -7.220 |
|---|
| x2-y2 | -0.996 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.998 |
0.000 |
0.000 |
| y |
0.000 |
6.109 |
0.000 |
| z |
0.000 |
0.000 |
4.851 |
<r2> (average value of r
2) Å
2
| <r2> |
67.139 |
| (<r2>)1/2 |
8.194 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -192.869446 |
| Energy at 298.15K | -192.876514 |
| HF Energy | -192.869446 |
| Nuclear repulsion energy | 127.243057 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3074 |
3057 |
24.93 |
|
|
|
| 2 |
A' |
3015 |
2997 |
27.15 |
|
|
|
| 3 |
A' |
2967 |
2949 |
99.75 |
|
|
|
| 4 |
A' |
2934 |
2917 |
6.88 |
|
|
|
| 5 |
A' |
1470 |
1462 |
0.01 |
|
|
|
| 6 |
A' |
1412 |
1403 |
2.35 |
|
|
|
| 7 |
A' |
1324 |
1317 |
3.54 |
|
|
|
| 8 |
A' |
1138 |
1132 |
0.85 |
|
|
|
| 9 |
A' |
1093 |
1087 |
0.90 |
|
|
|
| 10 |
A' |
1037 |
1031 |
4.89 |
|
|
|
| 11 |
A' |
912 |
907 |
24.88 |
|
|
|
| 12 |
A' |
792 |
787 |
3.81 |
|
|
|
| 13 |
A' |
734 |
730 |
0.57 |
|
|
|
| 14 |
A' |
25 |
25 |
1.34 |
|
|
|
| 15 |
A" |
2968 |
2951 |
3.21 |
|
|
|
| 16 |
A" |
2921 |
2904 |
175.70 |
|
|
|
| 17 |
A" |
1437 |
1429 |
4.25 |
|
|
|
| 18 |
A" |
1251 |
1244 |
0.02 |
|
|
|
| 19 |
A" |
1210 |
1203 |
4.93 |
|
|
|
| 20 |
A" |
1183 |
1177 |
0.13 |
|
|
|
| 21 |
A" |
1117 |
1110 |
0.02 |
|
|
|
| 22 |
A" |
1020 |
1015 |
95.29 |
|
|
|
| 23 |
A" |
939 |
934 |
10.37 |
|
|
|
| 24 |
A" |
811 |
806 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18390.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 18284.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.042 |
-1.078 |
0.000 |
| C2 |
-0.011 |
1.078 |
0.000 |
| C3 |
-0.011 |
-0.064 |
1.038 |
| C4 |
-0.011 |
-0.064 |
-1.038 |
| H5 |
0.897 |
1.710 |
0.000 |
| H6 |
-0.906 |
1.729 |
0.000 |
| H7 |
0.872 |
-0.112 |
1.714 |
| H8 |
-0.934 |
-0.148 |
1.658 |
| H9 |
0.872 |
-0.112 |
-1.714 |
| H10 |
-0.934 |
-0.148 |
-1.658 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1567 | 1.4525 | 1.4525 | 2.9160 | 2.9627 | 2.1359 | 2.1364 | 2.1359 | 2.1364 |
C2 | 2.1567 | | 1.5432 | 1.5432 | 1.1062 | 1.1062 | 2.2662 | 2.2591 | 2.2662 | 2.2591 | C3 | 1.4525 | 1.5432 | | 2.0766 | 2.2468 | 2.2565 | 1.1136 | 1.1145 | 2.8914 | 2.8507 | C4 | 1.4525 | 1.5432 | 2.0766 | | 2.2468 | 2.2565 | 2.8914 | 2.8507 | 1.1136 | 1.1145 | H5 | 2.9160 | 1.1062 | 2.2468 | 2.2468 | | 1.8023 | 2.5018 | 3.0908 | 2.5018 | 3.0908 | H6 | 2.9627 | 1.1062 | 2.2565 | 2.2565 | 1.8023 | | 3.0804 | 2.5048 | 3.0804 | 2.5048 | H7 | 2.1359 | 2.2662 | 1.1136 | 2.8914 | 2.5018 | 3.0804 | | 1.8075 | 3.4288 | 3.8255 | H8 | 2.1364 | 2.2591 | 1.1145 | 2.8507 | 3.0908 | 2.5048 | 1.8075 | | 3.8255 | 3.3152 | H9 | 2.1359 | 2.2662 | 2.8914 | 1.1136 | 2.5018 | 3.0804 | 3.4288 | 3.8255 | | 1.8075 | H10 | 2.1364 | 2.2591 | 2.8507 | 1.1145 | 3.0908 | 2.5048 | 3.8255 | 3.3152 | 1.8075 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
92.046 |
|
O1 |
C3 |
H7 |
112.003 |
| O1 |
C3 |
H8 |
111.992 |
|
O1 |
C4 |
C2 |
92.046 |
| O1 |
C4 |
H9 |
112.003 |
|
O1 |
C4 |
H10 |
111.992 |
| C2 |
C3 |
H7 |
116.136 |
|
C2 |
C3 |
H8 |
115.480 |
| C2 |
C4 |
H9 |
116.136 |
|
C2 |
C4 |
H10 |
115.480 |
| C3 |
O1 |
C4 |
91.258 |
|
C3 |
C2 |
C4 |
84.568 |
| C3 |
C2 |
H5 |
115.004 |
|
C3 |
C2 |
H6 |
115.813 |
| C4 |
C2 |
H5 |
115.004 |
|
C4 |
C2 |
H6 |
115.813 |
| H5 |
C2 |
H6 |
109.101 |
|
H7 |
C3 |
H8 |
108.431 |
| H9 |
C4 |
H10 |
108.431 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.309 |
|
|
|
| 2 |
C |
-0.124 |
|
|
|
| 3 |
C |
0.137 |
|
|
|
| 4 |
C |
0.137 |
|
|
|
| 5 |
H |
0.033 |
|
|
|
| 6 |
H |
0.031 |
|
|
|
| 7 |
H |
0.024 |
|
|
|
| 8 |
H |
0.023 |
|
|
|
| 9 |
H |
0.024 |
|
|
|
| 10 |
H |
0.023 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.062 |
1.719 |
0.000 |
1.720 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.296 |
0.115 |
0.000 |
| y |
0.115 |
-27.397 |
0.000 |
| z |
0.000 |
0.000 |
-23.290 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.047 |
0.115 |
0.000 |
| y |
0.115 |
-3.604 |
0.000 |
| z |
0.000 |
0.000 |
2.557 |
|
| Polar |
|---|
| 3z2-r2 | 5.113 |
|---|
| x2-y2 | 3.100 |
|---|
| xy | 0.115 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.000 |
-0.000 |
0.000 |
| y |
-0.000 |
4.853 |
0.000 |
| z |
0.000 |
0.000 |
6.108 |
<r2> (average value of r
2) Å
2
| <r2> |
67.129 |
| (<r2>)1/2 |
8.193 |
