return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (1,3-Dioxane)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-306.834929
Energy at 298.15K-306.846303
Nuclear repulsion energy264.360942
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3171 3042 35.37      
2 A' 3141 3013 31.32      
3 A' 3130 3003 25.37      
4 A' 3072 2947 19.64      
5 A' 3005 2883 112.81      
6 A' 2960 2840 71.53      
7 A' 1516 1455 4.14      
8 A' 1501 1440 0.17      
9 A' 1468 1408 6.57      
10 A' 1424 1366 28.86      
11 A' 1326 1272 6.79      
12 A' 1228 1178 105.04      
13 A' 1205 1156 85.10      
14 A' 1122 1077 8.82      
15 A' 1020 978 38.14      
16 A' 939 901 7.97      
17 A' 863 828 12.23      
18 A' 657 630 5.03      
19 A' 491 472 0.65      
20 A' 447 429 10.76      
21 A' 269 259 1.56      
22 A" 3136 3009 45.01      
23 A" 2997 2875 20.70      
24 A" 1500 1439 5.06      
25 A" 1457 1398 21.20      
26 A" 1403 1346 0.52      
27 A" 1374 1318 0.93      
28 A" 1345 1291 1.13      
29 A" 1267 1216 34.48      
30 A" 1235 1185 0.21      
31 A" 1127 1082 75.95      
32 A" 1059 1016 54.72      
33 A" 936 898 11.53      
34 A" 909 872 0.05      
35 A" 466 448 6.32      
36 A" 272 261 2.22      

Unscaled Zero Point Vibrational Energy (zpe) 27219.7 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 26114.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.16669 0.15950 0.09185

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.629 -1.196 0.000
O2 0.016 -0.767 1.173
O3 0.016 -0.767 -1.173
C4 0.016 0.657 1.242
C5 0.016 0.657 -1.242
C6 0.696 1.243 0.000
H7 -0.604 -2.295 0.000
H8 -1.684 -0.831 0.000
H9 0.544 0.929 2.171
H10 -1.030 1.032 1.314
H11 0.544 0.929 -2.171
H12 -1.030 1.032 -1.314
H13 1.762 0.952 0.000
H14 0.630 2.348 0.000

Atom - Atom Distances (Å)
  C1 O2 O3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C11.40601.40602.32242.32242.77601.09951.11643.25652.61723.25652.61723.21443.7606
O21.40602.34631.42582.80402.42482.02392.06702.03732.08573.78673.24282.71653.3843
O31.40602.34632.80401.42582.42482.02392.06703.78673.24282.03732.08572.71653.3843
C42.32241.42582.80402.48451.53273.26252.57891.10221.11363.46462.78732.16302.1858
C52.32242.80401.42582.48451.53273.26252.57893.46462.78731.10221.11362.16302.1858
C62.77602.42482.42481.53271.53273.76973.15772.19912.17962.19912.17961.10511.1063
H71.09952.02392.02393.26253.26253.76971.81934.05293.60234.05293.60234.01784.8039
H81.11642.06702.06702.57892.57893.15771.81933.57442.37193.57442.37193.88063.9321
H93.25652.03733.78671.10223.46462.19914.05293.57441.79484.34243.82522.49002.5950
H102.61722.08573.24281.11362.78732.17963.60232.37191.79483.82522.62783.08692.4926
H113.25653.78672.03733.46461.10222.19914.05293.57444.34243.82521.79482.49002.5950
H122.61723.24282.08572.78731.11362.17963.60232.37193.82522.62781.79483.08692.4926
H133.21442.71652.71652.16302.16301.10514.01783.88062.49003.08692.49003.08691.7971
H143.76063.38433.38432.18582.18581.10634.80393.93212.59502.49262.59502.49261.7971

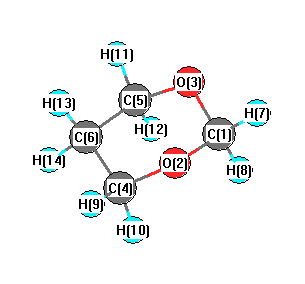
picture of 1,3-Dioxane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 C4 110.193 C1 O3 C5 110.193
O2 C1 O3 113.103 O2 C1 H7 107.127
O2 C1 H8 109.527 O2 C4 C6 110.039
O2 C4 H9 106.692 O2 C4 H10 109.829
O3 C1 H7 107.127 O3 C1 H8 109.527
O3 C5 C6 110.039 O3 C5 H11 106.692
O3 C5 H12 109.829 C4 C6 C5 108.294
C4 C6 H13 109.096 C4 C6 H14 110.807
C5 C6 H13 109.096 C5 C6 H14 110.807
C6 C4 H9 112.121 C6 C4 H10 109.890
C6 C5 H11 112.121 C6 C5 H12 109.890
H7 C1 H8 110.374 H9 C4 H10 108.195
H11 C5 H12 108.195 H13 C6 H14 108.710
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability