return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (1,3-Dioxane)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-306.826222
Energy at 298.15K-306.837619
Nuclear repulsion energy264.656295
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3178 3040 35.02      
2 A' 3150 3014 31.02      
3 A' 3141 3005 23.94      
4 A' 3083 2949 17.67      
5 A' 3017 2886 106.86      
6 A' 2973 2844 70.01      
7 A' 1523 1457 3.60      
8 A' 1506 1441 0.28      
9 A' 1472 1408 6.92      
10 A' 1431 1369 29.77      
11 A' 1332 1274 7.09      
12 A' 1239 1185 141.50      
13 A' 1214 1161 51.91      
14 A' 1128 1079 6.13      
15 A' 1029 984 38.98      
16 A' 944 903 7.94      
17 A' 870 832 10.87      
18 A' 662 633 5.12      
19 A' 494 472 0.66      
20 A' 447 427 11.05      
21 A' 269 257 1.62      
22 A" 3146 3010 43.41      
23 A" 3009 2879 20.22      
24 A" 1506 1441 6.63      
25 A" 1465 1402 20.86      
26 A" 1409 1348 0.76      
27 A" 1380 1320 1.07      
28 A" 1352 1294 1.19      
29 A" 1274 1219 39.72      
30 A" 1241 1188 0.04      
31 A" 1150 1100 75.72      
32 A" 1068 1022 46.99      
33 A" 945 904 10.05      
34 A" 913 873 0.16      
35 A" 470 449 6.31      
36 A" 272 260 2.28      

Unscaled Zero Point Vibrational Energy (zpe) 27349.4 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 26165.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.16702 0.15990 0.09202

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.626 -1.196 0.000
O2 0.016 -0.765 1.171
O3 0.016 -0.765 -1.171
C4 0.016 0.656 1.242
C5 0.016 0.656 -1.242
C6 0.694 1.244 0.000
H7 -0.597 -2.295 0.000
H8 -1.683 -0.839 0.000
H9 0.545 0.927 2.170
H10 -1.029 1.031 1.317
H11 0.545 0.927 -2.170
H12 -1.029 1.031 -1.317
H13 1.760 0.956 0.000
H14 0.626 2.348 0.000

Atom - Atom Distances (Å)
  C1 O2 O3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C11.40311.40312.32032.32032.77441.09921.11583.25382.61853.25382.61853.21353.7583
O21.40312.34141.42272.79992.42232.02142.06502.03522.08323.78173.24162.71573.3811
O31.40312.34142.79991.42272.42232.02142.06503.78173.24162.03522.08322.71573.3811
C42.32031.42272.79992.48381.53233.25952.58161.10181.11293.46292.78972.16212.1855
C52.32032.79991.42272.48381.53233.25952.58163.46292.78971.10181.11292.16212.1855
C62.77442.42232.42231.53231.53233.76723.16092.19762.17932.19762.17931.10471.1056
H71.09922.02142.02143.25953.25953.76721.81654.04883.60304.04883.60304.01564.8008
H81.11582.06502.06502.58162.58163.16091.81653.57672.37903.57672.37903.88353.9353
H93.25382.03523.78171.10183.46292.19764.04883.57671.79294.33913.82712.48692.5945
H102.61852.08323.24161.11292.78972.17933.60302.37901.79293.82712.63473.08572.4914
H113.25383.78172.03523.46291.10182.19764.04883.57674.33913.82711.79292.48692.5945
H122.61853.24162.08322.78971.11292.17933.60302.37903.82712.63471.79293.08572.4914
H133.21352.71572.71572.16212.16211.10474.01563.88352.48693.08572.48693.08571.7958
H143.75833.38113.38112.18552.18551.10564.80083.93532.59452.49142.59452.49141.7958

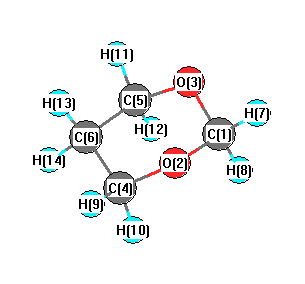
picture of 1,3-Dioxane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 C4 110.389 C1 O3 C5 110.389
O2 C1 O3 113.097 O2 C1 H7 107.144
O2 C1 H8 109.604 O2 C4 C6 110.060
O2 C4 H9 106.751 O2 C4 H10 109.877
O3 C1 H7 107.144 O3 C1 H8 109.604
O3 C5 C6 110.060 O3 C5 H11 106.751
O3 C5 H12 109.877 C4 C6 C5 108.282
C4 C6 H13 109.073 C4 C6 H14 110.853
C5 C6 H13 109.073 C5 C6 H14 110.853
C6 C4 H9 112.048 C6 C4 H10 109.931
C6 C5 H11 112.048 C6 C5 H12 109.931
H7 C1 H8 110.179 H9 C4 H10 108.100
H11 C5 H12 108.100 H13 C6 H14 108.673
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability