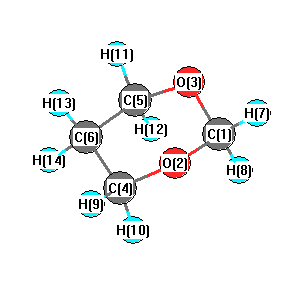
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3055 |
3037 |
29.90 |
|
|
|
| 2 |
A' |
3038 |
3021 |
27.44 |
|
|
|
| 3 |
A' |
3030 |
3012 |
21.83 |
|
|
|
| 4 |
A' |
2977 |
2960 |
19.27 |
|
|
|
| 5 |
A' |
2878 |
2862 |
111.70 |
|
|
|
| 6 |
A' |
2812 |
2796 |
92.55 |
|
|
|
| 7 |
A' |
1423 |
1414 |
5.59 |
|
|
|
| 8 |
A' |
1410 |
1401 |
1.09 |
|
|
|
| 9 |
A' |
1385 |
1377 |
6.62 |
|
|
|
| 10 |
A' |
1348 |
1340 |
19.55 |
|
|
|
| 11 |
A' |
1264 |
1257 |
5.75 |
|
|
|
| 12 |
A' |
1157 |
1150 |
56.20 |
|
|
|
| 13 |
A' |
1133 |
1126 |
115.87 |
|
|
|
| 14 |
A' |
1067 |
1061 |
9.82 |
|
|
|
| 15 |
A' |
972 |
966 |
35.84 |
|
|
|
| 16 |
A' |
897 |
892 |
9.06 |
|
|
|
| 17 |
A' |
826 |
821 |
12.55 |
|
|
|
| 18 |
A' |
625 |
622 |
3.53 |
|
|
|
| 19 |
A' |
477 |
474 |
0.42 |
|
|
|
| 20 |
A' |
427 |
425 |
9.16 |
|
|
|
| 21 |
A' |
263 |
261 |
1.36 |
|
|
|
| 22 |
A" |
3032 |
3014 |
41.95 |
|
|
|
| 23 |
A" |
2870 |
2854 |
22.77 |
|
|
|
| 24 |
A" |
1411 |
1402 |
9.05 |
|
|
|
| 25 |
A" |
1374 |
1366 |
11.08 |
|
|
|
| 26 |
A" |
1327 |
1320 |
0.48 |
|
|
|
| 27 |
A" |
1305 |
1298 |
0.62 |
|
|
|
| 28 |
A" |
1274 |
1267 |
0.90 |
|
|
|
| 29 |
A" |
1206 |
1199 |
23.58 |
|
|
|
| 30 |
A" |
1175 |
1168 |
0.64 |
|
|
|
| 31 |
A" |
1032 |
1026 |
12.43 |
|
|
|
| 32 |
A" |
991 |
986 |
112.44 |
|
|
|
| 33 |
A" |
891 |
886 |
24.34 |
|
|
|
| 34 |
A" |
866 |
861 |
0.02 |
|
|
|
| 35 |
A" |
450 |
448 |
5.59 |
|
|
|
| 36 |
A" |
259 |
257 |
1.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25963.5 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 25812.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.233 |
|
|
|
| 2 |
O |
-0.271 |
|
|
|
| 3 |
O |
-0.271 |
|
|
|
| 4 |
C |
0.090 |
|
|
|
| 5 |
C |
0.090 |
|
|
|
| 6 |
C |
-0.044 |
|
|
|
| 7 |
H |
0.034 |
|
|
|
| 8 |
H |
-0.003 |
|
|
|
| 9 |
H |
0.030 |
|
|
|
| 10 |
H |
0.012 |
|
|
|
| 11 |
H |
0.030 |
|
|
|
| 12 |
H |
0.012 |
|
|
|
| 13 |
H |
0.038 |
|
|
|
| 14 |
H |
0.019 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.511 |
1.625 |
0.000 |
1.703 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.296 |
1.533 |
0.000 |
| y |
1.533 |
-35.128 |
0.000 |
| z |
0.000 |
0.000 |
-38.565 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.551 |
1.533 |
0.000 |
| y |
1.533 |
2.303 |
0.000 |
| z |
0.000 |
0.000 |
-2.853 |
|
| Polar |
|---|
| 3z2-r2 | -5.707 |
|---|
| x2-y2 | -1.168 |
|---|
| xy | 1.533 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.124 |
0.546 |
0.000 |
| y |
0.546 |
8.142 |
0.000 |
| z |
0.000 |
0.000 |
7.619 |
<r2> (average value of r
2) Å
2
| <r2> |
140.722 |
| (<r2>)1/2 |
11.863 |