Jump to
S1C2
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -133.256206 |
| Energy at 298.15K | -133.260368 |
| HF Energy | -133.256206 |
| Nuclear repulsion energy | 64.695048 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3436 |
3316 |
0.25 |
|
|
|
| 2 |
A' |
3284 |
3169 |
2.87 |
|
|
|
| 3 |
A' |
3163 |
3052 |
2.89 |
|
|
|
| 4 |
A' |
3049 |
2942 |
50.91 |
|
|
|
| 5 |
A' |
1504 |
1452 |
9.03 |
|
|
|
| 6 |
A' |
1446 |
1395 |
2.94 |
|
|
|
| 7 |
A' |
1354 |
1306 |
36.65 |
|
|
|
| 8 |
A' |
1250 |
1206 |
6.42 |
|
|
|
| 9 |
A' |
1118 |
1079 |
21.78 |
|
|
|
| 10 |
A' |
985 |
951 |
10.19 |
|
|
|
| 11 |
A' |
482 |
465 |
10.47 |
|
|
|
| 12 |
A" |
1045 |
1008 |
0.00 |
|
|
|
| 13 |
A" |
806 |
778 |
8.10 |
|
|
|
| 14 |
A" |
669 |
645 |
92.44 |
|
|
|
| 15 |
A" |
526 |
508 |
5.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12058.0 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 11636.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.418 |
0.000 |
| C2 |
1.153 |
-0.380 |
0.000 |
| N3 |
-1.194 |
-0.135 |
0.000 |
| H4 |
0.150 |
1.513 |
0.000 |
| H5 |
2.151 |
0.061 |
0.000 |
| H6 |
1.047 |
-1.467 |
0.000 |
| H7 |
-1.906 |
0.608 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4017 | 1.3155 | 1.1052 | 2.1805 | 2.1565 | 1.9159 |
C2 | 1.4017 | | 2.3594 | 2.1420 | 1.0914 | 1.0925 | 3.2145 | N3 | 1.3155 | 2.3594 | | 2.1257 | 3.3507 | 2.6076 | 1.0290 | H4 | 1.1052 | 2.1420 | 2.1257 | | 2.4726 | 3.1124 | 2.2463 | H5 | 2.1805 | 1.0914 | 3.3507 | 2.4726 | | 1.8851 | 4.0942 | H6 | 2.1565 | 1.0925 | 2.6076 | 3.1124 | 1.8851 | | 3.6099 | H7 | 1.9159 | 3.2145 | 1.0290 | 2.2463 | 4.0942 | 3.6099 | |
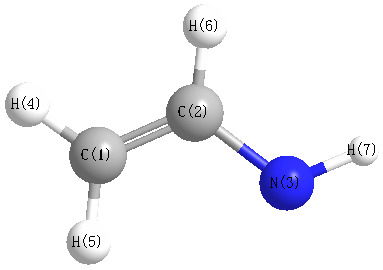 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.496 |
|
C1 |
C2 |
H6 |
119.154 |
| C1 |
N3 |
H7 |
108.998 |
|
C2 |
C1 |
N3 |
120.493 |
| C2 |
C1 |
H4 |
116.906 |
|
N3 |
C1 |
H4 |
122.601 |
| H5 |
C2 |
H6 |
119.350 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.050 |
|
|
|
| 2 |
C |
-0.004 |
|
|
|
| 3 |
N |
-0.173 |
|
|
|
| 4 |
H |
0.012 |
|
|
|
| 5 |
H |
0.057 |
|
|
|
| 6 |
H |
0.059 |
|
|
|
| 7 |
H |
0.100 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.928 |
1.485 |
0.000 |
1.751 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.676 |
-2.595 |
0.000 |
| y |
-2.595 |
-17.535 |
0.000 |
| z |
0.000 |
0.000 |
-19.805 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.994 |
-2.595 |
0.000 |
| y |
-2.595 |
0.706 |
0.000 |
| z |
0.000 |
0.000 |
-2.700 |
|
| Polar |
|---|
| 3z2-r2 | -5.399 |
|---|
| x2-y2 | 0.859 |
|---|
| xy | -2.595 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.488 |
-0.474 |
0.000 |
| y |
-0.474 |
4.252 |
0.000 |
| z |
0.000 |
0.000 |
2.049 |
<r2> (average value of r
2) Å
2
| <r2> |
45.431 |
| (<r2>)1/2 |
6.740 |
Jump to
S1C1
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -133.255967 |
| Energy at 298.15K | -133.260112 |
| HF Energy | -133.255967 |
| Nuclear repulsion energy | 64.694480 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3382 |
3263 |
7.41 |
|
|
|
| 2 |
A' |
3262 |
3148 |
6.48 |
|
|
|
| 3 |
A' |
3146 |
3036 |
3.45 |
|
|
|
| 4 |
A' |
3111 |
3002 |
22.53 |
|
|
|
| 5 |
A' |
1482 |
1430 |
5.34 |
|
|
|
| 6 |
A' |
1445 |
1394 |
3.88 |
|
|
|
| 7 |
A' |
1381 |
1333 |
12.09 |
|
|
|
| 8 |
A' |
1245 |
1202 |
32.85 |
|
|
|
| 9 |
A' |
1146 |
1106 |
29.93 |
|
|
|
| 10 |
A' |
978 |
943 |
0.79 |
|
|
|
| 11 |
A' |
485 |
468 |
7.79 |
|
|
|
| 12 |
A" |
1072 |
1035 |
46.19 |
|
|
|
| 13 |
A" |
812 |
783 |
56.57 |
|
|
|
| 14 |
A" |
678 |
654 |
5.71 |
|
|
|
| 15 |
A" |
483 |
466 |
0.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12053.6 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 11631.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.447 |
0.000 |
| C2 |
1.130 |
-0.384 |
0.000 |
| N3 |
-1.260 |
0.071 |
0.000 |
| H4 |
0.172 |
1.535 |
0.000 |
| H5 |
2.141 |
0.029 |
0.000 |
| H6 |
1.020 |
-1.474 |
0.000 |
| H7 |
-1.292 |
-0.962 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4026 | 1.3148 | 1.1013 | 2.1813 | 2.1742 | 1.9108 |
C2 | 1.4026 | | 2.4328 | 2.1450 | 1.0923 | 1.0946 | 2.4892 | N3 | 1.3148 | 2.4328 | | 2.0472 | 3.4011 | 2.7537 | 1.0331 | H4 | 1.1013 | 2.1450 | 2.0472 | | 2.4790 | 3.1254 | 2.8933 | H5 | 2.1813 | 1.0923 | 3.4011 | 2.4790 | | 1.8747 | 3.5725 | H6 | 2.1742 | 1.0946 | 2.7537 | 3.1254 | 1.8747 | | 2.3673 | H7 | 1.9108 | 2.4892 | 1.0331 | 2.8933 | 3.5725 | 2.3673 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.420 |
|
C1 |
C2 |
H6 |
120.569 |
| C1 |
N3 |
H7 |
108.354 |
|
C2 |
C1 |
N3 |
127.055 |
| C2 |
C1 |
H4 |
117.381 |
|
N3 |
C1 |
H4 |
115.564 |
| H5 |
C2 |
H6 |
118.011 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.036 |
|
|
|
| 2 |
C |
-0.034 |
|
|
|
| 3 |
N |
-0.170 |
|
|
|
| 4 |
H |
0.040 |
|
|
|
| 5 |
H |
0.060 |
|
|
|
| 6 |
H |
0.053 |
|
|
|
| 7 |
H |
0.088 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.948 |
-1.027 |
0.000 |
2.202 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.358 |
1.996 |
0.000 |
| y |
1.996 |
-16.218 |
0.000 |
| z |
0.000 |
0.000 |
-19.811 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.344 |
1.996 |
0.000 |
| y |
1.996 |
3.867 |
0.000 |
| z |
0.000 |
0.000 |
-1.523 |
|
| Polar |
|---|
| 3z2-r2 | -3.046 |
|---|
| x2-y2 | -4.140 |
|---|
| xy | 1.996 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.159 |
-0.115 |
0.000 |
| y |
-0.115 |
4.401 |
0.000 |
| z |
0.000 |
0.000 |
2.061 |
<r2> (average value of r
2) Å
2
| <r2> |
45.403 |
| (<r2>)1/2 |
6.738 |
