Jump to
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -132.917694 |
| Energy at 298.15K | -132.921850 |
| HF Energy | -132.478162 |
| Nuclear repulsion energy | 64.276208 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3445 |
3305 |
1.86 |
|
|
|
| 2 |
A' |
3299 |
3165 |
4.08 |
|
|
|
| 3 |
A' |
3182 |
3053 |
3.74 |
|
|
|
| 4 |
A' |
3100 |
2975 |
42.50 |
|
|
|
| 5 |
A' |
1517 |
1455 |
9.83 |
|
|
|
| 6 |
A' |
1460 |
1401 |
2.19 |
|
|
|
| 7 |
A' |
1367 |
1311 |
28.17 |
|
|
|
| 8 |
A' |
1243 |
1193 |
8.99 |
|
|
|
| 9 |
A' |
1112 |
1067 |
20.22 |
|
|
|
| 10 |
A' |
994 |
953 |
8.62 |
|
|
|
| 11 |
A' |
482 |
462 |
10.72 |
|
|
|
| 12 |
A" |
1046 |
1003 |
0.01 |
|
|
|
| 13 |
A" |
774 |
743 |
3.60 |
|
|
|
| 14 |
A" |
682 |
654 |
97.54 |
|
|
|
| 15 |
A" |
521 |
500 |
3.86 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12111.6 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 11619.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.425 |
0.000 |
| C2 |
1.160 |
-0.387 |
0.000 |
| N3 |
-1.202 |
-0.135 |
0.000 |
| H4 |
0.146 |
1.521 |
0.000 |
| H5 |
2.164 |
0.051 |
0.000 |
| H6 |
1.048 |
-1.476 |
0.000 |
| H7 |
-1.909 |
0.617 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4160 | 1.3254 | 1.1057 | 2.1956 | 2.1707 | 1.9182 |
C2 | 1.4160 | | 2.3754 | 2.1605 | 1.0947 | 1.0953 | 3.2289 | N3 | 1.3254 | 2.3754 | | 2.1346 | 3.3703 | 2.6191 | 1.0319 | H4 | 1.1057 | 2.1605 | 2.1346 | | 2.4961 | 3.1299 | 2.2445 | H5 | 2.1956 | 1.0947 | 3.3703 | 2.4961 | | 1.8916 | 4.1113 | H6 | 2.1707 | 1.0953 | 2.6191 | 3.1299 | 1.8916 | | 3.6224 | H7 | 1.9182 | 3.2289 | 1.0319 | 2.2445 | 4.1113 | 3.6224 | |
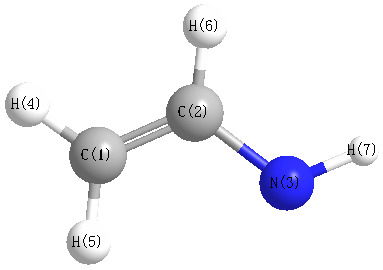 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.448 |
|
C1 |
C2 |
H6 |
119.071 |
| C1 |
N3 |
H7 |
108.274 |
|
C2 |
C1 |
N3 |
120.065 |
| C2 |
C1 |
H4 |
117.384 |
|
N3 |
C1 |
H4 |
122.551 |
| H5 |
C2 |
H6 |
119.481 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -132.917591 |
| Energy at 298.15K | -132.921737 |
| HF Energy | -132.477740 |
| Nuclear repulsion energy | 64.291280 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3397 |
3259 |
9.69 |
|
|
|
| 2 |
A' |
3280 |
3147 |
7.63 |
|
|
|
| 3 |
A' |
3168 |
3039 |
2.62 |
|
|
|
| 4 |
A' |
3156 |
3028 |
22.09 |
|
|
|
| 5 |
A' |
1505 |
1444 |
5.37 |
|
|
|
| 6 |
A' |
1457 |
1398 |
3.47 |
|
|
|
| 7 |
A' |
1384 |
1328 |
19.58 |
|
|
|
| 8 |
A' |
1241 |
1191 |
27.25 |
|
|
|
| 9 |
A' |
1137 |
1091 |
21.31 |
|
|
|
| 10 |
A' |
985 |
945 |
0.39 |
|
|
|
| 11 |
A' |
487 |
468 |
8.89 |
|
|
|
| 12 |
A" |
1080 |
1036 |
49.48 |
|
|
|
| 13 |
A" |
784 |
752 |
56.00 |
|
|
|
| 14 |
A" |
682 |
654 |
1.02 |
|
|
|
| 15 |
A" |
489 |
469 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12115.8 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 11623.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.456 |
0.000 |
| C2 |
1.136 |
-0.392 |
0.000 |
| N3 |
-1.268 |
0.072 |
0.000 |
| H4 |
0.173 |
1.544 |
0.000 |
| H5 |
2.152 |
0.018 |
0.000 |
| H6 |
1.016 |
-1.483 |
0.000 |
| H7 |
-1.279 |
-0.964 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4171 | 1.3249 | 1.1020 | 2.1957 | 2.1886 | 1.9109 |
C2 | 1.4171 | | 2.4480 | 2.1621 | 1.0952 | 1.0975 | 2.4815 | N3 | 1.3249 | 2.4480 | | 2.0602 | 3.4200 | 2.7626 | 1.0358 | H4 | 1.1020 | 2.1621 | 2.0602 | | 2.4990 | 3.1421 | 2.8981 | H5 | 2.1957 | 1.0952 | 3.4200 | 2.4990 | | 1.8818 | 3.5681 | H6 | 2.1886 | 1.0975 | 2.7626 | 3.1421 | 1.8818 | | 2.3526 | H7 | 1.9109 | 2.4815 | 1.0358 | 2.8981 | 3.5681 | 2.3526 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.311 |
|
C1 |
C2 |
H6 |
120.463 |
| C1 |
N3 |
H7 |
107.456 |
|
C2 |
C1 |
N3 |
126.411 |
| C2 |
C1 |
H4 |
117.706 |
|
N3 |
C1 |
H4 |
115.883 |
| H5 |
C2 |
H6 |
118.226 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
