Jump to
S1C2
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -132.905444 |
| Energy at 298.15K | -132.909657 |
| HF Energy | -132.478371 |
| Nuclear repulsion energy | 64.629390 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3478 |
3327 |
1.08 |
|
|
|
| 2 |
A' |
3316 |
3173 |
2.94 |
|
|
|
| 3 |
A' |
3198 |
3059 |
4.04 |
|
|
|
| 4 |
A' |
3112 |
2977 |
39.83 |
|
|
|
| 5 |
A' |
1525 |
1459 |
16.98 |
|
|
|
| 6 |
A' |
1463 |
1399 |
12.35 |
|
|
|
| 7 |
A' |
1378 |
1318 |
15.27 |
|
|
|
| 8 |
A' |
1230 |
1177 |
24.69 |
|
|
|
| 9 |
A' |
1085 |
1038 |
5.13 |
|
|
|
| 10 |
A' |
1008 |
964 |
9.12 |
|
|
|
| 11 |
A' |
491 |
469 |
11.56 |
|
|
|
| 12 |
A" |
1099 |
1052 |
0.03 |
|
|
|
| 13 |
A" |
813 |
778 |
5.05 |
|
|
|
| 14 |
A" |
715 |
684 |
97.98 |
|
|
|
| 15 |
A" |
545 |
521 |
3.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12226.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 11697.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.421 |
0.000 |
| C2 |
1.152 |
-0.392 |
0.000 |
| N3 |
-1.193 |
-0.124 |
0.000 |
| H4 |
0.154 |
1.515 |
0.000 |
| H5 |
2.156 |
0.040 |
0.000 |
| H6 |
1.034 |
-1.480 |
0.000 |
| H7 |
-1.899 |
0.626 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4095 | 1.3121 | 1.1056 | 2.1898 | 2.1638 | 1.9103 |
C2 | 1.4095 | | 2.3602 | 2.1527 | 1.0939 | 1.0945 | 3.2161 | N3 | 1.3121 | 2.3602 | | 2.1225 | 3.3540 | 2.6074 | 1.0301 | H4 | 1.1056 | 2.1527 | 2.1225 | | 2.4874 | 3.1223 | 2.2377 | H5 | 2.1898 | 1.0939 | 3.3540 | 2.4874 | | 1.8899 | 4.0978 | H6 | 2.1638 | 1.0945 | 2.6074 | 3.1223 | 1.8899 | | 3.6108 | H7 | 1.9103 | 3.2161 | 1.0301 | 2.2377 | 4.0978 | 3.6108 | |
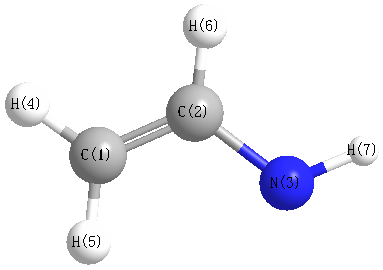 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.517 |
|
C1 |
C2 |
H6 |
119.039 |
| C1 |
N3 |
H7 |
108.701 |
|
C2 |
C1 |
N3 |
120.236 |
| C2 |
C1 |
H4 |
117.210 |
|
N3 |
C1 |
H4 |
122.554 |
| H5 |
C2 |
H6 |
119.444 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -132.905148 |
| Energy at 298.15K | -132.909361 |
| HF Energy | -132.477990 |
| Nuclear repulsion energy | 64.639887 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3429 |
3281 |
7.76 |
|
|
|
| 2 |
A' |
3298 |
3156 |
5.78 |
|
|
|
| 3 |
A' |
3185 |
3047 |
2.68 |
|
|
|
| 4 |
A' |
3165 |
3028 |
21.18 |
|
|
|
| 5 |
A' |
1516 |
1451 |
5.89 |
|
|
|
| 6 |
A' |
1458 |
1395 |
2.39 |
|
|
|
| 7 |
A' |
1381 |
1321 |
29.87 |
|
|
|
| 8 |
A' |
1230 |
1177 |
28.99 |
|
|
|
| 9 |
A' |
1109 |
1061 |
22.99 |
|
|
|
| 10 |
A' |
1000 |
956 |
0.41 |
|
|
|
| 11 |
A' |
497 |
475 |
8.96 |
|
|
|
| 12 |
A" |
1125 |
1076 |
48.17 |
|
|
|
| 13 |
A" |
825 |
789 |
60.50 |
|
|
|
| 14 |
A" |
719 |
687 |
1.45 |
|
|
|
| 15 |
A" |
521 |
499 |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12228.5 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 11699.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.452 |
0.000 |
| C2 |
1.127 |
-0.395 |
0.000 |
| N3 |
-1.258 |
0.078 |
0.000 |
| H4 |
0.181 |
1.539 |
0.000 |
| H5 |
2.143 |
0.012 |
0.000 |
| H6 |
1.005 |
-1.484 |
0.000 |
| H7 |
-1.282 |
-0.956 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4096 | 1.3128 | 1.1020 | 2.1879 | 2.1820 | 1.9042 |
C2 | 1.4096 | | 2.4315 | 2.1530 | 1.0944 | 1.0965 | 2.4734 | N3 | 1.3128 | 2.4315 | | 2.0510 | 3.4020 | 2.7503 | 1.0338 | H4 | 1.1020 | 2.1530 | 2.0510 | | 2.4868 | 3.1341 | 2.8922 | H5 | 2.1879 | 1.0944 | 3.4020 | 2.4868 | | 1.8796 | 3.5591 | H6 | 2.1820 | 1.0965 | 2.7503 | 3.1341 | 1.8796 | | 2.3476 | H7 | 1.9042 | 2.4734 | 1.0338 | 2.8922 | 3.5591 | 2.3476 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.278 |
|
C1 |
C2 |
H6 |
120.558 |
| C1 |
N3 |
H7 |
107.889 |
|
C2 |
C1 |
N3 |
126.501 |
| C2 |
C1 |
H4 |
117.489 |
|
N3 |
C1 |
H4 |
116.010 |
| H5 |
C2 |
H6 |
118.164 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
