Jump to
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -133.130861 |
| Energy at 298.15K | -133.134955 |
| HF Energy | -133.130861 |
| Nuclear repulsion energy | 64.280411 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3313 |
3294 |
1.73 |
|
|
|
| 2 |
A' |
3206 |
3187 |
3.59 |
|
|
|
| 3 |
A' |
3085 |
3067 |
2.57 |
|
|
|
| 4 |
A' |
2950 |
2933 |
59.61 |
|
|
|
| 5 |
A' |
1471 |
1462 |
4.25 |
|
|
|
| 6 |
A' |
1403 |
1395 |
4.16 |
|
|
|
| 7 |
A' |
1311 |
1303 |
38.51 |
|
|
|
| 8 |
A' |
1226 |
1218 |
4.42 |
|
|
|
| 9 |
A' |
1098 |
1092 |
23.04 |
|
|
|
| 10 |
A' |
953 |
948 |
11.83 |
|
|
|
| 11 |
A' |
468 |
466 |
9.31 |
|
|
|
| 12 |
A" |
1009 |
1003 |
0.00 |
|
|
|
| 13 |
A" |
785 |
780 |
7.61 |
|
|
|
| 14 |
A" |
635 |
632 |
82.77 |
|
|
|
| 15 |
A" |
512 |
509 |
5.62 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11712.0 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 11644.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.421 |
0.000 |
| C2 |
1.159 |
-0.380 |
0.000 |
| N3 |
-1.201 |
-0.140 |
0.000 |
| H4 |
0.155 |
1.525 |
0.000 |
| H5 |
2.164 |
0.065 |
0.000 |
| H6 |
1.051 |
-1.475 |
0.000 |
| H7 |
-1.917 |
0.614 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4085 | 1.3253 | 1.1145 | 2.1928 | 2.1674 | 1.9265 |
C2 | 1.4085 | | 2.3720 | 2.1524 | 1.0989 | 1.1003 | 3.2321 | N3 | 1.3253 | 2.3720 | | 2.1469 | 3.3709 | 2.6179 | 1.0391 | H4 | 1.1145 | 2.1524 | 2.1469 | | 2.4825 | 3.1300 | 2.2637 | H5 | 2.1928 | 1.0989 | 3.3709 | 2.4825 | | 1.8999 | 4.1172 | H6 | 2.1674 | 1.1003 | 2.6179 | 3.1300 | 1.8999 | | 3.6288 | H7 | 1.9265 | 3.2321 | 1.0391 | 2.2637 | 4.1172 | 3.6288 | |
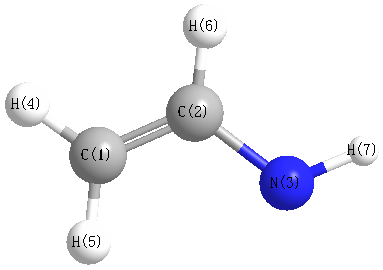 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.483 |
|
C1 |
C2 |
H6 |
119.007 |
| C1 |
N3 |
H7 |
108.525 |
|
C2 |
C1 |
N3 |
120.343 |
| C2 |
C1 |
H4 |
116.619 |
|
N3 |
C1 |
H4 |
123.038 |
| H5 |
C2 |
H6 |
119.510 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.045 |
|
|
|
| 2 |
C |
0.025 |
|
|
|
| 3 |
N |
-0.145 |
|
|
|
| 4 |
H |
-0.006 |
|
|
|
| 5 |
H |
0.042 |
|
|
|
| 6 |
H |
0.041 |
|
|
|
| 7 |
H |
0.089 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.875 |
1.408 |
0.000 |
1.658 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.831 |
-2.501 |
0.000 |
| y |
-2.501 |
-17.780 |
0.000 |
| z |
0.000 |
0.000 |
-19.862 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.990 |
-2.501 |
0.000 |
| y |
-2.501 |
0.566 |
0.000 |
| z |
0.000 |
0.000 |
-2.557 |
|
| Polar |
|---|
| 3z2-r2 | -5.113 |
|---|
| x2-y2 | 0.949 |
|---|
| xy | -2.501 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.716 |
-0.513 |
0.000 |
| y |
-0.513 |
4.470 |
0.000 |
| z |
0.000 |
0.000 |
2.067 |
<r2> (average value of r
2) Å
2
| <r2> |
45.923 |
| (<r2>)1/2 |
6.777 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -133.130867 |
| Energy at 298.15K | -133.134929 |
| HF Energy | -133.130867 |
| Nuclear repulsion energy | 64.261241 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3248 |
3229 |
14.90 |
|
|
|
| 2 |
A' |
3180 |
3162 |
8.30 |
|
|
|
| 3 |
A' |
3066 |
3048 |
4.18 |
|
|
|
| 4 |
A' |
3021 |
3003 |
23.72 |
|
|
|
| 5 |
A' |
1442 |
1434 |
5.31 |
|
|
|
| 6 |
A' |
1407 |
1399 |
4.20 |
|
|
|
| 7 |
A' |
1350 |
1342 |
10.83 |
|
|
|
| 8 |
A' |
1221 |
1214 |
29.32 |
|
|
|
| 9 |
A' |
1122 |
1115 |
30.73 |
|
|
|
| 10 |
A' |
948 |
943 |
0.56 |
|
|
|
| 11 |
A' |
472 |
469 |
7.05 |
|
|
|
| 12 |
A" |
1039 |
1033 |
42.39 |
|
|
|
| 13 |
A" |
788 |
784 |
49.48 |
|
|
|
| 14 |
A" |
646 |
643 |
6.32 |
|
|
|
| 15 |
A" |
450 |
447 |
0.50 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11699.6 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 11631.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.446 |
0.000 |
| C2 |
1.138 |
-0.385 |
0.000 |
| N3 |
-1.270 |
0.073 |
0.000 |
| H4 |
0.173 |
1.543 |
0.000 |
| H5 |
2.156 |
0.033 |
0.000 |
| H6 |
1.028 |
-1.482 |
0.000 |
| H7 |
-1.294 |
-0.971 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4095 | 1.3241 | 1.1101 | 2.1951 | 2.1850 | 1.9196 |
C2 | 1.4095 | | 2.4516 | 2.1561 | 1.1003 | 1.1025 | 2.5022 | N3 | 1.3241 | 2.4516 | | 2.0602 | 3.4264 | 2.7744 | 1.0443 | H4 | 1.1101 | 2.1561 | 2.0602 | | 2.4923 | 3.1433 | 2.9110 | H5 | 2.1951 | 1.1003 | 3.4264 | 2.4923 | | 1.8892 | 3.5935 | H6 | 2.1850 | 1.1025 | 2.7744 | 3.1433 | 1.8892 | | 2.3774 | H7 | 1.9196 | 2.5022 | 1.0443 | 2.9110 | 3.5935 | 2.3774 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.516 |
|
C1 |
C2 |
H6 |
120.387 |
| C1 |
N3 |
H7 |
107.709 |
|
C2 |
C1 |
N3 |
127.465 |
| C2 |
C1 |
H4 |
117.186 |
|
N3 |
C1 |
H4 |
115.349 |
| H5 |
C2 |
H6 |
118.098 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.036 |
|
|
|
| 2 |
C |
-0.004 |
|
|
|
| 3 |
N |
-0.140 |
|
|
|
| 4 |
H |
0.023 |
|
|
|
| 5 |
H |
0.044 |
|
|
|
| 6 |
H |
0.037 |
|
|
|
| 7 |
H |
0.076 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.862 |
-0.983 |
0.000 |
2.105 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.431 |
1.909 |
0.000 |
| y |
1.909 |
-16.470 |
0.000 |
| z |
0.000 |
0.000 |
-19.869 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.261 |
1.909 |
0.000 |
| y |
1.909 |
3.680 |
0.000 |
| z |
0.000 |
0.000 |
-1.419 |
|
| Polar |
|---|
| 3z2-r2 | -2.837 |
|---|
| x2-y2 | -3.960 |
|---|
| xy | 1.909 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.382 |
-0.119 |
0.000 |
| y |
-0.119 |
4.610 |
0.000 |
| z |
0.000 |
0.000 |
2.079 |
<r2> (average value of r
2) Å
2
| <r2> |
45.940 |
| (<r2>)1/2 |
6.778 |
