Jump to
S1C2
Energy calculated at MP3=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -132.908152 |
| Energy at 298.15K | -132.912377 |
| HF Energy | -132.478503 |
| Nuclear repulsion energy | 64.747740 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3505 |
3296 |
0.54 |
|
|
|
| 2 |
A' |
3340 |
3140 |
2.50 |
|
|
|
| 3 |
A' |
3220 |
3028 |
3.65 |
|
|
|
| 4 |
A' |
3131 |
2944 |
39.87 |
|
|
|
| 5 |
A' |
1530 |
1439 |
17.65 |
|
|
|
| 6 |
A' |
1466 |
1379 |
13.95 |
|
|
|
| 7 |
A' |
1383 |
1300 |
14.84 |
|
|
|
| 8 |
A' |
1233 |
1160 |
25.88 |
|
|
|
| 9 |
A' |
1081 |
1016 |
4.12 |
|
|
|
| 10 |
A' |
1011 |
951 |
9.49 |
|
|
|
| 11 |
A' |
492 |
463 |
11.81 |
|
|
|
| 12 |
A" |
1110 |
1043 |
0.02 |
|
|
|
| 13 |
A" |
822 |
773 |
5.77 |
|
|
|
| 14 |
A" |
720 |
677 |
98.74 |
|
|
|
| 15 |
A" |
549 |
516 |
3.91 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12296.1 cm
-1
Scaled (by 0.9403) Zero Point Vibrational Energy (zpe) 11562.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3=FULL/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.420 |
0.000 |
| C2 |
1.149 |
-0.392 |
0.000 |
| N3 |
-1.191 |
-0.123 |
0.000 |
| H4 |
0.155 |
1.513 |
0.000 |
| H5 |
2.153 |
0.038 |
0.000 |
| H6 |
1.030 |
-1.478 |
0.000 |
| H7 |
-1.895 |
0.625 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4075 | 1.3094 | 1.1035 | 2.1863 | 2.1595 | 1.9065 |
C2 | 1.4075 | | 2.3561 | 2.1488 | 1.0917 | 1.0924 | 3.2104 | N3 | 1.3094 | 2.3561 | | 2.1188 | 3.3479 | 2.6016 | 1.0278 | H4 | 1.1035 | 2.1488 | 2.1188 | | 2.4829 | 3.1160 | 2.2343 | H5 | 2.1863 | 1.0917 | 3.3479 | 2.4829 | | 1.8868 | 4.0905 | H6 | 2.1595 | 1.0924 | 2.6016 | 3.1160 | 1.8868 | | 3.6029 | H7 | 1.9065 | 3.2104 | 1.0278 | 2.2343 | 4.0905 | 3.6029 | |
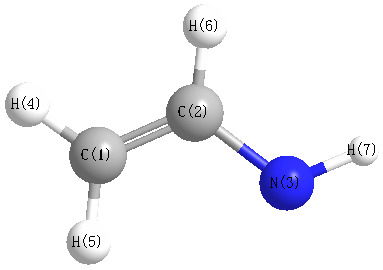 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.531 |
|
C1 |
C2 |
H6 |
118.962 |
| C1 |
N3 |
H7 |
108.722 |
|
C2 |
C1 |
N3 |
120.226 |
| C2 |
C1 |
H4 |
117.173 |
|
N3 |
C1 |
H4 |
122.601 |
| H5 |
C2 |
H6 |
119.507 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP3=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -132.907880 |
| Energy at 298.15K | -132.912102 |
| HF Energy | -132.478126 |
| Nuclear repulsion energy | 64.760821 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3453 |
3247 |
7.03 |
|
|
|
| 2 |
A' |
3322 |
3123 |
5.27 |
|
|
|
| 3 |
A' |
3206 |
3015 |
2.34 |
|
|
|
| 4 |
A' |
3186 |
2996 |
20.59 |
|
|
|
| 5 |
A' |
1521 |
1431 |
6.26 |
|
|
|
| 6 |
A' |
1461 |
1374 |
2.37 |
|
|
|
| 7 |
A' |
1384 |
1302 |
30.07 |
|
|
|
| 8 |
A' |
1233 |
1159 |
30.92 |
|
|
|
| 9 |
A' |
1104 |
1039 |
23.96 |
|
|
|
| 10 |
A' |
1003 |
943 |
0.36 |
|
|
|
| 11 |
A' |
498 |
468 |
9.10 |
|
|
|
| 12 |
A" |
1134 |
1066 |
48.63 |
|
|
|
| 13 |
A" |
833 |
783 |
61.63 |
|
|
|
| 14 |
A" |
725 |
682 |
1.64 |
|
|
|
| 15 |
A" |
525 |
494 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12294.4 cm
-1
Scaled (by 0.9403) Zero Point Vibrational Energy (zpe) 11560.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3=FULL/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.452 |
0.000 |
| C2 |
1.125 |
-0.394 |
0.000 |
| N3 |
-1.256 |
0.078 |
0.000 |
| H4 |
0.181 |
1.537 |
0.000 |
| H5 |
2.139 |
0.010 |
0.000 |
| H6 |
1.001 |
-1.482 |
0.000 |
| H7 |
-1.279 |
-0.954 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4075 | 1.3103 | 1.0998 | 2.1843 | 2.1775 | 1.9002 |
C2 | 1.4075 | | 2.4268 | 2.1492 | 1.0922 | 1.0945 | 2.4676 | N3 | 1.3103 | 2.4268 | | 2.0477 | 3.3956 | 2.7434 | 1.0318 | H4 | 1.0998 | 2.1492 | 2.0477 | | 2.4827 | 3.1279 | 2.8867 | H5 | 2.1843 | 1.0922 | 3.3956 | 2.4827 | | 1.8765 | 3.5512 | H6 | 2.1775 | 1.0945 | 2.7434 | 3.1279 | 1.8765 | | 2.3403 | H7 | 1.9002 | 2.4676 | 1.0318 | 2.8867 | 3.5512 | 2.3403 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.300 |
|
C1 |
C2 |
H6 |
120.483 |
| C1 |
N3 |
H7 |
107.864 |
|
C2 |
C1 |
N3 |
126.452 |
| C2 |
C1 |
H4 |
117.480 |
|
N3 |
C1 |
H4 |
116.068 |
| H5 |
C2 |
H6 |
118.218 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
