Jump to
S1C2
Energy calculated at ROHF/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -132.459629 |
| Energy at 298.15K | -132.463716 |
| HF Energy | -132.459629 |
| Nuclear repulsion energy | 65.128210 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at ROHF/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3712 |
3195 |
1.57 |
|
|
|
| 2 |
A' |
3443 |
2963 |
4.17 |
|
|
|
| 3 |
A' |
3321 |
2859 |
3.28 |
|
|
|
| 4 |
A' |
3217 |
2769 |
55.53 |
|
|
|
| 5 |
A' |
1818 |
1565 |
61.49 |
|
|
|
| 6 |
A' |
1586 |
1365 |
24.63 |
|
|
|
| 7 |
A' |
1518 |
1306 |
30.04 |
|
|
|
| 8 |
A' |
1366 |
1176 |
13.55 |
|
|
|
| 9 |
A' |
1134 |
976 |
56.30 |
|
|
|
| 10 |
A' |
1034 |
890 |
1.16 |
|
|
|
| 11 |
A' |
515 |
444 |
15.94 |
|
|
|
| 12 |
A" |
1215 |
1045 |
6.85 |
|
|
|
| 13 |
A" |
822 |
707 |
44.06 |
|
|
|
| 14 |
A" |
592 |
509 |
65.57 |
|
|
|
| 15 |
A" |
387 |
333 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12839.9 cm
-1
Scaled (by 0.8607) Zero Point Vibrational Energy (zpe) 11051.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at ROHF/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.416 |
0.000 |
| C2 |
1.134 |
-0.483 |
0.000 |
| N3 |
-1.185 |
-0.024 |
0.000 |
| H4 |
0.234 |
1.485 |
0.000 |
| H5 |
2.146 |
-0.106 |
0.000 |
| H6 |
0.958 |
-1.549 |
0.000 |
| H7 |
-1.848 |
0.738 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4471 | 1.2638 | 1.0937 | 2.2089 | 2.1862 | 1.8758 |
C2 | 1.4471 | | 2.3637 | 2.1637 | 1.0801 | 1.0802 | 3.2221 | N3 | 1.2638 | 2.3637 | | 2.0705 | 3.3320 | 2.6303 | 1.0100 | H4 | 1.0937 | 2.1637 | 2.0705 | | 2.4878 | 3.1187 | 2.2114 | H5 | 2.2089 | 1.0801 | 3.3320 | 2.4878 | | 1.8687 | 4.0824 | H6 | 2.1862 | 1.0802 | 2.6303 | 3.1187 | 1.8687 | | 3.6200 | H7 | 1.8758 | 3.2221 | 1.0100 | 2.2114 | 4.0824 | 3.6200 | |
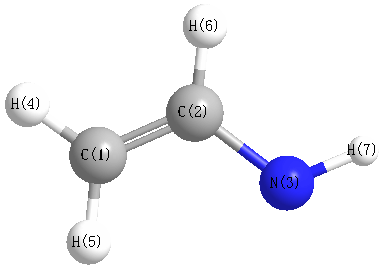 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.181 |
|
C1 |
C2 |
H6 |
119.058 |
| C1 |
N3 |
H7 |
110.669 |
|
C2 |
C1 |
N3 |
121.216 |
| C2 |
C1 |
H4 |
116.080 |
|
N3 |
C1 |
H4 |
122.705 |
| H5 |
C2 |
H6 |
119.761 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at ROHF/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.014 |
|
|
|
| 2 |
C |
-0.027 |
|
|
|
| 3 |
N |
-0.233 |
|
|
|
| 4 |
H |
0.017 |
|
|
|
| 5 |
H |
0.060 |
|
|
|
| 6 |
H |
0.068 |
|
|
|
| 7 |
H |
0.101 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.276 |
1.634 |
0.000 |
2.073 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.578 |
-2.592 |
-0.003 |
| y |
-2.592 |
-17.205 |
0.001 |
| z |
-0.003 |
0.001 |
-20.100 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.074 |
-2.592 |
-0.003 |
| y |
-2.592 |
1.635 |
0.001 |
| z |
-0.003 |
0.001 |
-2.709 |
|
| Polar |
|---|
| 3z2-r2 | -5.418 |
|---|
| x2-y2 | -0.373 |
|---|
| xy | -2.592 |
|---|
| xz | -0.003 |
|---|
| yz | 0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
45.561 |
| (<r2>)1/2 |
6.750 |
Jump to
S1C1
Energy calculated at ROHF/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -132.458834 |
| Energy at 298.15K | -132.462902 |
| HF Energy | -132.458834 |
| Nuclear repulsion energy | 65.143724 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at ROHF/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3669 |
3158 |
0.54 |
|
|
|
| 2 |
A' |
3427 |
2950 |
6.91 |
|
|
|
| 3 |
A' |
3309 |
2848 |
1.98 |
|
|
|
| 4 |
A' |
3271 |
2815 |
33.36 |
|
|
|
| 5 |
A' |
1823 |
1569 |
62.93 |
|
|
|
| 6 |
A' |
1578 |
1359 |
9.47 |
|
|
|
| 7 |
A' |
1522 |
1310 |
8.30 |
|
|
|
| 8 |
A' |
1381 |
1189 |
90.70 |
|
|
|
| 9 |
A' |
1145 |
986 |
13.29 |
|
|
|
| 10 |
A' |
1020 |
878 |
1.01 |
|
|
|
| 11 |
A' |
520 |
448 |
9.90 |
|
|
|
| 12 |
A" |
1232 |
1060 |
57.97 |
|
|
|
| 13 |
A" |
865 |
745 |
41.39 |
|
|
|
| 14 |
A" |
571 |
492 |
23.48 |
|
|
|
| 15 |
A" |
364 |
313 |
1.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12849.1 cm
-1
Scaled (by 0.8607) Zero Point Vibrational Energy (zpe) 11059.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at ROHF/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.454 |
0.000 |
| C2 |
1.109 |
-0.482 |
0.000 |
| N3 |
-1.230 |
0.168 |
0.000 |
| H4 |
0.264 |
1.511 |
0.000 |
| H5 |
2.130 |
-0.132 |
0.000 |
| H6 |
0.927 |
-1.549 |
0.000 |
| H7 |
-1.364 |
-0.836 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4509 | 1.2625 | 1.0900 | 2.2094 | 2.2067 | 1.8772 |
C2 | 1.4509 | | 2.4273 | 2.1651 | 1.0803 | 1.0822 | 2.4980 | N3 | 1.2625 | 2.4273 | | 2.0089 | 3.3736 | 2.7566 | 1.0131 | H4 | 1.0900 | 2.1651 | 2.0089 | | 2.4864 | 3.1312 | 2.8566 | H5 | 2.2094 | 1.0803 | 3.3736 | 2.4864 | | 1.8598 | 3.5648 | H6 | 2.2067 | 1.0822 | 2.7566 | 3.1312 | 1.8598 | | 2.3991 | H7 | 1.8772 | 2.4980 | 1.0131 | 2.8566 | 3.5648 | 2.3991 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
120.887 |
|
C1 |
C2 |
H6 |
120.480 |
| C1 |
N3 |
H7 |
110.686 |
|
C2 |
C1 |
N3 |
126.760 |
| C2 |
C1 |
H4 |
116.154 |
|
N3 |
C1 |
H4 |
117.086 |
| H5 |
C2 |
H6 |
118.633 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at ROHF/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.031 |
|
|
|
| 2 |
C |
-0.057 |
|
|
|
| 3 |
N |
-0.233 |
|
|
|
| 4 |
H |
0.042 |
|
|
|
| 5 |
H |
0.066 |
|
|
|
| 6 |
H |
0.060 |
|
|
|
| 7 |
H |
0.090 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.220 |
-1.193 |
0.000 |
2.520 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.650 |
2.654 |
-0.002 |
| y |
2.654 |
-16.667 |
-0.001 |
| z |
-0.002 |
-0.001 |
-20.096 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.269 |
2.654 |
-0.002 |
| y |
2.654 |
3.706 |
-0.001 |
| z |
-0.002 |
-0.001 |
-1.437 |
|
| Polar |
|---|
| 3z2-r2 | -2.875 |
|---|
| x2-y2 | -3.984 |
|---|
| xy | 2.654 |
|---|
| xz | -0.002 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
45.471 |
| (<r2>)1/2 |
6.743 |
