Jump to
S1C2
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -168.158463 |
| Energy at 298.15K | |
| HF Energy | -167.639015 |
| Nuclear repulsion energy | 59.699306 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3501 |
3336 |
|
|
|
|
| 2 |
Σ |
2272 |
2165 |
|
|
|
|
| 3 |
Σ |
1266 |
1207 |
|
|
|
|
| 4 |
Π |
546 |
521 |
|
|
|
|
| 4 |
Π |
546 |
521 |
|
|
|
|
| 5 |
Π |
357i |
340i |
|
|
|
|
| 5 |
Π |
357i |
341i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3708.4 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 3534.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
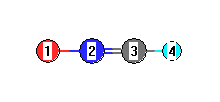
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.196 |
| N2 |
0.000 |
0.000 |
-0.015 |
| C3 |
0.000 |
0.000 |
-1.197 |
| H4 |
0.000 |
0.000 |
-2.273 |
Atom - Atom Distances (Å)
| |
O1 |
N2 |
C3 |
H4 |
| O1 | | 1.2111 | 2.3932 | 3.4684 |
N2 | 1.2111 | | 1.1821 | 2.2573 | C3 | 2.3932 | 1.1821 | | 1.0752 | H4 | 3.4684 | 2.2573 | 1.0752 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
N2 |
C3 |
180.000 |
|
N2 |
C3 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -168.159746 |
| Energy at 298.15K | -168.160515 |
| HF Energy | -167.632277 |
| Nuclear repulsion energy | 59.634340 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3423 |
3262 |
|
|
|
|
| 2 |
A' |
2207 |
2103 |
|
|
|
|
| 3 |
A' |
1277 |
1217 |
|
|
|
|
| 4 |
A' |
538 |
513 |
|
|
|
|
| 5 |
A' |
465 |
443 |
|
|
|
|
| 6 |
A" |
542 |
517 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4225.7 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 4027.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.069 |
-0.521 |
0.000 |
| N2 |
0.000 |
0.037 |
0.000 |
| C3 |
1.130 |
0.434 |
0.000 |
| H4 |
1.771 |
1.305 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
N2 |
C3 |
H4 |
| O1 | | 1.2056 | 2.3966 | 3.3757 |
N2 | 1.2056 | | 1.1972 | 2.1778 | C3 | 2.3966 | 1.1972 | | 1.0816 | H4 | 3.3757 | 2.1778 | 1.0816 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
N2 |
C3 |
171.767 |
|
N2 |
C3 |
H4 |
145.703 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
