Jump to
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -168.137882 |
| Energy at 298.15K | |
| HF Energy | -167.640995 |
| Nuclear repulsion energy | 59.899282 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3520 |
3377 |
248.75 |
|
|
|
| 2 |
Σ |
2321 |
2227 |
297.83 |
|
|
|
| 3 |
Σ |
1269 |
1218 |
107.20 |
|
|
|
| 4 |
Π |
559 |
537 |
0.94 |
|
|
|
| 4 |
Π |
559 |
537 |
0.94 |
|
|
|
| 5 |
Π |
203i |
194i |
77.98 |
|
|
|
| 5 |
Π |
203i |
194i |
77.98 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3911.7 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 3752.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.193 |
| N2 |
0.000 |
0.000 |
-0.018 |
| C3 |
0.000 |
0.000 |
-1.191 |
| H4 |
0.000 |
0.000 |
-2.266 |
Atom - Atom Distances (Å)
| |
O1 |
N2 |
C3 |
H4 |
| O1 | | 1.2109 | 2.3841 | 3.4582 |
N2 | 1.2109 | | 1.1732 | 2.2473 | C3 | 2.3841 | 1.1732 | | 1.0741 | H4 | 3.4582 | 2.2473 | 1.0741 | |
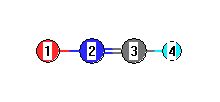 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
N2 |
C3 |
180.000 |
|
N2 |
C3 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -168.138024 |
| Energy at 298.15K | -168.138620 |
| HF Energy | -167.638741 |
| Nuclear repulsion energy | 59.895500 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3493 |
3351 |
204.69 |
|
|
|
| 2 |
A' |
2298 |
2205 |
315.09 |
|
|
|
| 3 |
A' |
1277 |
1225 |
92.86 |
|
|
|
| 4 |
A' |
558 |
535 |
2.97 |
|
|
|
| 5 |
A' |
276 |
265 |
130.82 |
|
|
|
| 6 |
A" |
558 |
535 |
1.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4229.9 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 4058.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.832 |
-0.851 |
0.000 |
| N2 |
0.000 |
0.026 |
0.000 |
| C3 |
0.878 |
0.811 |
0.000 |
| H4 |
1.387 |
1.759 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
N2 |
C3 |
H4 |
| O1 | | 1.2085 | 2.3845 | 3.4259 |
N2 | 1.2085 | | 1.1780 | 2.2202 | C3 | 2.3845 | 1.1780 | | 1.0763 | H4 | 3.4259 | 2.2202 | 1.0763 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
N2 |
C3 |
175.308 |
|
N2 |
C3 |
H4 |
160.023 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
