Jump to
S1C2
Energy calculated at QCISD(T)=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -168.164923 |
| Energy at 298.15K | |
| HF Energy | -167.639225 |
| Nuclear repulsion energy | 59.736375 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3507 |
3363 |
|
|
|
|
| 2 |
Σ |
2276 |
2182 |
|
|
|
|
| 3 |
Σ |
1269 |
1217 |
|
|
|
|
| 4 |
Π |
548 |
526 |
|
|
|
|
| 4 |
Π |
548 |
526 |
|
|
|
|
| 5 |
Π |
351i |
337i |
|
|
|
|
| 5 |
Π |
351i |
337i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3722.8 cm
-1
Scaled (by 0.959) Zero Point Vibrational Energy (zpe) 3570.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)=FULL/cc-pVDZ
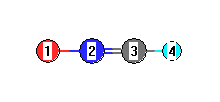
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.195 |
| N2 |
0.000 |
0.000 |
-0.015 |
| C3 |
0.000 |
0.000 |
-1.197 |
| H4 |
0.000 |
0.000 |
-2.271 |
Atom - Atom Distances (Å)
| |
O1 |
N2 |
C3 |
H4 |
| O1 | | 1.2103 | 2.3917 | 3.4662 |
N2 | 1.2103 | | 1.1815 | 2.2559 | C3 | 2.3917 | 1.1815 | | 1.0745 | H4 | 3.4662 | 2.2559 | 1.0745 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
N2 |
C3 |
180.000 |
|
N2 |
C3 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -168.166130 |
| Energy at 298.15K | -168.166896 |
| HF Energy | -167.632695 |
| Nuclear repulsion energy | 59.675643 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3431 |
3290 |
|
|
|
|
| 2 |
A' |
2213 |
2122 |
|
|
|
|
| 3 |
A' |
1279 |
1227 |
|
|
|
|
| 4 |
A' |
540 |
518 |
|
|
|
|
| 5 |
A' |
459 |
440 |
|
|
|
|
| 6 |
A" |
544 |
522 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4232.8 cm
-1
Scaled (by 0.959) Zero Point Vibrational Energy (zpe) 4059.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)=FULL/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.063 |
-0.530 |
0.000 |
| N2 |
0.000 |
0.036 |
0.000 |
| C3 |
1.124 |
0.445 |
0.000 |
| H4 |
1.762 |
1.317 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
N2 |
C3 |
H4 |
| O1 | | 1.2048 | 2.3950 | 3.3757 |
N2 | 1.2048 | | 1.1961 | 2.1784 | C3 | 2.3950 | 1.1961 | | 1.0806 | H4 | 3.3757 | 2.1784 | 1.0806 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
N2 |
C3 |
171.935 |
|
N2 |
C3 |
H4 |
146.162 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
